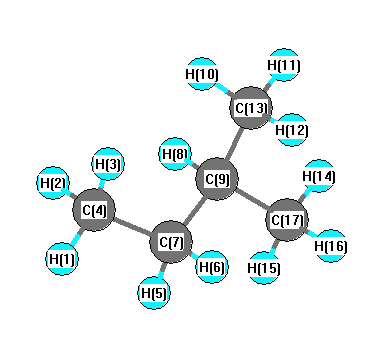
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3135 |
3000 |
31.15 |
|
|
|
| 2 |
A |
3125 |
2991 |
66.84 |
|
|
|
| 3 |
A |
3122 |
2988 |
14.32 |
|
|
|
| 4 |
A |
3120 |
2986 |
53.49 |
|
|
|
| 5 |
A |
3115 |
2982 |
52.13 |
|
|
|
| 6 |
A |
3111 |
2977 |
6.37 |
|
|
|
| 7 |
A |
3069 |
2938 |
21.16 |
|
|
|
| 8 |
A |
3052 |
2921 |
22.55 |
|
|
|
| 9 |
A |
3047 |
2916 |
37.09 |
|
|
|
| 10 |
A |
3043 |
2912 |
32.75 |
|
|
|
| 11 |
A |
3033 |
2903 |
31.80 |
|
|
|
| 12 |
A |
3020 |
2890 |
4.76 |
|
|
|
| 13 |
A |
1514 |
1449 |
5.35 |
|
|
|
| 14 |
A |
1505 |
1441 |
9.76 |
|
|
|
| 15 |
A |
1503 |
1439 |
3.95 |
|
|
|
| 16 |
A |
1496 |
1432 |
11.87 |
|
|
|
| 17 |
A |
1493 |
1429 |
2.42 |
|
|
|
| 18 |
A |
1487 |
1423 |
0.42 |
|
|
|
| 19 |
A |
1479 |
1416 |
0.08 |
|
|
|
| 20 |
A |
1416 |
1355 |
6.40 |
|
|
|
| 21 |
A |
1405 |
1344 |
4.65 |
|
|
|
| 22 |
A |
1399 |
1339 |
7.00 |
|
|
|
| 23 |
A |
1383 |
1324 |
0.79 |
|
|
|
| 24 |
A |
1367 |
1309 |
0.86 |
|
|
|
| 25 |
A |
1322 |
1265 |
1.66 |
|
|
|
| 26 |
A |
1293 |
1238 |
1.29 |
|
|
|
| 27 |
A |
1205 |
1154 |
0.65 |
|
|
|
| 28 |
A |
1200 |
1148 |
0.45 |
|
|
|
| 29 |
A |
1176 |
1126 |
2.34 |
|
|
|
| 30 |
A |
1066 |
1021 |
0.25 |
|
|
|
| 31 |
A |
1029 |
985 |
2.69 |
|
|
|
| 32 |
A |
980 |
938 |
3.52 |
|
|
|
| 33 |
A |
969 |
928 |
0.36 |
|
|
|
| 34 |
A |
930 |
890 |
0.60 |
|
|
|
| 35 |
A |
924 |
884 |
1.02 |
|
|
|
| 36 |
A |
801 |
767 |
0.12 |
|
|
|
| 37 |
A |
767 |
734 |
2.59 |
|
|
|
| 38 |
A |
447 |
428 |
0.17 |
|
|
|
| 39 |
A |
401 |
384 |
0.12 |
|
|
|
| 40 |
A |
361 |
346 |
0.02 |
|
|
|
| 41 |
A |
270 |
259 |
0.01 |
|
|
|
| 42 |
A |
255 |
244 |
0.01 |
|
|
|
| 43 |
A |
220 |
211 |
0.01 |
|
|
|
| 44 |
A |
205 |
196 |
0.03 |
|
|
|
| 45 |
A |
91 |
87 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35176.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 33666.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.120 |
|
|
|
| 2 |
H |
0.114 |
|
|
|
| 3 |
H |
0.117 |
|
|
|
| 4 |
C |
-0.379 |
|
|
|
| 5 |
H |
0.112 |
|
|
|
| 6 |
H |
0.111 |
|
|
|
| 7 |
C |
-0.198 |
|
|
|
| 8 |
H |
0.114 |
|
|
|
| 9 |
C |
-0.065 |
|
|
|
| 10 |
H |
0.120 |
|
|
|
| 11 |
H |
0.117 |
|
|
|
| 12 |
H |
0.113 |
|
|
|
| 13 |
C |
-0.372 |
|
|
|
| 14 |
H |
0.115 |
|
|
|
| 15 |
H |
0.114 |
|
|
|
| 16 |
H |
0.112 |
|
|
|
| 17 |
C |
-0.364 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.032 |
-0.043 |
-0.077 |
0.094 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.618 |
0.290 |
-0.234 |
| y |
0.290 |
-35.351 |
0.080 |
| z |
-0.234 |
0.080 |
-34.757 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.564 |
0.290 |
-0.234 |
| y |
0.290 |
-0.163 |
0.080 |
| z |
-0.234 |
0.080 |
0.727 |
|
| Polar |
|---|
| 3z2-r2 | 1.454 |
|---|
| x2-y2 | -0.267 |
|---|
| xy | 0.290 |
|---|
| xz | -0.234 |
|---|
| yz | 0.080 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.182 |
-0.045 |
0.021 |
| y |
-0.045 |
9.193 |
-0.032 |
| z |
0.021 |
-0.032 |
8.298 |
<r2> (average value of r
2) Å
2
| <r2> |
155.785 |
| (<r2>)1/2 |
12.481 |