return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (1-Buten-3-yne, 2-methyl-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-193.692929
Energy at 298.15K-193.697528
HF Energy-192.815175
Nuclear repulsion energy143.544075
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3498 3321 66.12      
2 A' 3293 3126 3.93      
3 A' 3206 3043 1.68      
4 A' 3181 3020 11.34      
5 A' 3090 2933 13.22      
6 A' 2140 2031 0.06      
7 A' 1688 1603 13.38      
8 A' 1517 1441 11.99      
9 A' 1455 1381 0.16      
10 A' 1417 1345 5.53      
11 A' 1309 1243 9.34      
12 A' 1039 986 3.63      
13 A' 988 938 0.31      
14 A' 788 749 1.28      
15 A' 659 625 41.08      
16 A' 558 530 6.13      
17 A' 395 375 0.21      
18 A' 180 170 1.37      
19 A" 3158 2998 8.07      
20 A" 1502 1426 7.65      
21 A" 1082 1027 0.23      
22 A" 959 910 40.38      
23 A" 760 721 0.19      
24 A" 652 619 39.74      
25 A" 549 521 1.45      
26 A" 276 262 8.26      
27 A" 196 186 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 19765.7 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18765.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.31711 0.13481 0.09626

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.457 0.780 0.000
H2 -1.602 1.855 0.000
H3 -1.941 0.357 0.876
H4 -1.941 0.357 -0.876
C5 0.518 -2.139 0.000
H6 0.703 -3.179 0.000
C7 0.320 -0.943 0.000
C8 0.000 0.446 0.000
C9 0.962 1.371 0.000
H10 2.003 1.097 0.000
H11 0.717 2.420 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 C8 C9 H10 H11
C11.08461.08591.08593.52464.50992.47561.49492.49053.47502.7231
H21.08461.76761.76764.52195.53663.39502.13352.60993.68432.3871
H31.08591.76761.75133.61164.50102.75122.13093.19704.10723.4761
H41.08591.76761.75133.61164.50102.75122.13093.19704.10723.4761
C53.52464.52193.61163.61161.05631.21192.63653.53773.56084.5629
H64.50995.53664.50104.50101.05632.26823.69264.55714.46985.5986
C72.47563.39502.75122.75121.21192.26821.42592.40152.64533.3863
C81.49492.13352.13092.13092.63653.69261.42591.33452.10652.0997
C92.49052.60993.19703.19703.53774.55712.40151.33451.07631.0772
H103.47503.68434.10724.10723.56084.46982.64532.10651.07631.8447
H112.72312.38713.47613.47614.56295.59863.38632.09971.07721.8447

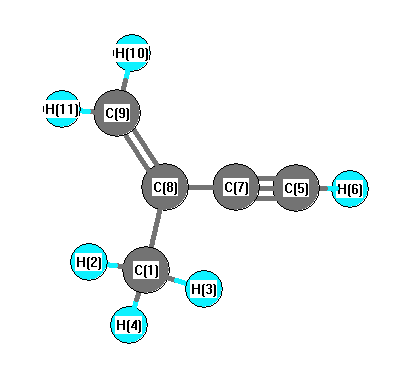
picture of 1-Buten-3-yne, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C8 C7 115.881 C1 C8 C9 123.238
H2 C1 H3 109.045 H2 C1 H4 109.045
H2 C1 C8 110.596 H3 C1 H4 107.479
H3 C1 C8 110.302 H4 C1 C8 110.302
C5 C7 C8 176.434 H6 C5 C7 179.346
C7 C8 C9 120.880 C8 C9 H10 121.433
C8 C9 H11 120.693 H10 C9 H11 117.875
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability