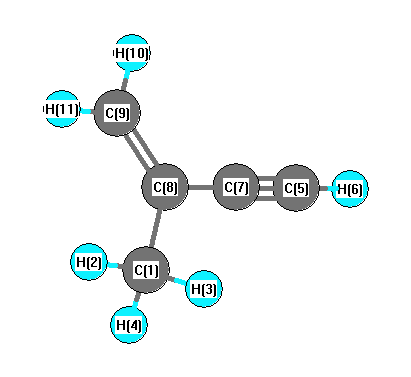
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3490 |
3340 |
70.23 |
|
|
|
| 2 |
A' |
3266 |
3126 |
5.92 |
|
|
|
| 3 |
A' |
3171 |
3035 |
3.02 |
|
|
|
| 4 |
A' |
3152 |
3017 |
14.56 |
|
|
|
| 5 |
A' |
3059 |
2928 |
15.68 |
|
|
|
| 6 |
A' |
2224 |
2129 |
1.83 |
|
|
|
| 7 |
A' |
1702 |
1629 |
16.56 |
|
|
|
| 8 |
A' |
1492 |
1428 |
12.67 |
|
|
|
| 9 |
A' |
1437 |
1376 |
0.20 |
|
|
|
| 10 |
A' |
1406 |
1346 |
5.44 |
|
|
|
| 11 |
A' |
1299 |
1243 |
11.33 |
|
|
|
| 12 |
A' |
1028 |
984 |
3.84 |
|
|
|
| 13 |
A' |
974 |
932 |
0.52 |
|
|
|
| 14 |
A' |
786 |
753 |
1.24 |
|
|
|
| 15 |
A' |
686 |
657 |
41.39 |
|
|
|
| 16 |
A' |
570 |
545 |
9.33 |
|
|
|
| 17 |
A' |
388 |
372 |
0.22 |
|
|
|
| 18 |
A' |
187 |
179 |
1.61 |
|
|
|
| 19 |
A" |
3121 |
2988 |
11.01 |
|
|
|
| 20 |
A" |
1474 |
1411 |
7.51 |
|
|
|
| 21 |
A" |
1069 |
1023 |
0.62 |
|
|
|
| 22 |
A" |
946 |
906 |
40.52 |
|
|
|
| 23 |
A" |
752 |
720 |
0.07 |
|
|
|
| 24 |
A" |
667 |
638 |
43.39 |
|
|
|
| 25 |
A" |
549 |
526 |
1.55 |
|
|
|
| 26 |
A" |
272 |
260 |
8.23 |
|
|
|
| 27 |
A" |
180 |
173 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19673.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 18829.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.326 |
|
|
|
| 2 |
H |
0.110 |
|
|
|
| 3 |
H |
0.127 |
|
|
|
| 4 |
H |
0.127 |
|
|
|
| 5 |
C |
-0.290 |
|
|
|
| 6 |
H |
0.148 |
|
|
|
| 7 |
C |
0.076 |
|
|
|
| 8 |
C |
0.099 |
|
|
|
| 9 |
C |
-0.335 |
|
|
|
| 10 |
H |
0.136 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.386 |
0.369 |
0.000 |
0.534 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.418 |
-0.925 |
0.000 |
| y |
-0.925 |
-25.185 |
0.000 |
| z |
0.000 |
0.000 |
-32.684 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.483 |
-0.925 |
0.000 |
| y |
-0.925 |
5.866 |
0.000 |
| z |
0.000 |
0.000 |
-5.383 |
|
| Polar |
|---|
| 3z2-r2 | -10.766 |
|---|
| x2-y2 | -4.233 |
|---|
| xy | -0.925 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.122 |
0.269 |
0.000 |
| y |
0.269 |
12.221 |
0.000 |
| z |
0.000 |
0.000 |
5.169 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |