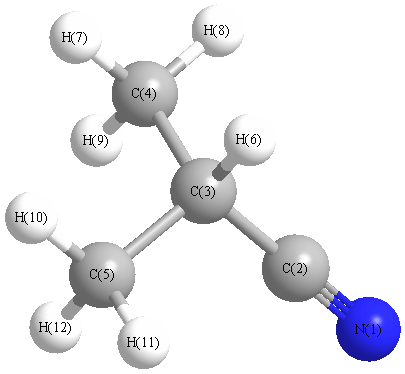
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3150 |
3021 |
17.78 |
|
|
|
| 2 |
A' |
3143 |
3015 |
29.28 |
|
|
|
| 3 |
A' |
3065 |
2940 |
15.28 |
|
|
|
| 4 |
A' |
3058 |
2933 |
2.11 |
|
|
|
| 5 |
A' |
2377 |
2280 |
11.67 |
|
|
|
| 6 |
A' |
1512 |
1450 |
10.05 |
|
|
|
| 7 |
A' |
1501 |
1440 |
9.66 |
|
|
|
| 8 |
A' |
1424 |
1366 |
3.46 |
|
|
|
| 9 |
A' |
1354 |
1299 |
5.65 |
|
|
|
| 10 |
A' |
1200 |
1151 |
3.12 |
|
|
|
| 11 |
A' |
1130 |
1084 |
4.60 |
|
|
|
| 12 |
A' |
941 |
903 |
1.00 |
|
|
|
| 13 |
A' |
781 |
749 |
0.51 |
|
|
|
| 14 |
A' |
557 |
534 |
1.26 |
|
|
|
| 15 |
A' |
349 |
335 |
0.22 |
|
|
|
| 16 |
A' |
281 |
269 |
1.06 |
|
|
|
| 17 |
A' |
217 |
208 |
1.93 |
|
|
|
| 18 |
A" |
3148 |
3020 |
11.29 |
|
|
|
| 19 |
A" |
3140 |
3011 |
0.19 |
|
|
|
| 20 |
A" |
3063 |
2938 |
14.29 |
|
|
|
| 21 |
A" |
1490 |
1429 |
3.39 |
|
|
|
| 22 |
A" |
1488 |
1427 |
0.98 |
|
|
|
| 23 |
A" |
1403 |
1346 |
5.00 |
|
|
|
| 24 |
A" |
1325 |
1271 |
0.25 |
|
|
|
| 25 |
A" |
1141 |
1094 |
2.56 |
|
|
|
| 26 |
A" |
979 |
939 |
0.27 |
|
|
|
| 27 |
A" |
940 |
901 |
2.22 |
|
|
|
| 28 |
A" |
582 |
558 |
0.00 |
|
|
|
| 29 |
A" |
222 |
213 |
0.02 |
|
|
|
| 30 |
A" |
188 |
180 |
4.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22573.3 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 21652.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.087 |
|
|
|
| 2 |
C |
-0.068 |
|
|
|
| 3 |
C |
-0.065 |
|
|
|
| 4 |
C |
-0.236 |
|
|
|
| 5 |
C |
-0.236 |
|
|
|
| 6 |
H |
0.114 |
|
|
|
| 7 |
H |
0.089 |
|
|
|
| 8 |
H |
0.102 |
|
|
|
| 9 |
H |
0.098 |
|
|
|
| 10 |
H |
0.089 |
|
|
|
| 11 |
H |
0.102 |
|
|
|
| 12 |
H |
0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.164 |
3.923 |
0.000 |
4.092 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.438 |
1.722 |
0.000 |
| y |
1.722 |
-39.191 |
0.000 |
| z |
0.000 |
0.000 |
-29.934 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.125 |
1.722 |
0.000 |
| y |
1.722 |
-9.005 |
0.000 |
| z |
0.000 |
0.000 |
4.880 |
|
| Polar |
|---|
| 3z2-r2 | 9.760 |
|---|
| x2-y2 | 8.753 |
|---|
| xy | 1.722 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.297 |
-0.547 |
0.000 |
| y |
-0.547 |
8.841 |
0.000 |
| z |
0.000 |
0.000 |
7.124 |
<r2> (average value of r
2) Å
2
| <r2> |
127.900 |
| (<r2>)1/2 |
11.309 |