return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (1-Propanol, 2-methyl-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-233.269402
Energy at 298.15K-233.280663
HF Energy-232.237693
Nuclear repulsion energy194.573998
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3887 3690 29.43      
2 A 3163 3003 27.08      
3 A 3159 2999 18.77      
4 A 3152 2992 46.85      
5 A 3146 2987 14.72      
6 A 3137 2979 20.69      
7 A 3097 2940 16.02      
8 A 3079 2923 17.97      
9 A 3073 2918 18.18      
10 A 3035 2882 40.88      
11 A 1541 1463 14.51      
12 A 1530 1452 5.85      
13 A 1525 1448 0.24      
14 A 1518 1441 4.27      
15 A 1510 1434 0.78      
16 A 1434 1362 23.91      
17 A 1418 1346 2.35      
18 A 1410 1339 15.29      
19 A 1404 1333 4.12      
20 A 1380 1310 4.23      
21 A 1338 1270 0.11      
22 A 1261 1198 11.38      
23 A 1216 1155 0.16      
24 A 1178 1119 1.83      
25 A 1136 1079 3.17      
26 A 1097 1042 119.36      
27 A 989 939 0.45      
28 A 968 919 6.58      
29 A 937 890 0.47      
30 A 914 868 7.60      
31 A 841 798 6.35      
32 A 498 473 5.38      
33 A 417 396 0.95      
34 A 355 337 3.08      
35 A 287 272 56.18      
36 A 265 251 1.20      
37 A 242 229 53.90      
38 A 222 211 8.97      
39 A 126 120 3.56      

Unscaled Zero Point Vibrational Energy (zpe) 30441.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 28901.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.25688 0.11896 0.08967

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.446 0.004 -0.354
C2 -0.763 -0.692 0.236
C3 1.704 -0.750 0.030
C4 0.519 1.446 0.104
O5 -1.991 -0.117 -0.157
H6 0.332 -0.015 -1.436
H7 1.661 -1.784 -0.301
H8 2.581 -0.289 -0.412
H9 1.834 -0.747 1.110
H10 -0.359 2.006 -0.204
H11 0.599 1.495 1.189
H12 1.389 1.941 -0.316
H13 -0.787 -1.720 -0.109
H14 -0.659 -0.714 1.324
H15 -2.095 0.710 0.314

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.51391.51601.51522.44781.08752.16232.15662.15272.16312.15182.15492.13342.13362.7201
C21.51392.47552.49571.41242.10992.71213.43002.73992.76292.74633.44431.08541.09281.9355
C31.51602.47552.49593.75352.13761.08651.08561.08773.45012.75772.73072.67692.69424.0792
C41.51522.49572.49592.96872.13103.44952.74402.74711.08611.08861.08573.43162.74592.7233
O52.44781.41243.75352.96872.65394.01744.58314.07842.67843.33443.96062.00562.07960.9577
H61.08752.10992.13762.13102.65392.48652.48633.04472.46583.04002.48912.43293.01443.0784
H72.16232.71211.08653.44954.01742.48651.75891.75934.29563.75513.73432.45673.02804.5502
H82.15663.43001.08562.74404.58312.48631.75891.75673.73573.11132.53013.67263.70104.8365
H92.15272.73991.08772.74714.07843.04471.75931.75673.75642.56103.07453.05022.50234.2649
H102.16312.76293.45011.08612.67842.46584.29563.73573.75641.76551.75293.75183.13382.2270
H112.15182.74632.75771.08863.33443.04003.75513.11132.56101.76551.75773.73432.54552.9385
H122.15493.44432.73071.08573.96062.48913.73432.53013.07451.75291.75774.26383.73253.7480
H132.13341.08542.67693.43162.00562.43292.45673.67263.05023.75183.73434.26381.75612.7921
H142.13361.09282.69422.74592.07963.01443.02803.70102.50233.13382.54553.73251.75612.2603
H152.72011.93554.07922.72330.95773.07844.55024.83654.26492.22702.93853.74802.79212.2603

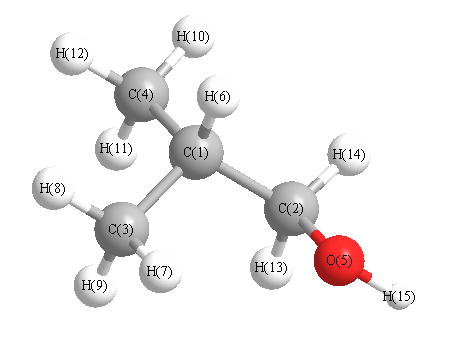
picture of 1-Propanol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O5 113.497 C1 C2 H13 109.214
C1 C2 H14 108.797 C1 C3 H7 111.302
C1 C3 H8 110.901 C1 C3 H9 110.460
C1 C4 H10 111.458 C1 C4 H11 110.388
C1 C4 H12 110.812 C2 C1 C3 109.580
C2 C1 C4 110.961 C2 C1 H6 107.264
C2 O5 H15 107.957 C3 C1 C4 110.854
C3 C1 H6 109.277 C4 C1 H6 108.819
O5 C2 H13 106.079 O5 C2 H14 111.583
H7 C3 H8 108.156 H7 C3 H9 108.034
H8 C3 H9 107.863 H10 C4 H11 108.548
H10 C4 H12 107.638 H11 C4 H12 107.872
H13 C2 H14 107.456
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability