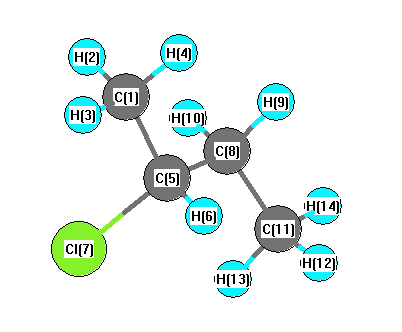
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3155 |
3020 |
17.44 |
|
|
|
| 2 |
A |
3146 |
3012 |
21.93 |
|
|
|
| 3 |
A |
3134 |
3000 |
27.84 |
|
|
|
| 4 |
A |
3130 |
2996 |
25.94 |
|
|
|
| 5 |
A |
3105 |
2971 |
10.16 |
|
|
|
| 6 |
A |
3084 |
2951 |
2.64 |
|
|
|
| 7 |
A |
3060 |
2929 |
15.83 |
|
|
|
| 8 |
A |
3059 |
2928 |
30.10 |
|
|
|
| 9 |
A |
3044 |
2913 |
18.70 |
|
|
|
| 10 |
A |
1508 |
1443 |
7.04 |
|
|
|
| 11 |
A |
1499 |
1435 |
8.12 |
|
|
|
| 12 |
A |
1495 |
1431 |
1.66 |
|
|
|
| 13 |
A |
1484 |
1421 |
9.66 |
|
|
|
| 14 |
A |
1471 |
1408 |
0.99 |
|
|
|
| 15 |
A |
1417 |
1357 |
5.35 |
|
|
|
| 16 |
A |
1409 |
1348 |
7.49 |
|
|
|
| 17 |
A |
1394 |
1334 |
0.52 |
|
|
|
| 18 |
A |
1324 |
1267 |
7.04 |
|
|
|
| 19 |
A |
1316 |
1259 |
9.09 |
|
|
|
| 20 |
A |
1261 |
1207 |
12.70 |
|
|
|
| 21 |
A |
1184 |
1133 |
7.74 |
|
|
|
| 22 |
A |
1140 |
1092 |
2.12 |
|
|
|
| 23 |
A |
1096 |
1049 |
2.97 |
|
|
|
| 24 |
A |
1052 |
1007 |
2.74 |
|
|
|
| 25 |
A |
1007 |
963 |
9.26 |
|
|
|
| 26 |
A |
972 |
930 |
5.93 |
|
|
|
| 27 |
A |
861 |
824 |
9.22 |
|
|
|
| 28 |
A |
796 |
762 |
9.98 |
|
|
|
| 29 |
A |
626 |
599 |
22.04 |
|
|
|
| 30 |
A |
457 |
437 |
1.00 |
|
|
|
| 31 |
A |
386 |
369 |
2.16 |
|
|
|
| 32 |
A |
317 |
304 |
1.17 |
|
|
|
| 33 |
A |
246 |
235 |
0.11 |
|
|
|
| 34 |
A |
234 |
224 |
0.76 |
|
|
|
| 35 |
A |
223 |
213 |
0.32 |
|
|
|
| 36 |
A |
116 |
111 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27102.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 25940.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.341 |
|
|
|
| 2 |
H |
0.129 |
|
|
|
| 3 |
H |
0.135 |
|
|
|
| 4 |
H |
0.114 |
|
|
|
| 5 |
C |
-0.051 |
|
|
|
| 6 |
H |
0.150 |
|
|
|
| 7 |
Cl |
-0.183 |
|
|
|
| 8 |
C |
-0.205 |
|
|
|
| 9 |
H |
0.117 |
|
|
|
| 10 |
H |
0.133 |
|
|
|
| 11 |
C |
-0.360 |
|
|
|
| 12 |
H |
0.111 |
|
|
|
| 13 |
H |
0.132 |
|
|
|
| 14 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.764 |
1.992 |
0.362 |
2.164 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.758 |
-1.163 |
-0.320 |
| y |
-1.163 |
-40.238 |
0.380 |
| z |
-0.320 |
0.380 |
-39.055 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.112 |
-1.163 |
-0.320 |
| y |
-1.163 |
-0.831 |
0.380 |
| z |
-0.320 |
0.380 |
0.943 |
|
| Polar |
|---|
| 3z2-r2 | 1.885 |
|---|
| x2-y2 | 0.479 |
|---|
| xy | -1.163 |
|---|
| xz | -0.320 |
|---|
| yz | 0.380 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.715 |
0.092 |
0.054 |
| y |
0.092 |
9.812 |
0.125 |
| z |
0.054 |
0.125 |
7.624 |
<r2> (average value of r
2) Å
2
| <r2> |
179.180 |
| (<r2>)1/2 |
13.386 |