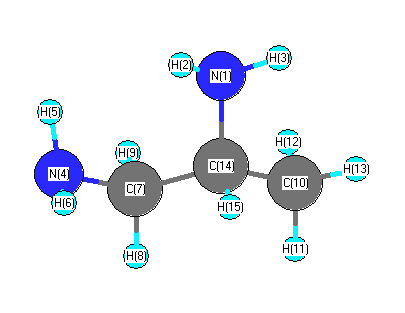
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3621 |
3466 |
0.80 |
|
|
|
| 2 |
A |
3609 |
3454 |
0.61 |
|
|
|
| 3 |
A |
3531 |
3379 |
0.97 |
|
|
|
| 4 |
A |
3530 |
3378 |
0.33 |
|
|
|
| 5 |
A |
3134 |
2999 |
28.04 |
|
|
|
| 6 |
A |
3121 |
2987 |
31.09 |
|
|
|
| 7 |
A |
3095 |
2963 |
27.22 |
|
|
|
| 8 |
A |
3052 |
2922 |
42.10 |
|
|
|
| 9 |
A |
3048 |
2917 |
21.58 |
|
|
|
| 10 |
A |
2937 |
2811 |
75.69 |
|
|
|
| 11 |
A |
1656 |
1585 |
3.38 |
|
|
|
| 12 |
A |
1645 |
1574 |
57.66 |
|
|
|
| 13 |
A |
1500 |
1435 |
3.04 |
|
|
|
| 14 |
A |
1489 |
1425 |
6.58 |
|
|
|
| 15 |
A |
1475 |
1411 |
0.71 |
|
|
|
| 16 |
A |
1417 |
1356 |
1.26 |
|
|
|
| 17 |
A |
1403 |
1343 |
16.05 |
|
|
|
| 18 |
A |
1393 |
1333 |
3.18 |
|
|
|
| 19 |
A |
1372 |
1313 |
8.45 |
|
|
|
| 20 |
A |
1340 |
1282 |
7.22 |
|
|
|
| 21 |
A |
1263 |
1209 |
1.06 |
|
|
|
| 22 |
A |
1206 |
1154 |
2.95 |
|
|
|
| 23 |
A |
1178 |
1128 |
6.37 |
|
|
|
| 24 |
A |
1125 |
1077 |
16.39 |
|
|
|
| 25 |
A |
1048 |
1003 |
9.80 |
|
|
|
| 26 |
A |
1028 |
984 |
1.07 |
|
|
|
| 27 |
A |
959 |
918 |
5.34 |
|
|
|
| 28 |
A |
896 |
858 |
63.15 |
|
|
|
| 29 |
A |
877 |
840 |
64.12 |
|
|
|
| 30 |
A |
832 |
797 |
89.16 |
|
|
|
| 31 |
A |
788 |
754 |
110.87 |
|
|
|
| 32 |
A |
490 |
469 |
1.30 |
|
|
|
| 33 |
A |
460 |
440 |
18.12 |
|
|
|
| 34 |
A |
359 |
344 |
11.30 |
|
|
|
| 35 |
A |
344 |
329 |
38.89 |
|
|
|
| 36 |
A |
249 |
239 |
6.32 |
|
|
|
| 37 |
A |
233 |
223 |
28.85 |
|
|
|
| 38 |
A |
217 |
207 |
17.11 |
|
|
|
| 39 |
A |
125 |
119 |
3.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30522.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29212.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.351 |
|
|
|
| 2 |
H |
0.130 |
|
|
|
| 3 |
H |
0.144 |
|
|
|
| 4 |
N |
-0.370 |
|
|
|
| 5 |
H |
0.151 |
|
|
|
| 6 |
H |
0.144 |
|
|
|
| 7 |
C |
-0.121 |
|
|
|
| 8 |
H |
0.111 |
|
|
|
| 9 |
H |
0.116 |
|
|
|
| 10 |
C |
-0.365 |
|
|
|
| 11 |
H |
0.110 |
|
|
|
| 12 |
H |
0.122 |
|
|
|
| 13 |
H |
0.110 |
|
|
|
| 14 |
C |
-0.025 |
|
|
|
| 15 |
H |
0.094 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.875 |
1.010 |
1.569 |
2.061 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.535 |
-2.464 |
-0.805 |
| y |
-2.464 |
-31.332 |
1.422 |
| z |
-0.805 |
1.422 |
-33.395 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.172 |
-2.464 |
-0.805 |
| y |
-2.464 |
3.133 |
1.422 |
| z |
-0.805 |
1.422 |
0.038 |
|
| Polar |
|---|
| 3z2-r2 | 0.077 |
|---|
| x2-y2 | -4.203 |
|---|
| xy | -2.464 |
|---|
| xz | -0.805 |
|---|
| yz | 1.422 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.846 |
-0.205 |
-0.071 |
| y |
-0.205 |
8.128 |
0.020 |
| z |
-0.071 |
0.020 |
7.189 |
<r2> (average value of r
2) Å
2
| <r2> |
141.964 |
| (<r2>)1/2 |
11.915 |