Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -232.432679 |
| Energy at 298.15K | -232.440981 |
| Nuclear repulsion energy | 177.273582 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3177 |
3041 |
9.00 |
|
|
|
| 2 |
A |
3151 |
3016 |
20.78 |
|
|
|
| 3 |
A |
3147 |
3012 |
19.71 |
|
|
|
| 4 |
A |
3125 |
2991 |
8.41 |
|
|
|
| 5 |
A |
3073 |
2941 |
21.64 |
|
|
|
| 6 |
A |
3073 |
2941 |
7.24 |
|
|
|
| 7 |
A |
3061 |
2929 |
7.42 |
|
|
|
| 8 |
A |
3044 |
2914 |
16.15 |
|
|
|
| 9 |
A |
1828 |
1749 |
154.13 |
|
|
|
| 10 |
A |
1503 |
1438 |
8.70 |
|
|
|
| 11 |
A |
1500 |
1436 |
7.83 |
|
|
|
| 12 |
A |
1475 |
1412 |
9.89 |
|
|
|
| 13 |
A |
1465 |
1402 |
21.96 |
|
|
|
| 14 |
A |
1450 |
1387 |
2.78 |
|
|
|
| 15 |
A |
1420 |
1359 |
6.91 |
|
|
|
| 16 |
A |
1392 |
1333 |
62.66 |
|
|
|
| 17 |
A |
1369 |
1310 |
9.98 |
|
|
|
| 18 |
A |
1282 |
1227 |
0.00 |
|
|
|
| 19 |
A |
1198 |
1146 |
64.27 |
|
|
|
| 20 |
A |
1131 |
1083 |
0.20 |
|
|
|
| 21 |
A |
1118 |
1070 |
0.87 |
|
|
|
| 22 |
A |
1011 |
967 |
3.95 |
|
|
|
| 23 |
A |
948 |
907 |
3.77 |
|
|
|
| 24 |
A |
945 |
904 |
8.94 |
|
|
|
| 25 |
A |
777 |
744 |
3.38 |
|
|
|
| 26 |
A |
751 |
719 |
2.91 |
|
|
|
| 27 |
A |
594 |
568 |
9.14 |
|
|
|
| 28 |
A |
473 |
452 |
0.02 |
|
|
|
| 29 |
A |
399 |
382 |
4.39 |
|
|
|
| 30 |
A |
257 |
246 |
4.67 |
|
|
|
| 31 |
A |
249 |
238 |
0.22 |
|
|
|
| 32 |
A |
106 |
101 |
0.07 |
|
|
|
| 33 |
A |
34 |
33 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24761.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23699.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.876 |
-0.489 |
0.000 |
| C2 |
-0.521 |
0.166 |
-0.000 |
| C3 |
0.674 |
-0.756 |
-0.001 |
| C4 |
1.996 |
-0.022 |
0.001 |
| O5 |
-0.396 |
1.364 |
-0.000 |
| H6 |
-2.653 |
0.267 |
-0.003 |
| H7 |
-1.985 |
-1.132 |
-0.872 |
| H8 |
-1.987 |
-1.126 |
0.878 |
| H9 |
2.829 |
-0.720 |
0.000 |
| H10 |
2.081 |
0.620 |
-0.872 |
| H11 |
2.080 |
0.617 |
0.876 |
| H12 |
0.583 |
-1.417 |
0.865 |
| H13 |
0.584 |
-1.415 |
-0.868 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5050 | 2.5644 | 3.9002 | 2.3717 | 1.0843 | 1.0895 | 1.0895 | 4.7113 | 4.2015 | 4.2006 | 2.7667 | 2.7681 |
C2 | 1.5050 | | 1.5099 | 2.5241 | 1.2050 | 2.1342 | 2.1421 | 2.1416 | 3.4657 | 2.7819 | 2.7820 | 2.1149 | 2.1148 | C3 | 2.5644 | 1.5099 | | 1.5122 | 2.3756 | 3.4812 | 2.8236 | 2.8265 | 2.1554 | 2.1524 | 2.1524 | 1.0927 | 1.0927 | C4 | 3.9002 | 2.5241 | 1.5122 | | 2.7648 | 4.6579 | 4.2243 | 4.2248 | 1.0874 | 1.0868 | 1.0868 | 2.1659 | 2.1658 | O5 | 2.3717 | 1.2050 | 2.3756 | 2.7648 | | 2.5091 | 3.0851 | 3.0822 | 3.8406 | 2.7300 | 2.7311 | 3.0731 | 3.0719 | H6 | 1.0843 | 2.1342 | 3.4812 | 4.6579 | 2.5091 | | 1.7779 | 1.7777 | 5.5706 | 4.8262 | 4.8268 | 3.7497 | 3.7488 | H7 | 1.0895 | 2.1421 | 2.8236 | 4.2243 | 3.0851 | 1.7779 | | 1.7500 | 4.9103 | 4.4278 | 4.7587 | 3.1130 | 2.5842 | H8 | 1.0895 | 2.1416 | 2.8265 | 4.2248 | 3.0822 | 1.7777 | 1.7500 | | 4.9121 | 4.7600 | 4.4248 | 2.5859 | 3.1207 | H9 | 4.7113 | 3.4657 | 2.1554 | 1.0874 | 3.8406 | 5.5706 | 4.9103 | 4.9121 | | 1.7654 | 1.7654 | 2.5061 | 2.5061 | H10 | 4.2015 | 2.7819 | 2.1524 | 1.0868 | 2.7300 | 4.8262 | 4.4278 | 4.7600 | 1.7654 | | 1.7480 | 3.0679 | 2.5264 | H11 | 4.2006 | 2.7820 | 2.1524 | 1.0868 | 2.7311 | 4.8268 | 4.7587 | 4.4248 | 1.7654 | 1.7480 | | 2.5265 | 3.0679 | H12 | 2.7667 | 2.1149 | 1.0927 | 2.1659 | 3.0731 | 3.7497 | 3.1130 | 2.5859 | 2.5061 | 3.0679 | 2.5265 | | 1.7328 | H13 | 2.7681 | 2.1148 | 1.0927 | 2.1658 | 3.0719 | 3.7488 | 2.5842 | 3.1207 | 2.5061 | 2.5264 | 3.0679 | 1.7328 | |
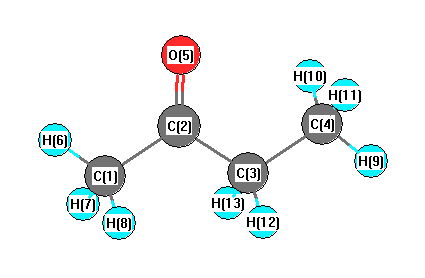 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
116.555 |
|
C1 |
C2 |
O5 |
121.744 |
| C2 |
C1 |
H6 |
109.960 |
|
C2 |
C1 |
H7 |
110.276 |
| C2 |
C1 |
H8 |
110.237 |
|
C2 |
C3 |
C4 |
113.281 |
| C2 |
C3 |
H12 |
107.626 |
|
C2 |
C3 |
H13 |
107.613 |
| C3 |
C2 |
O5 |
121.701 |
|
C3 |
C4 |
H9 |
110.966 |
| C3 |
C4 |
H10 |
110.765 |
|
C3 |
C4 |
H11 |
110.764 |
| C4 |
C3 |
H12 |
111.484 |
|
C4 |
C3 |
H13 |
111.478 |
| H6 |
C1 |
H7 |
109.745 |
|
H6 |
C1 |
H8 |
109.721 |
| H7 |
C1 |
H8 |
106.852 |
|
H9 |
C4 |
H10 |
108.580 |
| H9 |
C4 |
H11 |
108.576 |
|
H10 |
C4 |
H11 |
107.067 |
| H12 |
C3 |
H13 |
104.910 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.351 |
|
|
|
| 2 |
C |
0.210 |
|
|
|
| 3 |
C |
-0.224 |
|
|
|
| 4 |
C |
-0.362 |
|
|
|
| 5 |
O |
-0.265 |
|
|
|
| 6 |
H |
0.135 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.114 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.124 |
|
|
|
| 13 |
H |
0.124 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.238 |
-2.779 |
0.001 |
2.790 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.956 |
1.077 |
-0.002 |
| y |
1.077 |
-34.949 |
0.000 |
| z |
-0.002 |
0.000 |
-30.473 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.755 |
1.077 |
-0.002 |
| y |
1.077 |
-4.734 |
0.000 |
| z |
-0.002 |
0.000 |
1.979 |
|
| Polar |
|---|
| 3z2-r2 | 3.958 |
|---|
| x2-y2 | 4.992 |
|---|
| xy | 1.077 |
|---|
| xz | -0.002 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.590 |
0.212 |
-0.000 |
| y |
0.212 |
7.724 |
0.000 |
| z |
-0.000 |
0.000 |
6.075 |
<r2> (average value of r
2) Å
2
| <r2> |
135.167 |
| (<r2>)1/2 |
11.626 |
