Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1071.843263 |
| Energy at 298.15K | |
| HF Energy | -1070.833585 |
| Nuclear repulsion energy | 263.301699 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.394 |
-0.020 |
0.000 |
| C2 |
-0.172 |
1.394 |
0.000 |
| H3 |
1.478 |
-0.006 |
0.000 |
| Cl4 |
-0.172 |
-0.842 |
1.474 |
| Cl5 |
-0.172 |
-0.842 |
-1.474 |
| O6 |
0.537 |
2.369 |
0.000 |
| H7 |
-1.274 |
1.442 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5225 | 1.0844 | 1.7803 | 1.7803 | 2.3933 | 2.2178 |
C2 | 1.5225 | | 2.1634 | 2.6781 | 2.6781 | 1.2057 | 1.1030 | H3 | 1.0844 | 2.1634 | | 2.3653 | 2.3653 | 2.5546 | 3.1095 | Cl4 | 1.7803 | 2.6781 | 2.3653 | | 2.9487 | 3.6039 | 2.9338 | Cl5 | 1.7803 | 2.6781 | 2.3653 | 2.9487 | | 3.6039 | 2.9338 | O6 | 2.3933 | 1.2057 | 2.5546 | 3.6039 | 3.6039 | | 2.0340 | H7 | 2.2178 | 1.1030 | 3.1095 | 2.9338 | 2.9338 | 2.0340 | |
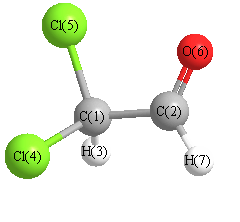 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.198 |
|
C1 |
C2 |
H7 |
114.332 |
| C2 |
C1 |
H3 |
111.058 |
|
C2 |
C1 |
Cl4 |
108.110 |
| C2 |
C1 |
Cl5 |
108.110 |
|
H3 |
C1 |
Cl4 |
108.879 |
| H3 |
C1 |
Cl5 |
108.879 |
|
Cl4 |
C1 |
Cl5 |
111.818 |
| O6 |
C2 |
H7 |
123.470 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1071.840269 |
| Energy at 298.15K | |
| HF Energy | -1070.830076 |
| Nuclear repulsion energy | 266.072569 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3120 |
3042 |
|
|
|
|
| 2 |
A |
2973 |
2898 |
|
|
|
|
| 3 |
A |
1806 |
1760 |
|
|
|
|
| 4 |
A |
1402 |
1367 |
|
|
|
|
| 5 |
A |
1272 |
1240 |
|
|
|
|
| 6 |
A |
1223 |
1192 |
|
|
|
|
| 7 |
A |
1037 |
1011 |
|
|
|
|
| 8 |
A |
943 |
919 |
|
|
|
|
| 9 |
A |
828 |
807 |
|
|
|
|
| 10 |
A |
652 |
636 |
|
|
|
|
| 11 |
A |
620 |
604 |
|
|
|
|
| 12 |
A |
343 |
334 |
|
|
|
|
| 13 |
A |
273 |
266 |
|
|
|
|
| 14 |
A |
218 |
212 |
|
|
|
|
| 15 |
A |
82 |
80 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8395.1 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 8183.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.100 |
-0.022 |
0.525 |
| C2 |
-0.704 |
-1.283 |
0.199 |
| H3 |
0.230 |
0.065 |
1.602 |
| Cl4 |
1.733 |
-0.254 |
-0.170 |
| Cl5 |
-0.681 |
1.446 |
-0.058 |
| O6 |
-1.789 |
-1.286 |
-0.320 |
| H7 |
-0.179 |
-2.208 |
0.494 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5311 | 1.0881 | 1.7899 | 1.7615 | 2.4246 | 2.2045 |
C2 | 1.5311 | | 2.1579 | 2.6708 | 2.7414 | 1.2024 | 1.1038 | H3 | 1.0881 | 2.1579 | | 2.3453 | 2.3436 | 3.0967 | 2.5612 | Cl4 | 1.7899 | 2.6708 | 2.3453 | | 2.9543 | 3.6729 | 2.8131 | Cl5 | 1.7615 | 2.7414 | 2.3436 | 2.9543 | | 2.9595 | 3.7295 | O6 | 2.4246 | 1.2024 | 3.0967 | 3.6729 | 2.9595 | | 2.0261 | H7 | 2.2045 | 1.1038 | 2.5612 | 2.8131 | 3.7295 | 2.0261 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.558 |
|
C1 |
C2 |
H7 |
112.566 |
| C2 |
C1 |
H3 |
109.799 |
|
C2 |
C1 |
Cl4 |
106.813 |
| C2 |
C1 |
Cl5 |
112.542 |
|
H3 |
C1 |
Cl4 |
106.597 |
| H3 |
C1 |
Cl5 |
108.330 |
|
Cl4 |
C1 |
Cl5 |
112.581 |
| O6 |
C2 |
H7 |
122.876 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
