Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1071.874152 |
| Energy at 298.15K | -1071.876718 |
| HF Energy | -1070.833968 |
| Nuclear repulsion energy | 265.334147 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3210 |
3048 |
1.35 |
|
|
|
| 2 |
A' |
3025 |
2872 |
35.69 |
|
|
|
| 3 |
A' |
1789 |
1698 |
112.26 |
|
|
|
| 4 |
A' |
1420 |
1348 |
1.28 |
|
|
|
| 5 |
A' |
1236 |
1174 |
6.28 |
|
|
|
| 6 |
A' |
1102 |
1046 |
25.06 |
|
|
|
| 7 |
A' |
813 |
772 |
18.84 |
|
|
|
| 8 |
A' |
450 |
427 |
4.06 |
|
|
|
| 9 |
A' |
324 |
308 |
21.30 |
|
|
|
| 10 |
A' |
259 |
246 |
1.92 |
|
|
|
| 11 |
A" |
1259 |
1196 |
20.57 |
|
|
|
| 12 |
A" |
1034 |
982 |
20.16 |
|
|
|
| 13 |
A" |
781 |
741 |
96.12 |
|
|
|
| 14 |
A" |
291 |
276 |
3.31 |
|
|
|
| 15 |
A" |
81 |
77 |
7.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8537.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 8105.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.338 |
0.207 |
0.000 |
| C2 |
-0.927 |
1.032 |
0.000 |
| H3 |
1.214 |
0.836 |
0.000 |
| Cl4 |
0.338 |
-0.782 |
1.461 |
| Cl5 |
0.338 |
-0.782 |
-1.461 |
| O6 |
-0.916 |
2.236 |
0.000 |
| H7 |
-1.846 |
0.434 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5102 | 1.0781 | 1.7649 | 1.7649 | 2.3850 | 2.1958 |
C2 | 1.5102 | | 2.1498 | 2.6511 | 2.6511 | 1.2038 | 1.0966 | H3 | 1.0781 | 2.1498 | | 2.3497 | 2.3497 | 2.5488 | 3.0862 | Cl4 | 1.7649 | 2.6511 | 2.3497 | | 2.9228 | 3.5802 | 2.8957 | Cl5 | 1.7649 | 2.6511 | 2.3497 | 2.9228 | | 3.5802 | 2.8957 | O6 | 2.3850 | 1.2038 | 2.5488 | 3.5802 | 3.5802 | | 2.0278 | H7 | 2.1958 | 1.0966 | 3.0862 | 2.8957 | 2.8957 | 2.0278 | |
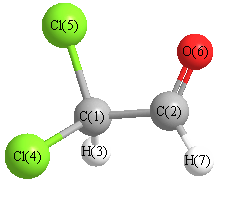 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.585 |
|
C1 |
C2 |
H7 |
113.830 |
| C2 |
C1 |
H3 |
111.220 |
|
C2 |
C1 |
Cl4 |
107.835 |
| C2 |
C1 |
Cl5 |
107.835 |
|
H3 |
C1 |
Cl4 |
109.080 |
| H3 |
C1 |
Cl5 |
109.080 |
|
Cl4 |
C1 |
Cl5 |
111.799 |
| O6 |
C2 |
H7 |
123.586 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1071.871242 |
| Energy at 298.15K | |
| HF Energy | -1070.830476 |
| Nuclear repulsion energy | 268.109053 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3160 |
3000 |
4.73 |
75.40 |
0.17 |
0.30 |
| 2 |
A |
3018 |
2865 |
49.15 |
131.24 |
0.28 |
0.43 |
| 3 |
A |
1799 |
1708 |
93.68 |
10.44 |
0.52 |
0.68 |
| 4 |
A |
1420 |
1348 |
10.26 |
4.58 |
0.32 |
0.49 |
| 5 |
A |
1284 |
1219 |
12.03 |
2.41 |
0.64 |
0.78 |
| 6 |
A |
1238 |
1176 |
10.51 |
5.48 |
0.73 |
0.85 |
| 7 |
A |
1058 |
1004 |
14.96 |
1.24 |
0.40 |
0.58 |
| 8 |
A |
962 |
913 |
6.68 |
4.02 |
0.66 |
0.79 |
| 9 |
A |
849 |
806 |
68.02 |
5.69 |
0.73 |
0.84 |
| 10 |
A |
669 |
636 |
27.54 |
8.95 |
0.07 |
0.13 |
| 11 |
A |
635 |
603 |
28.98 |
5.13 |
0.49 |
0.65 |
| 12 |
A |
349 |
332 |
1.27 |
2.22 |
0.19 |
0.33 |
| 13 |
A |
280 |
266 |
3.28 |
4.21 |
0.63 |
0.77 |
| 14 |
A |
222 |
211 |
2.71 |
1.11 |
0.68 |
0.81 |
| 15 |
A |
89 |
85 |
9.04 |
1.64 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 8515.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 8084.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.102 |
-0.017 |
0.526 |
| C2 |
-0.691 |
-1.273 |
0.207 |
| H3 |
0.234 |
0.071 |
1.596 |
| Cl4 |
1.717 |
-0.247 |
-0.172 |
| Cl5 |
-0.682 |
1.431 |
-0.058 |
| O6 |
-1.767 |
-1.284 |
-0.325 |
| H7 |
-0.169 |
-2.187 |
0.516 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5192 | 1.0820 | 1.7749 | 1.7465 | 2.4131 | 2.1871 |
C2 | 1.5192 | | 2.1431 | 2.6445 | 2.7171 | 1.2008 | 1.0969 | H3 | 1.0820 | 2.1431 | | 2.3293 | 2.3289 | 3.0877 | 2.5355 | Cl4 | 1.7749 | 2.6445 | 2.3293 | | 2.9301 | 3.6388 | 2.7914 | Cl5 | 1.7465 | 2.7171 | 2.3289 | 2.9301 | | 2.9365 | 3.6990 | O6 | 2.4131 | 1.2008 | 3.0877 | 3.6388 | 2.9365 | | 2.0192 | H7 | 2.1871 | 1.0969 | 2.5355 | 2.7914 | 3.6990 | 2.0192 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.625 |
|
C1 |
C2 |
H7 |
112.439 |
| C2 |
C1 |
H3 |
109.812 |
|
C2 |
C1 |
Cl4 |
106.538 |
| C2 |
C1 |
Cl5 |
112.428 |
|
H3 |
C1 |
Cl4 |
106.711 |
| H3 |
C1 |
Cl5 |
108.557 |
|
Cl4 |
C1 |
Cl5 |
112.623 |
| O6 |
C2 |
H7 |
122.936 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
