Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1073.164807 |
| Energy at 298.15K | -1073.167299 |
| HF Energy | -1073.164807 |
| Nuclear repulsion energy | 265.128631 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3186 |
3049 |
1.26 |
|
|
|
| 2 |
A' |
2977 |
2849 |
40.37 |
|
|
|
| 3 |
A' |
1859 |
1780 |
182.80 |
|
|
|
| 4 |
A' |
1398 |
1338 |
2.33 |
|
|
|
| 5 |
A' |
1214 |
1162 |
6.83 |
|
|
|
| 6 |
A' |
1078 |
1031 |
28.28 |
|
|
|
| 7 |
A' |
793 |
759 |
22.25 |
|
|
|
| 8 |
A' |
444 |
425 |
4.50 |
|
|
|
| 9 |
A' |
315 |
302 |
23.03 |
|
|
|
| 10 |
A' |
250 |
240 |
2.62 |
|
|
|
| 11 |
A" |
1236 |
1183 |
19.64 |
|
|
|
| 12 |
A" |
1018 |
974 |
18.24 |
|
|
|
| 13 |
A" |
741 |
710 |
116.55 |
|
|
|
| 14 |
A" |
271 |
260 |
4.03 |
|
|
|
| 15 |
A" |
86 |
83 |
7.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8433.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 8071.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
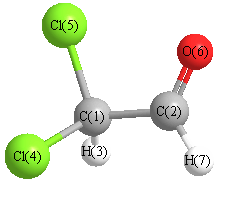
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.394 |
-0.022 |
0.000 |
| C2 |
-0.170 |
1.383 |
0.000 |
| H3 |
1.475 |
-0.012 |
0.000 |
| Cl4 |
-0.170 |
-0.835 |
1.466 |
| Cl5 |
-0.170 |
-0.835 |
-1.466 |
| O6 |
0.529 |
2.349 |
0.000 |
| H7 |
-1.271 |
1.427 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5142 | 1.0815 | 1.7686 | 1.7686 | 2.3750 | 2.2074 |
C2 | 1.5142 | | 2.1574 | 2.6588 | 2.6588 | 1.1923 | 1.1022 | H3 | 1.0815 | 2.1574 | | 2.3522 | 2.3522 | 2.5439 | 3.1008 | Cl4 | 1.7686 | 2.6588 | 2.3522 | | 2.9322 | 3.5743 | 2.9118 | Cl5 | 1.7686 | 2.6588 | 2.3522 | 2.9322 | | 3.5743 | 2.9118 | O6 | 2.3750 | 1.1923 | 2.5439 | 3.5743 | 3.5743 | | 2.0228 | H7 | 2.2074 | 1.1022 | 3.1008 | 2.9118 | 2.9118 | 2.0228 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.245 |
|
C1 |
C2 |
H7 |
114.135 |
| C2 |
C1 |
H3 |
111.341 |
|
C2 |
C1 |
Cl4 |
107.928 |
| C2 |
C1 |
Cl5 |
107.928 |
|
H3 |
C1 |
Cl4 |
108.842 |
| H3 |
C1 |
Cl5 |
108.842 |
|
Cl4 |
C1 |
Cl5 |
111.982 |
| O6 |
C2 |
H7 |
123.620 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.042 |
|
|
|
| 2 |
C |
0.138 |
|
|
|
| 3 |
H |
0.174 |
|
|
|
| 4 |
Cl |
-0.076 |
|
|
|
| 5 |
Cl |
-0.076 |
|
|
|
| 6 |
O |
-0.221 |
|
|
|
| 7 |
H |
0.104 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.323 |
-0.856 |
0.000 |
0.915 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.417 |
-2.832 |
0.000 |
| y |
-2.832 |
-48.039 |
0.000 |
| z |
0.000 |
0.000 |
-42.284 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.745 |
-2.832 |
0.000 |
| y |
-2.832 |
-6.689 |
0.000 |
| z |
0.000 |
0.000 |
1.944 |
|
| Polar |
|---|
| 3z2-r2 | 3.887 |
|---|
| x2-y2 | 7.623 |
|---|
| xy | -2.832 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.922 |
0.615 |
0.000 |
| y |
0.615 |
7.847 |
0.000 |
| z |
0.000 |
0.000 |
8.055 |
<r2> (average value of r
2) Å
2
| <r2> |
189.781 |
| (<r2>)1/2 |
13.776 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1073.161933 |
| Energy at 298.15K | |
| HF Energy | -1073.161933 |
| Nuclear repulsion energy | 267.904675 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3128 |
2994 |
5.24 |
81.17 |
0.17 |
0.29 |
| 2 |
A |
2962 |
2835 |
53.48 |
140.13 |
0.29 |
0.45 |
| 3 |
A |
1873 |
1793 |
150.55 |
16.07 |
0.47 |
0.64 |
| 4 |
A |
1395 |
1335 |
13.08 |
3.65 |
0.39 |
0.56 |
| 5 |
A |
1264 |
1210 |
11.52 |
2.54 |
0.73 |
0.85 |
| 6 |
A |
1212 |
1160 |
9.44 |
6.01 |
0.73 |
0.84 |
| 7 |
A |
1039 |
994 |
14.51 |
1.36 |
0.51 |
0.68 |
| 8 |
A |
942 |
902 |
9.47 |
4.02 |
0.63 |
0.77 |
| 9 |
A |
820 |
785 |
76.74 |
5.74 |
0.71 |
0.83 |
| 10 |
A |
657 |
629 |
36.07 |
9.58 |
0.08 |
0.15 |
| 11 |
A |
621 |
594 |
33.27 |
6.13 |
0.42 |
0.59 |
| 12 |
A |
340 |
325 |
1.62 |
2.59 |
0.19 |
0.32 |
| 13 |
A |
270 |
259 |
3.78 |
4.58 |
0.63 |
0.77 |
| 14 |
A |
213 |
204 |
2.77 |
1.28 |
0.69 |
0.82 |
| 15 |
A |
90 |
86 |
9.65 |
1.59 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 8413.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 8052.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.097 |
-0.016 |
0.521 |
| C2 |
-0.683 |
-1.282 |
0.200 |
| H3 |
0.229 |
0.074 |
1.595 |
| Cl4 |
1.724 |
-0.235 |
-0.169 |
| Cl5 |
-0.697 |
1.430 |
-0.059 |
| O6 |
-1.752 |
-1.301 |
-0.319 |
| H7 |
-0.145 |
-2.197 |
0.502 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5217 | 1.0858 | 1.7799 | 1.7480 | 2.4035 | 2.1946 |
C2 | 1.5217 | | 2.1486 | 2.6506 | 2.7242 | 1.1888 | 1.1034 | H3 | 1.0858 | 2.1486 | | 2.3319 | 2.3299 | 3.0789 | 2.5476 | Cl4 | 1.7799 | 2.6506 | 2.3319 | | 2.9395 | 3.6387 | 2.7915 | Cl5 | 1.7480 | 2.7242 | 2.3299 | 2.9395 | | 2.9386 | 3.7109 | O6 | 2.4035 | 1.1888 | 3.0789 | 3.6387 | 2.9386 | | 2.0154 | H7 | 2.1946 | 1.1034 | 2.5476 | 2.7915 | 3.7109 | 2.0154 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.472 |
|
C1 |
C2 |
H7 |
112.463 |
| C2 |
C1 |
H3 |
109.850 |
|
C2 |
C1 |
Cl4 |
106.541 |
| C2 |
C1 |
Cl5 |
112.669 |
|
H3 |
C1 |
Cl4 |
106.393 |
| H3 |
C1 |
Cl5 |
108.331 |
|
Cl4 |
C1 |
Cl5 |
112.859 |
| O6 |
C2 |
H7 |
123.065 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.053 |
|
|
|
| 2 |
C |
0.122 |
|
|
|
| 3 |
H |
0.173 |
|
|
|
| 4 |
Cl |
-0.087 |
|
|
|
| 5 |
Cl |
-0.056 |
|
|
|
| 6 |
O |
-0.201 |
|
|
|
| 7 |
H |
0.102 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.283 |
-0.673 |
1.927 |
2.411 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.927 |
-2.183 |
-1.209 |
| y |
-2.183 |
-41.387 |
-1.573 |
| z |
-1.209 |
-1.573 |
-39.886 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.291 |
-2.183 |
-1.209 |
| y |
-2.183 |
2.019 |
-1.573 |
| z |
-1.209 |
-1.573 |
4.271 |
|
| Polar |
|---|
| 3z2-r2 | 8.543 |
|---|
| x2-y2 | -5.540 |
|---|
| xy | -2.183 |
|---|
| xz | -1.209 |
|---|
| yz | -1.573 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.610 |
-0.637 |
0.175 |
| y |
-0.637 |
7.602 |
-0.298 |
| z |
0.175 |
-0.298 |
5.297 |
<r2> (average value of r
2) Å
2
| <r2> |
182.873 |
| (<r2>)1/2 |
13.523 |
