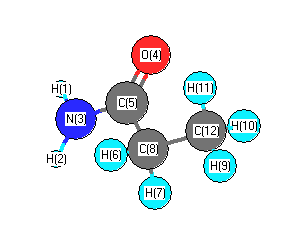
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3773 |
3611 |
38.46 |
|
|
|
| 2 |
A |
3633 |
3477 |
36.38 |
|
|
|
| 3 |
A |
3157 |
3021 |
19.59 |
|
|
|
| 4 |
A |
3148 |
3013 |
20.96 |
|
|
|
| 5 |
A |
3086 |
2954 |
9.88 |
|
|
|
| 6 |
A |
3074 |
2943 |
22.12 |
|
|
|
| 7 |
A |
3053 |
2922 |
22.72 |
|
|
|
| 8 |
A |
1812 |
1734 |
302.15 |
|
|
|
| 9 |
A |
1618 |
1548 |
121.31 |
|
|
|
| 10 |
A |
1501 |
1437 |
11.03 |
|
|
|
| 11 |
A |
1492 |
1428 |
7.51 |
|
|
|
| 12 |
A |
1465 |
1403 |
8.14 |
|
|
|
| 13 |
A |
1418 |
1357 |
19.15 |
|
|
|
| 14 |
A |
1412 |
1351 |
68.26 |
|
|
|
| 15 |
A |
1288 |
1233 |
79.80 |
|
|
|
| 16 |
A |
1286 |
1231 |
7.14 |
|
|
|
| 17 |
A |
1129 |
1080 |
1.10 |
|
|
|
| 18 |
A |
1100 |
1053 |
0.04 |
|
|
|
| 19 |
A |
1073 |
1027 |
1.54 |
|
|
|
| 20 |
A |
1006 |
963 |
4.16 |
|
|
|
| 21 |
A |
816 |
781 |
7.36 |
|
|
|
| 22 |
A |
799 |
765 |
3.83 |
|
|
|
| 23 |
A |
635 |
607 |
3.75 |
|
|
|
| 24 |
A |
603 |
577 |
7.58 |
|
|
|
| 25 |
A |
504 |
483 |
3.05 |
|
|
|
| 26 |
A |
438 |
420 |
3.76 |
|
|
|
| 27 |
A |
255 |
244 |
9.08 |
|
|
|
| 28 |
A |
212 |
203 |
0.42 |
|
|
|
| 29 |
A |
126 |
121 |
185.41 |
|
|
|
| 30 |
A |
23 |
22 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22467.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 21503.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.175 |
|
|
|
| 2 |
H |
0.175 |
|
|
|
| 3 |
N |
-0.267 |
|
|
|
| 4 |
O |
-0.331 |
|
|
|
| 5 |
C |
0.243 |
|
|
|
| 6 |
H |
0.129 |
|
|
|
| 7 |
H |
0.125 |
|
|
|
| 8 |
C |
-0.257 |
|
|
|
| 9 |
H |
0.110 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.130 |
|
|
|
| 12 |
C |
-0.360 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.997 |
-3.545 |
0.033 |
3.683 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.342 |
2.420 |
-0.011 |
| y |
2.420 |
-32.829 |
-0.073 |
| z |
-0.011 |
-0.073 |
-31.176 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.660 |
2.420 |
-0.011 |
| y |
2.420 |
-4.571 |
-0.073 |
| z |
-0.011 |
-0.073 |
-2.090 |
|
| Polar |
|---|
| 3z2-r2 | -4.179 |
|---|
| x2-y2 | 7.487 |
|---|
| xy | 2.420 |
|---|
| xz | -0.011 |
|---|
| yz | -0.073 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.027 |
0.347 |
0.020 |
| y |
0.347 |
7.249 |
-0.001 |
| z |
0.020 |
-0.001 |
5.238 |
<r2> (average value of r
2) Å
2
| <r2> |
126.533 |
| (<r2>)1/2 |
11.249 |