return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HOCH2COOH (Hydroxyacetic acid)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-303.888864
Energy at 298.15K-303.894615
HF Energy-302.784018
Nuclear repulsion energy180.911436
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3787 3596 83.41      
2 A' 3777 3586 92.24      
3 A' 3094 2938 25.89      
4 A' 1832 1740 240.57      
5 A' 1525 1448 8.57      
6 A' 1500 1424 5.50      
7 A' 1375 1305 146.69      
8 A' 1315 1248 17.56      
9 A' 1189 1129 93.53      
10 A' 1141 1084 261.02      
11 A' 886 841 35.35      
12 A' 660 626 17.75      
13 A' 481 457 26.38      
14 A' 286 271 9.24      
15 A" 3123 2965 9.97      
16 A" 1281 1216 0.11      
17 A" 1058 1005 1.01      
18 A" 664 630 112.21      
19 A" 523 497 11.02      
20 A" 363 345 75.87      
21 A" 103 98 8.22      

Unscaled Zero Point Vibrational Energy (zpe) 14982.0 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 14224.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.35963 0.13753 0.10134

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.600 -0.857 0.000
C2 0.000 0.517 0.000
O3 -0.928 1.481 0.000
O4 1.190 0.722 0.000
O5 0.399 -1.834 0.000
H6 -1.237 -0.943 0.878
H7 -1.237 -0.943 -0.878
H8 1.233 -1.346 0.000
H9 -0.448 2.321 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 O4 O5 H6 H7 H8 H9
C11.49832.36062.38621.39751.08791.08791.89673.1810
C21.49831.33881.20762.38422.10482.10482.23361.8592
O32.36061.33882.25053.57112.59662.59663.55870.9673
O42.38621.20762.25052.67533.07103.07102.06832.2894
O51.39752.38423.57112.67532.05902.05900.96594.2404
H61.08792.10482.59663.07102.05901.75532.65153.4707
H71.08792.10482.59663.07102.05901.75532.65153.4707
H81.89672.23363.55872.06830.96592.65152.65154.0339
H93.18101.85920.96732.28944.24043.47073.47074.0339

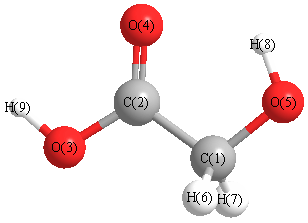
picture of Hydroxyacetic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 112.502 C1 C2 O4 123.378
C1 O5 H8 105.279 C2 C1 O5 110.789
C2 C1 H6 107.899 C2 C1 H7 107.899
C2 O3 H9 106.334 O3 C2 O4 124.120
O5 C1 H6 111.269 O5 C1 H7 111.269
H6 C1 H7 107.559
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability