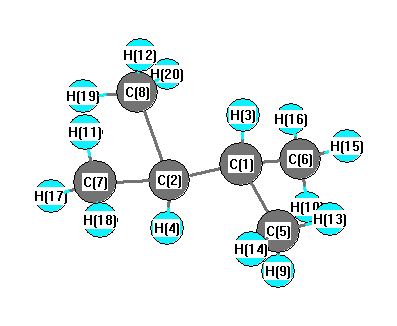
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3142 |
3007 |
0.00 |
|
|
|
| 2 |
Ag |
3116 |
2982 |
0.00 |
|
|
|
| 3 |
Ag |
3049 |
2918 |
0.00 |
|
|
|
| 4 |
Ag |
3003 |
2874 |
0.00 |
|
|
|
| 5 |
Ag |
1520 |
1455 |
0.00 |
|
|
|
| 6 |
Ag |
1502 |
1437 |
0.00 |
|
|
|
| 7 |
Ag |
1414 |
1354 |
0.00 |
|
|
|
| 8 |
Ag |
1373 |
1314 |
0.00 |
|
|
|
| 9 |
Ag |
1215 |
1163 |
0.00 |
|
|
|
| 10 |
Ag |
1193 |
1142 |
0.00 |
|
|
|
| 11 |
Ag |
957 |
916 |
0.00 |
|
|
|
| 12 |
Ag |
764 |
731 |
0.00 |
|
|
|
| 13 |
Ag |
487 |
466 |
0.00 |
|
|
|
| 14 |
Ag |
368 |
352 |
0.00 |
|
|
|
| 15 |
Ag |
227 |
218 |
0.00 |
|
|
|
| 16 |
Au |
3142 |
3007 |
55.01 |
|
|
|
| 17 |
Au |
3111 |
2978 |
2.20 |
|
|
|
| 18 |
Au |
3045 |
2914 |
45.14 |
|
|
|
| 19 |
Au |
1499 |
1435 |
3.85 |
|
|
|
| 20 |
Au |
1481 |
1417 |
1.65 |
|
|
|
| 21 |
Au |
1395 |
1335 |
12.85 |
|
|
|
| 22 |
Au |
1335 |
1277 |
2.45 |
|
|
|
| 23 |
Au |
1098 |
1051 |
1.73 |
|
|
|
| 24 |
Au |
971 |
930 |
0.65 |
|
|
|
| 25 |
Au |
921 |
881 |
2.10 |
|
|
|
| 26 |
Au |
287 |
275 |
0.00 |
|
|
|
| 27 |
Au |
187 |
179 |
0.00 |
|
|
|
| 28 |
Au |
56 |
54 |
0.00 |
|
|
|
| 29 |
Bg |
3125 |
2991 |
0.00 |
|
|
|
| 30 |
Bg |
3111 |
2977 |
0.00 |
|
|
|
| 31 |
Bg |
3043 |
2913 |
0.00 |
|
|
|
| 32 |
Bg |
1488 |
1424 |
0.00 |
|
|
|
| 33 |
Bg |
1479 |
1415 |
0.00 |
|
|
|
| 34 |
Bg |
1392 |
1333 |
0.00 |
|
|
|
| 35 |
Bg |
1350 |
1292 |
0.00 |
|
|
|
| 36 |
Bg |
1183 |
1132 |
0.00 |
|
|
|
| 37 |
Bg |
962 |
921 |
0.00 |
|
|
|
| 38 |
Bg |
911 |
872 |
0.00 |
|
|
|
| 39 |
Bg |
420 |
402 |
0.00 |
|
|
|
| 40 |
Bg |
179 |
172 |
0.00 |
|
|
|
| 41 |
Bu |
3128 |
2994 |
78.75 |
|
|
|
| 42 |
Bu |
3116 |
2982 |
131.69 |
|
|
|
| 43 |
Bu |
3046 |
2916 |
69.48 |
|
|
|
| 44 |
Bu |
3019 |
2890 |
31.38 |
|
|
|
| 45 |
Bu |
1505 |
1440 |
15.59 |
|
|
|
| 46 |
Bu |
1497 |
1432 |
22.71 |
|
|
|
| 47 |
Bu |
1409 |
1348 |
16.67 |
|
|
|
| 48 |
Bu |
1298 |
1243 |
5.02 |
|
|
|
| 49 |
Bu |
1177 |
1127 |
7.35 |
|
|
|
| 50 |
Bu |
991 |
949 |
5.80 |
|
|
|
| 51 |
Bu |
883 |
845 |
2.30 |
|
|
|
| 52 |
Bu |
389 |
373 |
0.10 |
|
|
|
| 53 |
Bu |
334 |
319 |
0.03 |
|
|
|
| 54 |
Bu |
198 |
189 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41244.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 39475.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.060 |
|
|
|
| 2 |
C |
-0.060 |
|
|
|
| 3 |
H |
0.120 |
|
|
|
| 4 |
H |
0.120 |
|
|
|
| 5 |
C |
-0.385 |
|
|
|
| 6 |
C |
-0.385 |
|
|
|
| 7 |
C |
-0.385 |
|
|
|
| 8 |
C |
-0.385 |
|
|
|
| 9 |
H |
0.116 |
|
|
|
| 10 |
H |
0.116 |
|
|
|
| 11 |
H |
0.116 |
|
|
|
| 12 |
H |
0.116 |
|
|
|
| 13 |
H |
0.115 |
|
|
|
| 14 |
H |
0.123 |
|
|
|
| 15 |
H |
0.115 |
|
|
|
| 16 |
H |
0.123 |
|
|
|
| 17 |
H |
0.115 |
|
|
|
| 18 |
H |
0.123 |
|
|
|
| 19 |
H |
0.115 |
|
|
|
| 20 |
H |
0.123 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.479 |
-0.444 |
0.000 |
| y |
-0.444 |
-42.074 |
0.000 |
| z |
0.000 |
0.000 |
-42.461 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.789 |
-0.444 |
0.000 |
| y |
-0.444 |
-0.104 |
0.000 |
| z |
0.000 |
0.000 |
-0.685 |
|
| Polar |
|---|
| 3z2-r2 | -1.371 |
|---|
| x2-y2 | 0.595 |
|---|
| xy | -0.444 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.665 |
0.288 |
0.000 |
| y |
0.288 |
11.815 |
0.000 |
| z |
0.000 |
0.000 |
11.306 |
<r2> (average value of r
2) Å
2
| <r2> |
205.660 |
| (<r2>)1/2 |
14.341 |