Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -216.809213 |
| Energy at 298.15K | |
| HF Energy | -216.003269 |
| Nuclear repulsion energy | 117.169479 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3305 |
3138 |
2.33 |
43.62 |
0.71 |
0.83 |
| 2 |
A' |
3230 |
3066 |
4.08 |
162.33 |
0.13 |
0.22 |
| 3 |
A' |
3209 |
3047 |
5.40 |
27.06 |
0.74 |
0.85 |
| 4 |
A' |
3094 |
2937 |
31.10 |
158.39 |
0.09 |
0.17 |
| 5 |
A' |
1725 |
1638 |
3.45 |
10.00 |
0.07 |
0.12 |
| 6 |
A' |
1533 |
1455 |
3.16 |
11.27 |
0.56 |
0.72 |
| 7 |
A' |
1470 |
1396 |
7.92 |
6.73 |
0.57 |
0.73 |
| 8 |
A' |
1444 |
1371 |
9.95 |
3.12 |
0.74 |
0.85 |
| 9 |
A' |
1325 |
1258 |
0.17 |
13.43 |
0.28 |
0.44 |
| 10 |
A' |
1163 |
1104 |
54.95 |
1.77 |
0.75 |
0.86 |
| 11 |
A' |
1027 |
975 |
28.74 |
3.93 |
0.73 |
0.84 |
| 12 |
A' |
936 |
889 |
1.67 |
4.18 |
0.09 |
0.16 |
| 13 |
A' |
616 |
585 |
5.86 |
1.33 |
0.74 |
0.85 |
| 14 |
A' |
272 |
258 |
2.33 |
0.96 |
0.51 |
0.68 |
| 15 |
A" |
3131 |
2973 |
20.69 |
76.12 |
0.75 |
0.86 |
| 16 |
A" |
1299 |
1233 |
0.05 |
5.03 |
0.75 |
0.86 |
| 17 |
A" |
1070 |
1016 |
16.01 |
0.62 |
0.75 |
0.86 |
| 18 |
A" |
1042 |
989 |
9.32 |
0.03 |
0.75 |
0.86 |
| 19 |
A" |
974 |
925 |
39.54 |
1.00 |
0.75 |
0.86 |
| 20 |
A" |
577 |
547 |
9.30 |
4.28 |
0.75 |
0.86 |
| 21 |
A" |
182 |
173 |
2.84 |
2.41 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16311.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 15486.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.940 |
-0.200 |
0.000 |
| C2 |
0.000 |
0.949 |
0.000 |
| C3 |
1.322 |
0.833 |
0.000 |
| F4 |
-0.268 |
-1.404 |
0.000 |
| H5 |
1.952 |
1.704 |
0.000 |
| H6 |
1.787 |
-0.136 |
0.000 |
| H7 |
-0.464 |
1.923 |
0.000 |
| H8 |
-1.577 |
-0.172 |
0.881 |
| H9 |
-1.577 |
-0.172 |
-0.881 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4851 | 2.4866 | 1.3790 | 3.4627 | 2.7279 | 2.1760 | 1.0875 | 1.0875 |
C2 | 1.4851 | | 1.3268 | 2.3688 | 2.0928 | 2.0908 | 1.0784 | 2.1262 | 2.1262 | C3 | 2.4866 | 1.3268 | | 2.7443 | 1.0752 | 1.0749 | 2.0918 | 3.1921 | 3.1921 | F4 | 1.3790 | 2.3688 | 2.7443 | | 3.8195 | 2.4147 | 3.3329 | 2.0023 | 2.0023 | H5 | 3.4627 | 2.0928 | 1.0752 | 3.8195 | | 1.8474 | 2.4256 | 4.0927 | 4.0927 | H6 | 2.7279 | 2.0908 | 1.0749 | 2.4147 | 1.8474 | | 3.0504 | 3.4779 | 3.4779 | H7 | 2.1760 | 1.0784 | 2.0918 | 3.3329 | 2.4256 | 3.0504 | | 2.5307 | 2.5307 | H8 | 1.0875 | 2.1262 | 3.1921 | 2.0023 | 4.0927 | 3.4779 | 2.5307 | | 1.7618 | H9 | 1.0875 | 2.1262 | 3.1921 | 2.0023 | 4.0927 | 3.4779 | 2.5307 | 1.7618 | |
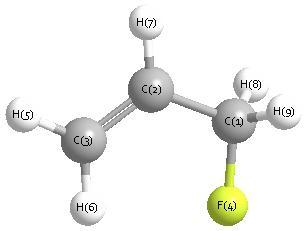 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.240 |
|
C1 |
C2 |
H7 |
115.258 |
| C2 |
C1 |
F4 |
111.544 |
|
C2 |
C1 |
H8 |
110.529 |
| C2 |
C1 |
H9 |
110.529 |
|
C2 |
C3 |
H5 |
120.851 |
| C2 |
C3 |
H6 |
120.688 |
|
C3 |
C2 |
H7 |
120.502 |
| F4 |
C1 |
H8 |
107.963 |
|
F4 |
C1 |
H9 |
107.963 |
| H5 |
C3 |
H6 |
118.461 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -216.808574 |
| Energy at 298.15K | |
| HF Energy | -216.002791 |
| Nuclear repulsion energy | 114.845365 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3286 |
3120 |
7.25 |
53.06 |
0.54 |
0.70 |
| 2 |
A |
3227 |
3064 |
2.56 |
123.27 |
0.23 |
0.37 |
| 3 |
A |
3202 |
3040 |
6.11 |
59.57 |
0.13 |
0.23 |
| 4 |
A |
3152 |
2992 |
19.57 |
59.80 |
0.75 |
0.85 |
| 5 |
A |
3104 |
2947 |
30.46 |
119.19 |
0.07 |
0.13 |
| 6 |
A |
1720 |
1633 |
0.37 |
10.13 |
0.08 |
0.15 |
| 7 |
A |
1537 |
1459 |
2.44 |
4.58 |
0.70 |
0.82 |
| 8 |
A |
1484 |
1409 |
19.93 |
7.32 |
0.66 |
0.80 |
| 9 |
A |
1417 |
1345 |
10.84 |
2.73 |
0.35 |
0.52 |
| 10 |
A |
1324 |
1257 |
0.10 |
10.53 |
0.37 |
0.54 |
| 11 |
A |
1298 |
1233 |
4.25 |
7.54 |
0.71 |
0.83 |
| 12 |
A |
1202 |
1142 |
3.70 |
1.22 |
0.64 |
0.78 |
| 13 |
A |
1085 |
1031 |
107.82 |
4.76 |
0.57 |
0.72 |
| 14 |
A |
1048 |
995 |
30.79 |
0.92 |
0.72 |
0.84 |
| 15 |
A |
1009 |
958 |
4.34 |
1.66 |
0.10 |
0.18 |
| 16 |
A |
984 |
934 |
40.39 |
0.86 |
0.66 |
0.79 |
| 17 |
A |
943 |
895 |
1.99 |
2.18 |
0.16 |
0.27 |
| 18 |
A |
675 |
641 |
6.56 |
2.61 |
0.52 |
0.69 |
| 19 |
A |
439 |
417 |
2.07 |
3.13 |
0.45 |
0.62 |
| 20 |
A |
338 |
321 |
6.65 |
2.25 |
0.75 |
0.86 |
| 21 |
A |
116 |
110 |
1.02 |
3.97 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16296.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 15472.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.603 |
0.419 |
0.331 |
| C2 |
0.640 |
-0.384 |
0.250 |
| C3 |
1.785 |
0.099 |
-0.221 |
| F4 |
-1.629 |
-0.216 |
-0.355 |
| H5 |
2.681 |
-0.497 |
-0.242 |
| H6 |
1.847 |
1.108 |
-0.592 |
| H7 |
0.583 |
-1.397 |
0.613 |
| H8 |
-0.931 |
0.532 |
1.361 |
| H9 |
-0.456 |
1.401 |
-0.107 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4826 | 2.4719 | 1.3880 | 3.4570 | 2.7074 | 2.1873 | 1.0865 | 1.0855 |
C2 | 1.4826 | | 1.3284 | 2.3544 | 2.1018 | 2.0957 | 1.0774 | 2.1314 | 2.1250 | C3 | 2.4719 | 1.3284 | | 3.4305 | 1.0758 | 1.0776 | 2.0920 | 3.1726 | 2.5941 | F4 | 1.3880 | 2.3544 | 3.4305 | | 4.3199 | 3.7266 | 2.6875 | 1.9984 | 2.0133 | H5 | 3.4570 | 2.1018 | 1.0758 | 4.3199 | | 1.8424 | 2.4375 | 4.0830 | 3.6683 | H6 | 2.7074 | 2.0957 | 1.0776 | 3.7266 | 1.8424 | | 3.0538 | 3.4441 | 2.3711 | H7 | 2.1873 | 1.0774 | 2.0920 | 2.6875 | 2.4375 | 3.0538 | | 2.5640 | 3.0702 | H8 | 1.0865 | 2.1314 | 3.1726 | 1.9984 | 4.0830 | 3.4441 | 2.5640 | | 1.7706 | H9 | 1.0855 | 2.1250 | 2.5941 | 2.0133 | 3.6683 | 2.3711 | 3.0702 | 1.7706 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.042 |
|
C1 |
C2 |
H7 |
116.495 |
| C2 |
C1 |
F4 |
110.163 |
|
C2 |
C1 |
H8 |
111.186 |
| C2 |
C1 |
H9 |
110.729 |
|
C2 |
C3 |
H5 |
121.549 |
| C2 |
C3 |
H6 |
120.806 |
|
C3 |
C2 |
H7 |
120.455 |
| F4 |
C1 |
H8 |
107.092 |
|
F4 |
C1 |
H9 |
108.342 |
| H5 |
C3 |
H6 |
117.644 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
