return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6 (Hexa-1,5-diene-3-yne)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-231.710562
Energy at 298.15K-231.714493
HF Energy-230.677487
Nuclear repulsion energy175.776217
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3296 3129 0.00      
2 Ag 3214 3051 0.00      
3 Ag 3201 3039 0.00      
4 Ag 2250 2136 0.00      
5 Ag 1655 1571 0.00      
6 Ag 1454 1380 0.00      
7 Ag 1330 1263 0.00      
8 Ag 1093 1038 0.00      
9 Ag 725 689 0.00      
10 Ag 519 492 0.00      
11 Ag 215 204 0.00      
12 Au 1033 981 40.27      
13 Au 965 916 73.88      
14 Au 709 673 10.24      
15 Au 185 176 6.70      
16 Au 44 42 0.09      
17 Bg 1030 978 0.00      
18 Bg 964 915 0.00      
19 Bg 743 705 0.00      
20 Bg 406 386 0.00      
21 Bu 3296 3129 8.50      
22 Bu 3211 3049 9.70      
23 Bu 3201 3039 9.86      
24 Bu 1684 1598 33.88      
25 Bu 1476 1402 5.83      
26 Bu 1327 1260 3.50      
27 Bu 1230 1168 13.15      
28 Bu 1028 976 0.22      
29 Bu 527 500 9.01      
30 Bu 108 102 2.23      

Unscaled Zero Point Vibrational Energy (zpe) 21058.8 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19993.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.87512 0.04588 0.04360

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.610 0.000
C2 0.000 -0.610 0.000
C3 0.000 2.022 0.000
C4 0.000 -2.022 0.000
C5 1.110 2.769 0.000
C6 -1.110 -2.769 0.000
H7 -0.962 2.511 0.000
H8 0.962 -2.511 0.000
H9 2.086 2.314 0.000
H10 1.046 3.843 0.000
H11 -2.086 -2.314 0.000
H12 -1.046 -3.843 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
C11.21921.41232.63152.42753.55582.13063.26532.69373.39823.59154.5736
C21.21922.63151.41233.55582.42753.26532.13063.59154.57362.69373.3982
C31.41232.63154.04381.33764.91741.07874.63372.10612.09994.81165.9572
C42.63151.41234.04384.91741.33764.63371.07874.81165.95722.10612.0999
C52.42753.55581.33764.91745.96552.08735.28161.07661.07606.00386.9540
C63.55582.42754.91741.33765.96555.28162.08736.00386.95401.07661.0760
H72.13063.26531.07874.63372.08735.28165.37743.05362.40934.95436.3542
H83.26532.13064.63371.07875.28162.08735.37744.95436.35423.05362.4093
H92.69373.59152.10614.81161.07666.00383.05364.95431.84866.23086.9077
H103.39824.57362.09995.95721.07606.95402.40936.35421.84866.90777.9652
H113.59152.69374.81162.10616.00381.07664.95433.05366.23086.90771.8486
H124.57363.39825.95722.09996.95401.07606.35422.40936.90777.96521.8486

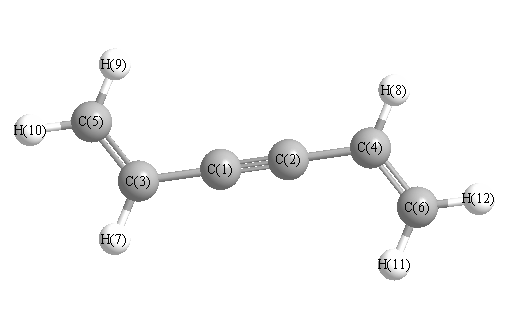
picture of Hexa-1,5-diene-3-yne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 180.000 C1 C3 C5 123.937
C1 C3 H7 116.954 C2 C1 C3 180.000
C2 C4 C6 123.937 C2 C4 H8 116.954
C3 C5 H9 121.095 C3 C5 H10 120.547
C4 C6 H11 121.095 C4 C6 H12 120.547
C5 C3 H7 119.109 C6 C4 H8 119.109
H9 C5 H10 118.358 H11 C6 H12 118.358
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability