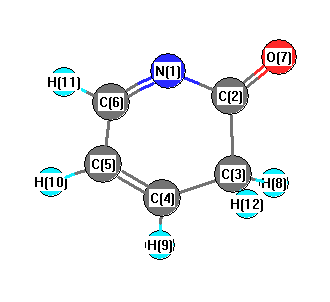
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3226 |
3087 |
10.23 |
|
|
|
| 2 |
A |
3193 |
3056 |
4.19 |
|
|
|
| 3 |
A |
3122 |
2988 |
18.42 |
|
|
|
| 4 |
A |
3121 |
2987 |
15.96 |
|
|
|
| 5 |
A |
3018 |
2889 |
0.54 |
|
|
|
| 6 |
A |
1831 |
1753 |
266.25 |
|
|
|
| 7 |
A |
1730 |
1656 |
69.25 |
|
|
|
| 8 |
A |
1634 |
1564 |
131.01 |
|
|
|
| 9 |
A |
1437 |
1375 |
28.36 |
|
|
|
| 10 |
A |
1412 |
1351 |
31.91 |
|
|
|
| 11 |
A |
1409 |
1349 |
12.27 |
|
|
|
| 12 |
A |
1331 |
1274 |
4.60 |
|
|
|
| 13 |
A |
1227 |
1175 |
40.03 |
|
|
|
| 14 |
A |
1173 |
1122 |
5.98 |
|
|
|
| 15 |
A |
1151 |
1102 |
27.30 |
|
|
|
| 16 |
A |
1024 |
980 |
1.60 |
|
|
|
| 17 |
A |
1018 |
974 |
4.20 |
|
|
|
| 18 |
A |
1005 |
962 |
3.12 |
|
|
|
| 19 |
A |
981 |
939 |
30.04 |
|
|
|
| 20 |
A |
965 |
923 |
5.73 |
|
|
|
| 21 |
A |
928 |
888 |
4.62 |
|
|
|
| 22 |
A |
789 |
755 |
5.54 |
|
|
|
| 23 |
A |
718 |
687 |
52.69 |
|
|
|
| 24 |
A |
620 |
593 |
1.14 |
|
|
|
| 25 |
A |
577 |
552 |
0.44 |
|
|
|
| 26 |
A |
528 |
505 |
1.70 |
|
|
|
| 27 |
A |
468 |
448 |
2.50 |
|
|
|
| 28 |
A |
428 |
410 |
4.58 |
|
|
|
| 29 |
A |
274 |
262 |
2.34 |
|
|
|
| 30 |
A |
80 |
76 |
2.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20206.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 19339.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.165 |
|
|
|
| 2 |
C |
0.228 |
|
|
|
| 3 |
C |
-0.238 |
|
|
|
| 4 |
C |
-0.110 |
|
|
|
| 5 |
C |
-0.176 |
|
|
|
| 6 |
C |
-0.024 |
|
|
|
| 7 |
O |
-0.260 |
|
|
|
| 8 |
H |
0.142 |
|
|
|
| 9 |
H |
0.154 |
|
|
|
| 10 |
H |
0.153 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
| 12 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.545 |
2.133 |
0.525 |
5.048 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.330 |
3.096 |
1.807 |
| y |
3.096 |
-37.576 |
0.794 |
| z |
1.807 |
0.794 |
-40.050 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.517 |
3.096 |
1.807 |
| y |
3.096 |
4.614 |
0.794 |
| z |
1.807 |
0.794 |
0.903 |
|
| Polar |
|---|
| 3z2-r2 | 1.806 |
|---|
| x2-y2 | -6.754 |
|---|
| xy | 3.096 |
|---|
| xz | 1.807 |
|---|
| yz | 0.794 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.430 |
0.441 |
0.133 |
| y |
0.441 |
10.200 |
0.048 |
| z |
0.133 |
0.048 |
5.699 |
<r2> (average value of r
2) Å
2
| <r2> |
172.847 |
| (<r2>)1/2 |
13.147 |