return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (propenalol)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-266.756006
Energy at 298.15K-266.761227
HF Energy-265.732976
Nuclear repulsion energy166.234512
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3297 3130 0.60      
2 A' 3232 3069 0.88      
3 A' 3125 2967 213.26      
4 A' 3051 2897 111.87      
5 A' 1719 1632 204.33      
6 A' 1670 1586 237.08      
7 A' 1502 1426 48.40      
8 A' 1434 1361 13.56      
9 A' 1425 1353 66.56      
10 A' 1311 1245 151.43      
11 A' 1131 1074 12.66      
12 A' 1026 974 52.86      
13 A' 900 854 6.06      
14 A' 525 499 18.14      
15 A' 294 279 6.01      
16 A" 1071 1017 10.63      
17 A" 1044 992 22.56      
18 A" 969 920 52.54      
19 A" 813 772 35.86      
20 A" 410 389 5.18      
21 A" 290 275 3.70      

Unscaled Zero Point Vibrational Energy (zpe) 15119.0 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 14354.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.33079 0.17600 0.11488

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.099 0.000
C2 1.223 0.360 0.000
C3 -1.178 0.425 0.000
O4 1.274 -0.878 0.000
O5 -1.271 -0.887 0.000
H6 -0.326 -1.208 0.000
H7 0.007 2.170 0.000
H8 2.159 0.926 0.000
H9 -2.131 0.929 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.42901.35702.35132.35792.32981.07142.16562.1375
C21.42902.40201.23912.78932.20442.18091.09293.4020
C31.35702.40202.77581.31501.84182.10933.37381.0783
O42.35131.23912.77582.54501.63293.30072.00913.8541
O52.35792.78931.31502.54500.99893.31353.87972.0092
H62.32982.20441.84181.63290.99893.39453.27492.7976
H71.07142.18092.10933.30073.31353.39452.48592.4714
H82.16561.09293.37382.00913.87973.27492.48594.2893
H92.13753.40201.07833.85412.00922.79762.47144.2893

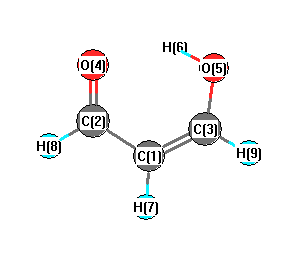
picture of propenalol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 123.436 C1 C2 H8 117.719
C1 C3 O5 123.871 C1 C3 H9 122.319
C2 C1 C3 119.097 C2 C1 H7 120.766
C3 C1 H7 120.136 C3 O5 H6 104.666
O4 C2 H8 118.844 O5 C3 H9 113.810
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/cc-pVTZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.365      
2 C 0.197      
3 C 0.135      
4 O -0.438      
5 O -0.341      
6 H 0.299      
7 H 0.183      
8 H 0.140      
9 H 0.191      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000