Jump to
S1C2
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -115.775612 |
| Energy at 298.15K | -115.777366 |
| HF Energy | -115.234279 |
| Nuclear repulsion energy | 58.015829 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3333 |
3209 |
|
|
|
|
| 2 |
A' |
3119 |
3003 |
|
|
|
|
| 3 |
A' |
1784 |
1718 |
|
|
|
|
| 4 |
A' |
1264 |
1217 |
|
|
|
|
| 5 |
A' |
1001 |
964 |
|
|
|
|
| 6 |
A' |
934 |
899 |
|
|
|
|
| 7 |
A' |
608 |
585 |
|
|
|
|
| 8 |
A" |
3496 |
3367 |
|
|
|
|
| 9 |
A" |
2381 |
2293 |
|
|
|
|
| 10 |
A" |
1080 |
1040 |
|
|
|
|
| 11 |
A" |
991 |
954 |
|
|
|
|
| 12 |
A" |
855 |
823 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10422.9 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 10036.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.037 |
0.887 |
0.000 |
| H2 |
0.732 |
1.649 |
0.000 |
| C3 |
-0.037 |
-0.418 |
0.659 |
| C4 |
-0.037 |
-0.418 |
-0.659 |
| H5 |
-0.036 |
-0.977 |
1.572 |
| H6 |
-0.036 |
-0.977 |
-1.572 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.0828 | 1.4617 | 1.4617 | 2.4384 | 2.4384 |
H2 | 1.0828 | | 2.3017 | 2.3017 | 3.1557 | 3.1557 | C3 | 1.4617 | 2.3017 | | 1.3174 | 1.0711 | 2.3000 | C4 | 1.4617 | 2.3017 | 1.3174 | | 2.3000 | 1.0711 | H5 | 2.4384 | 3.1557 | 1.0711 | 2.3000 | | 3.1448 | H6 | 2.4384 | 3.1557 | 2.3000 | 1.0711 | 3.1448 | |
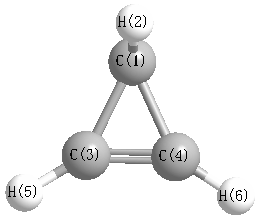 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.215 |
|
C1 |
C3 |
H5 |
148.237 |
| C1 |
C4 |
C3 |
63.215 |
|
C1 |
C4 |
H6 |
148.237 |
| H2 |
C1 |
C3 |
128.927 |
|
H2 |
C1 |
C4 |
128.927 |
| C3 |
C1 |
C4 |
53.571 |
|
C3 |
C4 |
H6 |
148.548 |
| C4 |
C3 |
H5 |
148.548 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
