All results from a given calculation for CH2OOH (CH2OOH radical)
using model chemistry: QCISD(T)/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C1 |
2A |
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.922646 |
| Energy at 298.15K | |
| HF Energy | -189.245482 |
| Nuclear repulsion energy | 74.530214 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.132 |
0.276 |
0.093 |
| O2 |
0.075 |
-0.576 |
-0.065 |
| O3 |
-1.139 |
0.223 |
-0.081 |
| H4 |
1.036 |
1.260 |
-0.346 |
| H5 |
2.065 |
-0.269 |
0.102 |
| H6 |
-1.378 |
0.180 |
0.856 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
H6 |
| C1 | | 1.3662 | 2.2782 | 1.0815 | 1.0813 | 2.6244 |
O2 | 1.3662 | | 1.4537 | 2.0907 | 2.0207 | 1.8789 | O3 | 2.2782 | 1.4537 | | 2.4242 | 3.2475 | 0.9672 | H4 | 1.0815 | 2.0907 | 2.4242 | | 1.8970 | 2.9041 | H5 | 1.0813 | 2.0207 | 3.2475 | 1.8970 | | 3.5531 | H6 | 2.6244 | 1.8789 | 0.9672 | 2.9041 | 3.5531 | |
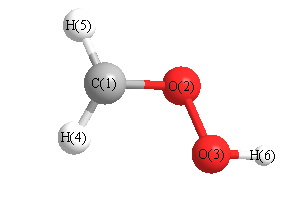 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
O3 |
107.742 |
|
O2 |
C1 |
H4 |
116.857 |
| O2 |
C1 |
H5 |
110.764 |
|
O2 |
O3 |
H6 |
99.857 |
| H4 |
C1 |
H5 |
122.588 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
