Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.621172 |
| Energy at 298.15K | -169.625200 |
| HF Energy | -168.972357 |
| Nuclear repulsion energy | 71.036019 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3890 |
3693 |
92.66 |
|
|
|
| 2 |
A' |
3578 |
3397 |
12.80 |
|
|
|
| 3 |
A' |
3122 |
2964 |
47.72 |
|
|
|
| 4 |
A' |
1763 |
1674 |
168.03 |
|
|
|
| 5 |
A' |
1435 |
1362 |
8.50 |
|
|
|
| 6 |
A' |
1344 |
1276 |
159.84 |
|
|
|
| 7 |
A' |
1202 |
1141 |
35.48 |
|
|
|
| 8 |
A' |
1072 |
1018 |
210.14 |
|
|
|
| 9 |
A' |
620 |
589 |
1.33 |
|
|
|
| 10 |
A" |
1058 |
1005 |
2.04 |
|
|
|
| 11 |
A" |
846 |
804 |
60.67 |
|
|
|
| 12 |
A" |
415 |
394 |
65.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10172.9 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 9658.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.374 |
0.000 |
| O2 |
-1.002 |
-0.529 |
0.000 |
| N3 |
1.196 |
-0.021 |
0.000 |
| H4 |
-0.330 |
1.409 |
0.000 |
| H5 |
-1.838 |
-0.058 |
0.000 |
| H6 |
1.810 |
0.784 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3493 | 1.2600 | 1.0860 | 1.8878 | 1.8554 |
O2 | 1.3493 | | 2.2561 | 2.0513 | 0.9596 | 3.1032 | N3 | 1.2600 | 2.2561 | | 2.0919 | 3.0341 | 1.0125 | H4 | 1.0860 | 2.0513 | 2.0919 | | 2.1031 | 2.2292 | H5 | 1.8878 | 0.9596 | 3.0341 | 2.1031 | | 3.7431 | H6 | 1.8554 | 3.1032 | 1.0125 | 2.2292 | 3.7431 | |
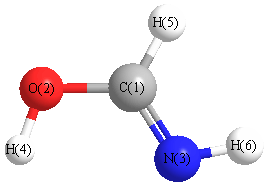 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.514 |
|
C1 |
N3 |
H6 |
108.973 |
| O2 |
C1 |
N3 |
119.643 |
|
O2 |
C1 |
H4 |
114.342 |
| N3 |
C1 |
H4 |
126.015 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.630259 |
| Energy at 298.15K | -169.634436 |
| HF Energy | -168.981856 |
| Nuclear repulsion energy | 71.385536 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3782 |
3590 |
65.82 |
|
|
|
| 2 |
A' |
3583 |
3402 |
12.66 |
|
|
|
| 3 |
A' |
3190 |
3029 |
24.31 |
|
|
|
| 4 |
A' |
1737 |
1649 |
230.79 |
|
|
|
| 5 |
A' |
1411 |
1340 |
18.32 |
|
|
|
| 6 |
A' |
1388 |
1318 |
4.22 |
|
|
|
| 7 |
A' |
1196 |
1136 |
103.82 |
|
|
|
| 8 |
A' |
1082 |
1027 |
174.41 |
|
|
|
| 9 |
A' |
591 |
561 |
51.44 |
|
|
|
| 10 |
A" |
1075 |
1021 |
0.92 |
|
|
|
| 11 |
A" |
848 |
805 |
29.31 |
|
|
|
| 12 |
A" |
639 |
606 |
168.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10261.1 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 9741.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.417 |
0.000 |
| O2 |
-1.110 |
-0.334 |
0.000 |
| N3 |
1.154 |
-0.097 |
0.000 |
| H4 |
-0.253 |
1.469 |
0.000 |
| H5 |
-0.803 |
-1.251 |
0.000 |
| H6 |
1.859 |
0.630 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3407 | 1.2637 | 1.0814 | 1.8515 | 1.8709 |
O2 | 1.3407 | | 2.2772 | 1.9964 | 0.9673 | 3.1217 | N3 | 1.2637 | 2.2772 | | 2.1048 | 2.2724 | 1.0120 | H4 | 1.0814 | 1.9964 | 2.1048 | | 2.7749 | 2.2719 | H5 | 1.8515 | 0.9673 | 2.2724 | 2.7749 | | 3.2592 | H6 | 1.8709 | 3.1217 | 1.0120 | 2.2719 | 3.2592 | |
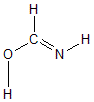 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
105.545 |
|
C1 |
N3 |
H6 |
110.107 |
| O2 |
C1 |
N3 |
121.920 |
|
O2 |
C1 |
H4 |
110.569 |
| N3 |
C1 |
H4 |
127.511 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.624841 |
| Energy at 298.15K | -169.628954 |
| HF Energy | -168.975892 |
| Nuclear repulsion energy | 70.981131 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3818 |
3625 |
46.72 |
|
|
|
| 2 |
A' |
3501 |
3323 |
6.84 |
|
|
|
| 3 |
A' |
3238 |
3074 |
8.72 |
|
|
|
| 4 |
A' |
1732 |
1644 |
228.79 |
|
|
|
| 5 |
A' |
1427 |
1355 |
1.31 |
|
|
|
| 6 |
A' |
1362 |
1293 |
31.91 |
|
|
|
| 7 |
A' |
1127 |
1070 |
269.15 |
|
|
|
| 8 |
A' |
1094 |
1038 |
31.93 |
|
|
|
| 9 |
A' |
597 |
567 |
33.83 |
|
|
|
| 10 |
A" |
1102 |
1046 |
63.20 |
|
|
|
| 11 |
A" |
863 |
820 |
51.40 |
|
|
|
| 12 |
A" |
526 |
499 |
69.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10193.5 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 9677.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.439 |
0.000 |
| O2 |
-1.099 |
-0.349 |
0.000 |
| N3 |
1.222 |
0.120 |
0.000 |
| H4 |
-0.294 |
1.477 |
0.000 |
| H5 |
-0.813 |
-1.270 |
0.000 |
| H6 |
1.348 |
-0.890 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3524 | 1.2626 | 1.0781 | 1.8923 | 1.8933 |
O2 | 1.3524 | | 2.3673 | 1.9950 | 0.9642 | 2.5059 | N3 | 1.2626 | 2.3673 | | 2.0343 | 2.4634 | 1.0180 | H4 | 1.0781 | 1.9950 | 2.0343 | | 2.7947 | 2.8807 | H5 | 1.8923 | 0.9642 | 2.4634 | 2.7947 | | 2.1935 | H6 | 1.8933 | 2.5059 | 1.0180 | 2.8807 | 2.1935 | |
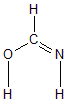 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.375 |
|
C1 |
N3 |
H6 |
111.787 |
| O2 |
C1 |
N3 |
129.687 |
|
O2 |
C1 |
H4 |
109.815 |
| N3 |
C1 |
H4 |
120.498 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
