Jump to
S1C2
S1C3
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.663775 |
| Energy at 298.15K | -169.667785 |
| HF Energy | -168.971812 |
| Nuclear repulsion energy | 70.821570 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3869 |
3725 |
|
|
|
|
| 2 |
A' |
3540 |
3409 |
|
|
|
|
| 3 |
A' |
3086 |
2971 |
|
|
|
|
| 4 |
A' |
1746 |
1681 |
|
|
|
|
| 5 |
A' |
1424 |
1372 |
|
|
|
|
| 6 |
A' |
1341 |
1292 |
|
|
|
|
| 7 |
A' |
1199 |
1155 |
|
|
|
|
| 8 |
A' |
1063 |
1024 |
|
|
|
|
| 9 |
A' |
613 |
590 |
|
|
|
|
| 10 |
A" |
1046 |
1007 |
|
|
|
|
| 11 |
A" |
838 |
807 |
|
|
|
|
| 12 |
A" |
408 |
392 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10086.1 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 9711.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.376 |
0.000 |
| O2 |
-1.005 |
-0.532 |
0.000 |
| N3 |
1.200 |
-0.020 |
0.000 |
| H4 |
-0.334 |
1.412 |
0.000 |
| H5 |
-1.840 |
-0.057 |
0.000 |
| H6 |
1.813 |
0.788 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3548 | 1.2638 | 1.0880 | 1.8899 | 1.8594 |
O2 | 1.3548 | | 2.2641 | 2.0566 | 0.9602 | 3.1123 | N3 | 1.2638 | 2.2641 | | 2.0981 | 3.0400 | 1.0144 | H4 | 1.0880 | 2.0566 | 2.0981 | | 2.1035 | 2.2353 | H5 | 1.8899 | 0.9602 | 3.0400 | 2.1035 | | 3.7492 | H6 | 1.8594 | 3.1123 | 1.0144 | 2.2353 | 3.7492 | |
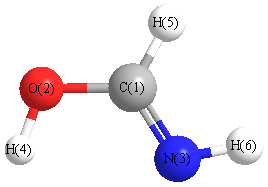 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.235 |
|
C1 |
N3 |
H6 |
108.912 |
| O2 |
C1 |
N3 |
119.640 |
|
O2 |
C1 |
H4 |
114.245 |
| N3 |
C1 |
H4 |
126.115 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.672655 |
| Energy at 298.15K | -169.676816 |
| HF Energy | -168.981362 |
| Nuclear repulsion energy | 71.176874 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3767 |
3627 |
|
|
|
|
| 2 |
A' |
3546 |
3415 |
|
|
|
|
| 3 |
A' |
3155 |
3038 |
|
|
|
|
| 4 |
A' |
1717 |
1654 |
|
|
|
|
| 5 |
A' |
1402 |
1350 |
|
|
|
|
| 6 |
A' |
1384 |
1333 |
|
|
|
|
| 7 |
A' |
1192 |
1148 |
|
|
|
|
| 8 |
A' |
1073 |
1033 |
|
|
|
|
| 9 |
A' |
584 |
563 |
|
|
|
|
| 10 |
A" |
1061 |
1022 |
|
|
|
|
| 11 |
A" |
840 |
809 |
|
|
|
|
| 12 |
A" |
629 |
606 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10175.5 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 9797.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| O2 |
-1.114 |
-0.337 |
0.000 |
| N3 |
1.158 |
-0.096 |
0.000 |
| H4 |
-0.255 |
1.472 |
0.000 |
| H5 |
-0.804 |
-1.253 |
0.000 |
| H6 |
1.862 |
0.633 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3459 | 1.2674 | 1.0833 | 1.8557 | 1.8746 |
O2 | 1.3459 | | 2.2846 | 2.0018 | 0.9676 | 3.1301 | N3 | 1.2674 | 2.2846 | | 2.1108 | 2.2782 | 1.0138 | H4 | 1.0833 | 2.0018 | 2.1108 | | 2.7799 | 2.2775 | H5 | 1.8557 | 0.9676 | 2.2782 | 2.7799 | | 3.2665 | H6 | 1.8746 | 3.1301 | 1.0138 | 2.2775 | 3.2665 | |
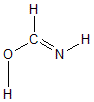 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
105.496 |
|
C1 |
N3 |
H6 |
110.023 |
| O2 |
C1 |
N3 |
121.872 |
|
O2 |
C1 |
H4 |
110.526 |
| N3 |
C1 |
H4 |
127.602 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.667633 |
| Energy at 298.15K | -169.671725 |
| HF Energy | -168.975273 |
| Nuclear repulsion energy | 70.759620 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3795 |
3655 |
|
|
|
|
| 2 |
A' |
3455 |
3326 |
|
|
|
|
| 3 |
A' |
3203 |
3084 |
|
|
|
|
| 4 |
A' |
1711 |
1647 |
|
|
|
|
| 5 |
A' |
1415 |
1362 |
|
|
|
|
| 6 |
A' |
1360 |
1310 |
|
|
|
|
| 7 |
A' |
1127 |
1085 |
|
|
|
|
| 8 |
A' |
1082 |
1042 |
|
|
|
|
| 9 |
A' |
587 |
565 |
|
|
|
|
| 10 |
A" |
1087 |
1047 |
|
|
|
|
| 11 |
A" |
853 |
821 |
|
|
|
|
| 12 |
A" |
516 |
497 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10095.5 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 9721.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.442 |
0.000 |
| O2 |
-1.102 |
-0.352 |
0.000 |
| N3 |
1.226 |
0.121 |
0.000 |
| H4 |
-0.296 |
1.480 |
0.000 |
| H5 |
-0.811 |
-1.272 |
0.000 |
| H6 |
1.348 |
-0.892 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3583 | 1.2667 | 1.0801 | 1.8958 | 1.8958 |
O2 | 1.3583 | | 2.3756 | 2.0018 | 0.9650 | 2.5088 | N3 | 1.2667 | 2.3756 | | 2.0402 | 2.4677 | 1.0204 | H4 | 1.0801 | 2.0018 | 2.0402 | | 2.8000 | 2.8859 | H5 | 1.8958 | 0.9650 | 2.4677 | 2.8000 | | 2.1921 | H6 | 1.8958 | 2.5088 | 1.0204 | 2.8859 | 2.1921 | |
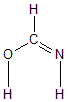 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.182 |
|
C1 |
N3 |
H6 |
111.518 |
| O2 |
C1 |
N3 |
129.608 |
|
O2 |
C1 |
H4 |
109.830 |
| N3 |
C1 |
H4 |
120.562 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
