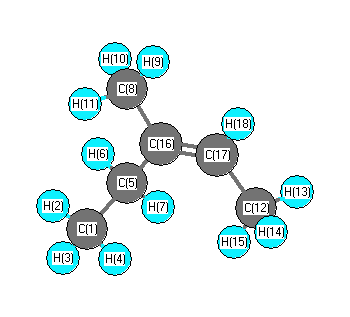
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3155 |
3019 |
50.76 |
|
|
|
| 2 |
A |
3138 |
3003 |
16.65 |
|
|
|
| 3 |
A |
3134 |
3000 |
26.87 |
|
|
|
| 4 |
A |
3133 |
2999 |
18.84 |
|
|
|
| 5 |
A |
3129 |
2994 |
36.62 |
|
|
|
| 6 |
A |
3105 |
2971 |
5.22 |
|
|
|
| 7 |
A |
3089 |
2956 |
0.82 |
|
|
|
| 8 |
A |
3088 |
2955 |
44.30 |
|
|
|
| 9 |
A |
3057 |
2926 |
28.60 |
|
|
|
| 10 |
A |
3048 |
2917 |
24.92 |
|
|
|
| 11 |
A |
3043 |
2912 |
44.91 |
|
|
|
| 12 |
A |
3037 |
2907 |
26.89 |
|
|
|
| 13 |
A |
1762 |
1686 |
0.82 |
|
|
|
| 14 |
A |
1514 |
1449 |
9.93 |
|
|
|
| 15 |
A |
1502 |
1438 |
5.56 |
|
|
|
| 16 |
A |
1498 |
1434 |
1.30 |
|
|
|
| 17 |
A |
1494 |
1430 |
8.17 |
|
|
|
| 18 |
A |
1481 |
1418 |
5.03 |
|
|
|
| 19 |
A |
1480 |
1416 |
5.02 |
|
|
|
| 20 |
A |
1474 |
1411 |
3.43 |
|
|
|
| 21 |
A |
1414 |
1353 |
1.79 |
|
|
|
| 22 |
A |
1412 |
1351 |
3.14 |
|
|
|
| 23 |
A |
1404 |
1344 |
6.29 |
|
|
|
| 24 |
A |
1386 |
1326 |
1.17 |
|
|
|
| 25 |
A |
1334 |
1277 |
2.90 |
|
|
|
| 26 |
A |
1298 |
1242 |
1.68 |
|
|
|
| 27 |
A |
1236 |
1183 |
1.78 |
|
|
|
| 28 |
A |
1143 |
1094 |
5.26 |
|
|
|
| 29 |
A |
1103 |
1056 |
3.03 |
|
|
|
| 30 |
A |
1079 |
1032 |
2.06 |
|
|
|
| 31 |
A |
1061 |
1016 |
1.91 |
|
|
|
| 32 |
A |
1053 |
1007 |
1.98 |
|
|
|
| 33 |
A |
1017 |
974 |
8.38 |
|
|
|
| 34 |
A |
1012 |
968 |
1.02 |
|
|
|
| 35 |
A |
926 |
886 |
6.01 |
|
|
|
| 36 |
A |
844 |
807 |
12.50 |
|
|
|
| 37 |
A |
793 |
759 |
1.47 |
|
|
|
| 38 |
A |
754 |
722 |
0.89 |
|
|
|
| 39 |
A |
570 |
546 |
3.03 |
|
|
|
| 40 |
A |
520 |
498 |
2.68 |
|
|
|
| 41 |
A |
391 |
374 |
0.30 |
|
|
|
| 42 |
A |
307 |
294 |
0.40 |
|
|
|
| 43 |
A |
271 |
259 |
1.22 |
|
|
|
| 44 |
A |
246 |
235 |
1.11 |
|
|
|
| 45 |
A |
217 |
207 |
0.15 |
|
|
|
| 46 |
A |
177 |
169 |
0.89 |
|
|
|
| 47 |
A |
131 |
125 |
0.18 |
|
|
|
| 48 |
A |
52 |
50 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36253.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34697.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.353 |
|
|
|
| 2 |
H |
0.117 |
|
|
|
| 3 |
H |
0.117 |
|
|
|
| 4 |
H |
0.121 |
|
|
|
| 5 |
C |
-0.230 |
|
|
|
| 6 |
H |
0.113 |
|
|
|
| 7 |
H |
0.122 |
|
|
|
| 8 |
C |
-0.354 |
|
|
|
| 9 |
H |
0.111 |
|
|
|
| 10 |
H |
0.116 |
|
|
|
| 11 |
H |
0.114 |
|
|
|
| 12 |
C |
-0.349 |
|
|
|
| 13 |
H |
0.120 |
|
|
|
| 14 |
H |
0.119 |
|
|
|
| 15 |
H |
0.110 |
|
|
|
| 16 |
C |
0.062 |
|
|
|
| 17 |
C |
-0.174 |
|
|
|
| 18 |
H |
0.119 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.127 |
-0.155 |
0.058 |
0.208 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.818 |
0.309 |
0.698 |
| y |
0.309 |
-39.403 |
-0.553 |
| z |
0.698 |
-0.553 |
-40.765 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.266 |
0.309 |
0.698 |
| y |
0.309 |
0.388 |
-0.553 |
| z |
0.698 |
-0.553 |
-1.654 |
|
| Polar |
|---|
| 3z2-r2 | -3.309 |
|---|
| x2-y2 | 0.585 |
|---|
| xy | 0.309 |
|---|
| xz | 0.698 |
|---|
| yz | -0.553 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.297 |
0.391 |
0.489 |
| y |
0.391 |
10.784 |
-0.144 |
| z |
0.489 |
-0.144 |
8.651 |
<r2> (average value of r
2) Å
2
| <r2> |
207.771 |
| (<r2>)1/2 |
14.414 |