All results from a given calculation for C6H10 (3-Hexyne)
using model chemistry: CCSD(T)/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2V |
1A1 |
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -234.160501 |
| Energy at 298.15K | |
| HF Energy | -233.059473 |
| Nuclear repulsion energy | 207.036699 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.607 |
-0.580 |
| C2 |
-0.000 |
-0.607 |
-0.580 |
| C3 |
0.000 |
2.076 |
-0.552 |
| C4 |
-0.000 |
-2.076 |
-0.552 |
| C5 |
0.000 |
2.630 |
0.879 |
| C6 |
-0.000 |
-2.630 |
0.879 |
| H7 |
0.877 |
2.444 |
-1.093 |
| H8 |
-0.876 |
2.444 |
-1.093 |
| H9 |
-0.877 |
-2.444 |
-1.093 |
| H10 |
0.876 |
-2.444 |
-1.093 |
| H11 |
0.000 |
3.722 |
0.866 |
| H12 |
-0.883 |
2.289 |
1.421 |
| H13 |
0.883 |
2.289 |
1.421 |
| H14 |
-0.000 |
-3.722 |
0.866 |
| H15 |
0.883 |
-2.289 |
1.421 |
| H16 |
-0.883 |
-2.289 |
1.421 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.2137 | 1.4698 | 2.6834 | 2.4947 | 3.5509 | 2.0992 | 2.0992 | 3.2157 | 3.2152 | 3.4348 | 2.7598 | 2.7598 | 4.5643 | 3.6293 | 3.6297 |
C2 | 1.2137 | | 2.6834 | 1.4698 | 3.5509 | 2.4947 | 3.2157 | 3.2152 | 2.0992 | 2.0992 | 4.5643 | 3.6293 | 3.6297 | 3.4348 | 2.7598 | 2.7598 | C3 | 1.4698 | 2.6834 | | 4.1528 | 1.5345 | 4.9196 | 1.0936 | 1.0936 | 4.6364 | 4.6361 | 2.1728 | 2.1726 | 2.1726 | 5.9696 | 4.8716 | 4.8719 | C4 | 2.6834 | 1.4698 | 4.1528 | | 4.9196 | 1.5345 | 4.6364 | 4.6361 | 1.0936 | 1.0936 | 5.9696 | 4.8716 | 4.8719 | 2.1728 | 2.1726 | 2.1726 | C5 | 2.4947 | 3.5509 | 1.5345 | 4.9196 | | 5.2609 | 2.1655 | 2.1655 | 5.5142 | 5.5139 | 1.0918 | 1.0911 | 1.0911 | 6.3526 | 5.0275 | 5.0279 | C6 | 3.5509 | 2.4947 | 4.9196 | 1.5345 | 5.2609 | | 5.5142 | 5.5139 | 2.1655 | 2.1655 | 6.3526 | 5.0275 | 5.0279 | 1.0918 | 1.0911 | 1.0911 | H7 | 2.0992 | 3.2157 | 1.0936 | 4.6364 | 2.1655 | 5.5142 | | 1.7537 | 5.1934 | 4.8880 | 2.4980 | 3.0726 | 2.5187 | 6.5291 | 5.3594 | 5.6413 | H8 | 2.0992 | 3.2152 | 1.0936 | 4.6361 | 2.1655 | 5.5139 | 1.7537 | | 4.8880 | 5.1928 | 2.4980 | 2.5187 | 3.0726 | 6.5289 | 5.6407 | 5.3594 | H9 | 3.2157 | 2.0992 | 4.6364 | 1.0936 | 5.5142 | 2.1655 | 5.1934 | 4.8880 | | 1.7537 | 6.5291 | 5.3594 | 5.6413 | 2.4980 | 3.0726 | 2.5187 | H10 | 3.2152 | 2.0992 | 4.6361 | 1.0936 | 5.5139 | 2.1655 | 4.8880 | 5.1928 | 1.7537 | | 6.5289 | 5.6407 | 5.3594 | 2.4980 | 2.5187 | 3.0726 | H11 | 3.4348 | 4.5643 | 2.1728 | 5.9696 | 1.0918 | 6.3526 | 2.4980 | 2.4980 | 6.5291 | 6.5289 | | 1.7723 | 1.7723 | 7.4443 | 6.1011 | 6.1014 | H12 | 2.7598 | 3.6293 | 2.1726 | 4.8716 | 1.0911 | 5.0275 | 3.0726 | 2.5187 | 5.3594 | 5.6407 | 1.7723 | | 1.7660 | 6.1011 | 4.9070 | 4.5785 | H13 | 2.7598 | 3.6297 | 2.1726 | 4.8719 | 1.0911 | 5.0279 | 2.5187 | 3.0726 | 5.6413 | 5.3594 | 1.7723 | 1.7660 | | 6.1014 | 4.5785 | 4.9076 | H14 | 4.5643 | 3.4348 | 5.9696 | 2.1728 | 6.3526 | 1.0918 | 6.5291 | 6.5289 | 2.4980 | 2.4980 | 7.4443 | 6.1011 | 6.1014 | | 1.7723 | 1.7723 | H15 | 3.6293 | 2.7598 | 4.8716 | 2.1726 | 5.0275 | 1.0911 | 5.3594 | 5.6407 | 3.0726 | 2.5187 | 6.1011 | 4.9070 | 4.5785 | 1.7723 | | 1.7660 | H16 | 3.6297 | 2.7598 | 4.8719 | 2.1726 | 5.0279 | 1.0911 | 5.6413 | 5.3594 | 2.5187 | 3.0726 | 6.1014 | 4.5785 | 4.9076 | 1.7723 | 1.7660 | |
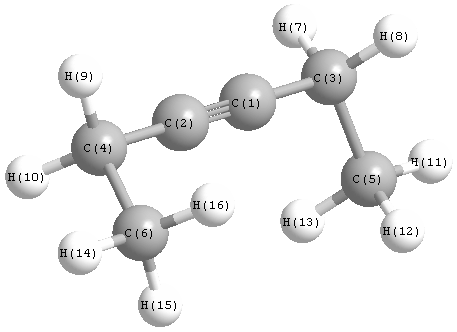 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
178.909 |
|
C1 |
C3 |
C5 |
112.255 |
| C1 |
C3 |
H7 |
109.067 |
|
C1 |
C3 |
H8 |
109.067 |
| C2 |
C1 |
C3 |
178.909 |
|
C2 |
C4 |
C6 |
112.255 |
| C2 |
C4 |
H9 |
109.067 |
|
C2 |
C4 |
H10 |
109.067 |
| C3 |
C5 |
H11 |
110.519 |
|
C3 |
C5 |
H12 |
110.540 |
| C3 |
C5 |
H13 |
110.540 |
|
C4 |
C6 |
H14 |
110.519 |
| C4 |
C6 |
H15 |
110.540 |
|
C4 |
C6 |
H16 |
110.540 |
| C5 |
C3 |
H7 |
109.838 |
|
C5 |
C3 |
H8 |
109.838 |
| C6 |
C4 |
H9 |
109.838 |
|
C6 |
C4 |
H10 |
109.838 |
| H7 |
C3 |
H8 |
106.610 |
|
H9 |
C4 |
H10 |
106.610 |
| H11 |
C5 |
H12 |
108.560 |
|
H11 |
C5 |
H13 |
108.560 |
| H12 |
C5 |
H13 |
108.046 |
|
H14 |
C6 |
H15 |
108.560 |
| H14 |
C6 |
H16 |
108.560 |
|
H15 |
C6 |
H15 |
0.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
