return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (3-Hexyne)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-234.157754
Energy at 298.15K-234.166950
HF Energy-233.059574
Nuclear repulsion energy208.553337
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3180 3019 0.00      
2 A 3175 3014 5.04      
3 A 3118 2960 0.00      
4 A 3099 2942 29.05      
5 A 3087 2931 16.00      
6 A 2299 2182 0.02      
7 A 1532 1455 1.03      
8 A 1524 1447 0.00      
9 A 1501 1425 5.01      
10 A 1420 1348 5.26      
11 A 1362 1293 0.12      
12 A 1307 1240 0.00      
13 A 1110 1054 0.00      
14 A 1109 1053 1.06      
15 A 1019 968 0.05      
16 A 793 753 0.00      
17 A 705 669 0.07      
18 A 429 407 6.56      
19 A 393 373 0.00      
20 A 242 230 0.00      
21 A 89 85 1.68      
22 A 9 8 0.00      
23 B 3180 3019 37.62      
24 B 3175 3014 45.78      
25 B 3118 2960 10.53      
26 B 3098 2942 17.57      
27 B 3087 2931 28.51      
28 B 1532 1454 3.61      
29 B 1525 1447 14.16      
30 B 1498 1422 0.63      
31 B 1420 1348 0.09      
32 B 1371 1301 27.87      
33 B 1309 1243 0.00      
34 B 1185 1125 0.14      
35 B 1124 1068 0.91      
36 B 1099 1043 4.46      
37 B 952 904 0.81      
38 B 795 755 2.69      
39 B 550 522 0.35      
40 B 307 292 3.61      
41 B 208 198 0.10      
42 B 162 154 4.87      

Unscaled Zero Point Vibrational Energy (zpe) 31596.6 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 29997.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.31220 0.04796 0.04288

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.607 -0.580
C2 -0.000 -0.607 -0.580
C3 0.000 2.060 -0.549
C4 -0.000 -2.060 -0.549
C5 -0.001 2.604 0.874
C6 0.001 -2.604 0.874
H7 0.872 2.430 -1.083
H8 -0.871 2.430 -1.084
H9 -0.872 -2.430 -1.083
H10 0.871 -2.430 -1.084
H11 -0.001 3.690 0.870
H12 -0.878 2.261 1.411
H13 0.877 2.261 1.412
H14 0.001 -3.690 0.870
H15 0.878 -2.261 1.411
H16 -0.877 -2.261 1.412

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.21341.45352.66672.47093.52502.08322.08323.19963.19963.40672.73352.73354.53423.59983.5999
C21.21342.66671.45353.52502.47093.19963.19962.08322.08324.53423.59983.59993.40672.73352.7335
C31.45352.66674.11961.52384.87641.08781.08784.60524.60522.16052.15702.15705.92184.82504.8251
C42.66671.45354.11964.87641.52384.60524.60521.08781.08785.92184.82504.82512.16052.15702.1570
C52.47093.52501.52384.87645.20852.15052.15055.47175.47221.08531.08461.08466.29384.97304.9726
C63.52502.47094.87641.52385.20855.47175.47222.15052.15056.29384.97304.97261.08531.08461.0846
H72.08323.19961.08784.60522.15055.47171.74335.16414.86082.48213.05192.50086.48285.31325.5940
H82.08323.19961.08784.60522.15055.47221.74334.86085.16382.48212.50093.05196.48325.59435.3140
H93.19962.08324.60521.08785.47172.15055.16414.86081.74336.48285.31325.59402.48213.05192.5008
H103.19962.08324.60521.08785.47222.15054.86085.16381.74336.48325.59435.31402.48212.50093.0519
H113.40674.53422.16055.92181.08536.29382.48212.48216.48286.48321.76201.76207.37926.03946.0391
H122.73353.59982.15704.82501.08464.97303.05192.50095.31325.59431.76201.75526.03944.85124.5219
H132.73353.59992.15704.82511.08464.97262.50083.05195.59405.31401.76201.75526.03914.52194.8501
H144.53423.40675.92182.16056.29381.08536.48286.48322.48212.48217.37926.03946.03911.76201.7620
H153.59982.73354.82502.15704.97301.08465.31325.59433.05192.50096.03944.85124.52191.76201.7552
H163.59992.73354.82512.15704.97261.08465.59405.31402.50083.05196.03914.52194.85011.76201.7552

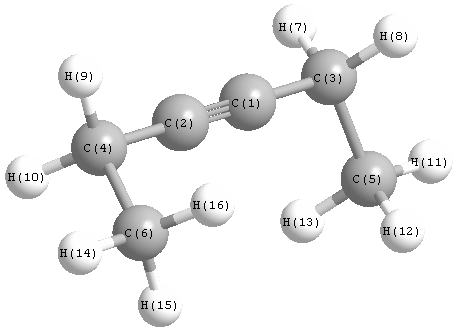
picture of 3-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 178.773 C1 C3 C5 112.161
C1 C3 H7 109.274 C1 C3 H8 109.274
C2 C1 C3 178.773 C2 C4 C6 112.161
C2 C4 H9 109.274 C2 C4 H10 109.274
C3 C5 H11 110.677 C3 C5 H12 110.442
C3 C5 H13 110.442 C4 C6 H14 110.677
C4 C6 H15 110.442 C4 C6 H16 110.442
C5 C3 H7 109.730 C5 C3 H8 109.732
C6 C4 H9 109.730 C6 C4 H10 109.732
H7 C3 H8 106.507 H9 C4 H10 106.507
H11 C5 H12 108.587 H11 C5 H13 108.586
H12 C5 H13 108.030 H14 C6 H15 108.587
H14 C6 H16 108.586 H15 C6 H15 0.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability