All results from a given calculation for C6H10 (3-Hexyne)
using model chemistry: MP4=FULL/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2V |
1A1 |
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -234.262072 |
| Energy at 298.15K | |
| HF Energy | -233.059601 |
| Nuclear repulsion energy | 207.785340 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.000 |
0.607 |
-0.575 |
| C2 |
0.000 |
-0.607 |
-0.575 |
| C3 |
-0.000 |
2.067 |
-0.550 |
| C4 |
0.000 |
-2.067 |
-0.550 |
| C5 |
-0.000 |
2.624 |
0.874 |
| C6 |
0.000 |
-2.624 |
0.874 |
| H7 |
0.874 |
2.435 |
-1.089 |
| H8 |
-0.874 |
2.435 |
-1.089 |
| H9 |
-0.874 |
-2.435 |
-1.089 |
| H10 |
0.874 |
-2.435 |
-1.089 |
| H11 |
-0.000 |
3.712 |
0.859 |
| H12 |
-0.880 |
2.285 |
1.415 |
| H13 |
0.880 |
2.285 |
1.415 |
| H14 |
0.000 |
-3.712 |
0.859 |
| H15 |
0.880 |
-2.285 |
1.415 |
| H16 |
-0.880 |
-2.285 |
1.415 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.2146 | 1.4603 | 2.6748 | 2.4832 | 3.5413 | 2.0898 | 2.0898 | 3.2065 | 3.2065 | 3.4201 | 2.7476 | 2.7476 | 4.5513 | 3.6193 | 3.6193 |
C2 | 1.2146 | | 2.6748 | 1.4603 | 3.5413 | 2.4832 | 3.2065 | 3.2065 | 2.0898 | 2.0898 | 4.5513 | 3.6193 | 3.6193 | 3.4201 | 2.7476 | 2.7476 | C3 | 1.4603 | 2.6748 | | 4.1348 | 1.5285 | 4.9027 | 1.0904 | 1.0904 | 4.6178 | 4.6178 | 2.1657 | 2.1639 | 2.1639 | 5.9488 | 4.8556 | 4.8556 | C4 | 2.6748 | 1.4603 | 4.1348 | | 4.9027 | 1.5285 | 4.6178 | 4.6178 | 1.0904 | 1.0904 | 5.9488 | 4.8556 | 4.8556 | 2.1657 | 2.1639 | 2.1639 | C5 | 2.4832 | 3.5413 | 1.5285 | 4.9027 | | 5.2481 | 2.1565 | 2.1565 | 5.4960 | 5.4960 | 1.0882 | 1.0875 | 1.0875 | 6.3362 | 5.0164 | 5.0164 | C6 | 3.5413 | 2.4832 | 4.9027 | 1.5285 | 5.2481 | | 5.4960 | 5.4960 | 2.1565 | 2.1565 | 6.3362 | 5.0164 | 5.0164 | 1.0882 | 1.0875 | 1.0875 | H7 | 2.0898 | 3.2065 | 1.0904 | 4.6178 | 2.1565 | 5.4960 | | 1.7478 | 5.1738 | 4.8696 | 2.4877 | 3.0606 | 2.5081 | 6.5071 | 5.3426 | 5.6231 | H8 | 2.0898 | 3.2065 | 1.0904 | 4.6178 | 2.1565 | 5.4960 | 1.7478 | | 4.8696 | 5.1738 | 2.4877 | 2.5081 | 3.0606 | 6.5071 | 5.6231 | 5.3426 | H9 | 3.2065 | 2.0898 | 4.6178 | 1.0904 | 5.4960 | 2.1565 | 5.1738 | 4.8696 | | 1.7478 | 6.5071 | 5.3426 | 5.6231 | 2.4877 | 3.0606 | 2.5081 | H10 | 3.2065 | 2.0898 | 4.6178 | 1.0904 | 5.4960 | 2.1565 | 4.8696 | 5.1738 | 1.7478 | | 6.5071 | 5.6231 | 5.3426 | 2.4877 | 2.5081 | 3.0606 | H11 | 3.4201 | 4.5513 | 2.1657 | 5.9488 | 1.0882 | 6.3362 | 2.4877 | 2.4877 | 6.5071 | 6.5071 | | 1.7666 | 1.7666 | 7.4243 | 6.0867 | 6.0866 | H12 | 2.7476 | 3.6193 | 2.1639 | 4.8556 | 1.0875 | 5.0164 | 3.0606 | 2.5081 | 5.3426 | 5.6231 | 1.7666 | | 1.7601 | 6.0867 | 4.8969 | 4.5696 | H13 | 2.7476 | 3.6193 | 2.1639 | 4.8556 | 1.0875 | 5.0164 | 2.5081 | 3.0606 | 5.6231 | 5.3426 | 1.7666 | 1.7601 | | 6.0866 | 4.5696 | 4.8969 | H14 | 4.5513 | 3.4201 | 5.9488 | 2.1657 | 6.3362 | 1.0882 | 6.5071 | 6.5071 | 2.4877 | 2.4877 | 7.4243 | 6.0867 | 6.0866 | | 1.7666 | 1.7666 | H15 | 3.6193 | 2.7476 | 4.8556 | 2.1639 | 5.0164 | 1.0875 | 5.3426 | 5.6231 | 3.0606 | 2.5081 | 6.0867 | 4.8969 | 4.5696 | 1.7666 | | 1.7601 | H16 | 3.6193 | 2.7476 | 4.8556 | 2.1639 | 5.0164 | 1.0875 | 5.6231 | 5.3426 | 2.5081 | 3.0606 | 6.0866 | 4.5696 | 4.8969 | 1.7666 | 1.7601 | |
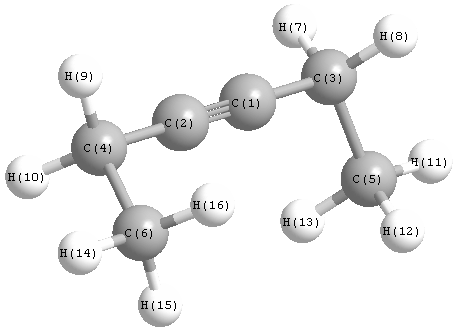 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
179.011 |
|
C1 |
C3 |
C5 |
112.346 |
| C1 |
C3 |
H7 |
109.168 |
|
C1 |
C3 |
H8 |
109.168 |
| C2 |
C1 |
C3 |
179.011 |
|
C2 |
C4 |
C6 |
112.346 |
| C2 |
C4 |
H9 |
109.168 |
|
C2 |
C4 |
H10 |
109.168 |
| C3 |
C5 |
H11 |
110.583 |
|
C3 |
C5 |
H12 |
110.486 |
| C3 |
C5 |
H13 |
110.486 |
|
C4 |
C6 |
H14 |
110.583 |
| C4 |
C6 |
H15 |
110.486 |
|
C4 |
C6 |
H16 |
110.486 |
| C5 |
C3 |
H7 |
109.723 |
|
C5 |
C3 |
H8 |
109.723 |
| C6 |
C4 |
H9 |
109.723 |
|
C6 |
C4 |
H10 |
109.723 |
| H7 |
C3 |
H8 |
106.541 |
|
H9 |
C4 |
H10 |
106.541 |
| H11 |
C5 |
H12 |
108.581 |
|
H11 |
C5 |
H13 |
108.581 |
| H12 |
C5 |
H13 |
108.049 |
|
H14 |
C6 |
H15 |
108.581 |
| H14 |
C6 |
H16 |
108.581 |
|
H15 |
C6 |
H15 |
0.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
