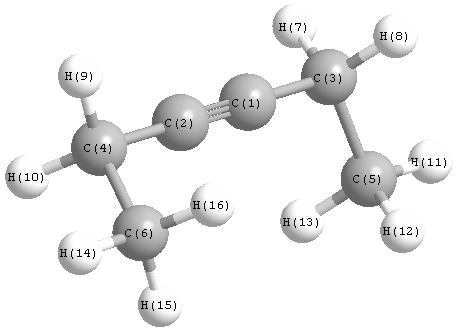
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3146 |
3011 |
0.00 |
|
|
|
| 2 |
A |
3140 |
3006 |
8.62 |
|
|
|
| 3 |
A |
3078 |
2946 |
0.00 |
|
|
|
| 4 |
A |
3065 |
2933 |
30.15 |
|
|
|
| 5 |
A |
3048 |
2917 |
24.74 |
|
|
|
| 6 |
A |
2376 |
2274 |
0.00 |
|
|
|
| 7 |
A |
1505 |
1441 |
1.49 |
|
|
|
| 8 |
A |
1493 |
1429 |
0.00 |
|
|
|
| 9 |
A |
1477 |
1414 |
5.19 |
|
|
|
| 10 |
A |
1405 |
1345 |
4.51 |
|
|
|
| 11 |
A |
1348 |
1290 |
0.04 |
|
|
|
| 12 |
A |
1285 |
1230 |
0.00 |
|
|
|
| 13 |
A |
1096 |
1049 |
0.00 |
|
|
|
| 14 |
A |
1093 |
1046 |
1.47 |
|
|
|
| 15 |
A |
1006 |
963 |
0.03 |
|
|
|
| 16 |
A |
783 |
750 |
0.00 |
|
|
|
| 17 |
A |
701 |
671 |
0.07 |
|
|
|
| 18 |
A |
424 |
405 |
6.94 |
|
|
|
| 19 |
A |
414 |
396 |
0.00 |
|
|
|
| 20 |
A |
224 |
215 |
0.00 |
|
|
|
| 21 |
A |
89 |
85 |
1.82 |
|
|
|
| 22 |
A |
15i |
14i |
0.00 |
|
|
|
| 23 |
B |
3146 |
3011 |
50.46 |
|
|
|
| 24 |
B |
3140 |
3005 |
55.16 |
|
|
|
| 25 |
B |
3077 |
2945 |
14.54 |
|
|
|
| 26 |
B |
3064 |
2933 |
29.39 |
|
|
|
| 27 |
B |
3049 |
2918 |
27.78 |
|
|
|
| 28 |
B |
1505 |
1440 |
3.49 |
|
|
|
| 29 |
B |
1494 |
1430 |
14.00 |
|
|
|
| 30 |
B |
1476 |
1413 |
0.83 |
|
|
|
| 31 |
B |
1405 |
1345 |
0.00 |
|
|
|
| 32 |
B |
1358 |
1299 |
30.13 |
|
|
|
| 33 |
B |
1288 |
1233 |
0.00 |
|
|
|
| 34 |
B |
1173 |
1122 |
1.07 |
|
|
|
| 35 |
B |
1110 |
1062 |
1.09 |
|
|
|
| 36 |
B |
1083 |
1036 |
4.88 |
|
|
|
| 37 |
B |
937 |
897 |
1.23 |
|
|
|
| 38 |
B |
783 |
749 |
3.55 |
|
|
|
| 39 |
B |
570 |
545 |
0.59 |
|
|
|
| 40 |
B |
283 |
271 |
4.63 |
|
|
|
| 41 |
B |
219 |
209 |
0.18 |
|
|
|
| 42 |
B |
155 |
149 |
4.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31246.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29906.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.015 |
|
|
|
| 2 |
C |
-0.015 |
|
|
|
| 3 |
C |
-0.253 |
|
|
|
| 4 |
C |
-0.253 |
|
|
|
| 5 |
C |
-0.342 |
|
|
|
| 6 |
C |
-0.342 |
|
|
|
| 7 |
H |
0.127 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
H |
0.127 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
H |
0.113 |
|
|
|
| 12 |
H |
0.121 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.113 |
|
|
|
| 15 |
H |
0.121 |
|
|
|
| 16 |
H |
0.121 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.113 |
0.113 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.353 |
0.000 |
0.000 |
| y |
0.000 |
-32.273 |
0.000 |
| z |
0.000 |
0.000 |
-39.196 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.618 |
0.000 |
0.000 |
| y |
0.000 |
6.501 |
0.000 |
| z |
0.000 |
0.000 |
-3.883 |
|
| Polar |
|---|
| 3z2-r2 | -7.765 |
|---|
| x2-y2 | -6.079 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.953 |
-0.001 |
0.000 |
| y |
-0.001 |
14.792 |
0.000 |
| z |
0.000 |
0.000 |
8.984 |
<r2> (average value of r
2) Å
2
| <r2> |
270.072 |
| (<r2>)1/2 |
16.434 |