return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6O (3-Methylfuran)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-268.894207
Energy at 298.15K-268.900942
HF Energy-267.758809
Nuclear repulsion energy222.340570
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3331 3163 0.22      
2 A' 3320 3152 0.27      
3 A' 3299 3132 0.60      
4 A' 3170 3009 12.50      
5 A' 3090 2934 26.62      
6 A' 1648 1565 0.78      
7 A' 1542 1464 10.72      
8 A' 1525 1448 10.80      
9 A' 1434 1361 5.27      
10 A' 1419 1348 0.92      
11 A' 1307 1241 2.10      
12 A' 1284 1219 1.54      
13 A' 1251 1187 23.37      
14 A' 1118 1061 16.95      
15 A' 1072 1018 15.49      
16 A' 1021 970 4.72      
17 A' 992 941 2.34      
18 A' 894 848 20.24      
19 A' 651 618 2.06      
20 A' 317 301 0.75      
21 A" 3155 2995 11.52      
22 A" 1510 1434 6.50      
23 A" 1076 1021 1.25      
24 A" 870 826 2.28      
25 A" 804 764 59.08      
26 A" 741 703 7.27      
27 A" 639 607 7.72      
28 A" 635 603 12.40      
29 A" 272 258 3.34      
30 A" 143 135 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 21764.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 20663.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.30117 0.11367 0.08379

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.691 -1.458 0.000
C2 -1.071 -0.157 0.000
C3 0.661 -1.451 0.000
C4 0.000 0.677 0.000
C5 1.142 -0.182 0.000
C6 -0.005 2.162 0.000
H7 -2.127 0.021 0.000
H8 1.135 -2.411 0.000
H9 2.172 0.112 0.000
H10 -1.020 2.544 0.000
H11 0.502 2.553 0.877
H12 0.502 2.553 -0.877

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
O11.35581.35142.24472.23293.68502.06172.05893.26484.01614.27624.2762
C21.35582.16191.35792.21292.55281.07063.15363.25412.70173.25443.2544
C31.35142.16192.22871.35693.67403.15231.07042.17394.33444.10234.1023
C42.24471.35792.22871.42911.48482.22593.29022.24422.12732.13082.1308
C52.23292.21291.35691.42912.60963.27472.22861.07133.47942.94292.9429
C63.68502.55283.67401.48482.60963.01494.71292.99011.08511.08581.0858
H72.06171.07063.15232.22593.27473.01494.06824.29962.75533.75433.7543
H82.05893.15361.07043.29022.22864.71294.06822.72755.40335.08055.0805
H93.26483.25412.17392.24421.07132.99014.29962.72754.01313.08503.0850
H104.01612.70174.33442.12733.47941.08512.75535.40334.01311.75711.7571
H114.27623.25444.10232.13082.94291.08583.75435.08053.08501.75711.7539
H124.27623.25444.10232.13082.94291.08583.75435.08053.08501.75711.7539

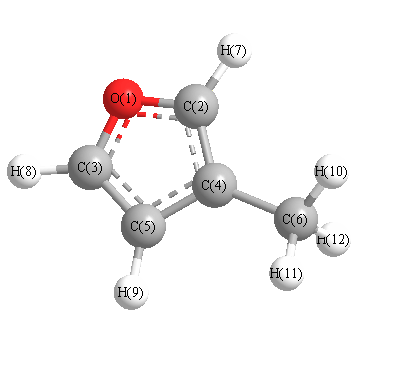
picture of 3-Methylfuran state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C4 111.623 O1 C2 H7 115.869
O1 C3 C5 111.065 O1 C3 H8 115.966
C2 O1 C3 105.988 C2 C4 C5 105.095
C2 C4 C6 127.746 C3 C5 C4 106.229
C3 C5 H9 126.683 C4 C2 H7 132.508
C4 C5 H9 127.088 C4 C6 H10 110.785
C4 C6 H11 111.022 C4 C6 H12 111.022
C5 C3 H8 132.969 C5 C4 C6 127.159
H10 C6 H11 108.073 H10 C6 H12 108.073
H11 C6 H12 107.735
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability