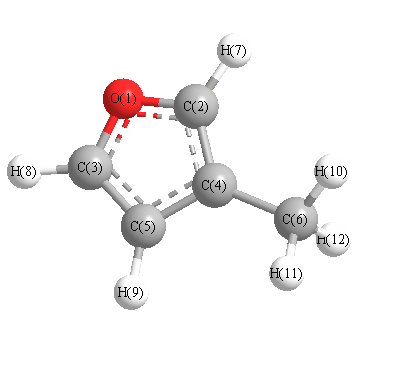
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3300 |
3159 |
0.66 |
|
|
|
| 2 |
A' |
3290 |
3149 |
0.95 |
|
|
|
| 3 |
A' |
3264 |
3124 |
1.61 |
|
|
|
| 4 |
A' |
3140 |
3005 |
16.92 |
|
|
|
| 5 |
A' |
3056 |
2924 |
31.35 |
|
|
|
| 6 |
A' |
1670 |
1598 |
1.91 |
|
|
|
| 7 |
A' |
1564 |
1497 |
14.74 |
|
|
|
| 8 |
A' |
1500 |
1436 |
7.14 |
|
|
|
| 9 |
A' |
1428 |
1367 |
5.90 |
|
|
|
| 10 |
A' |
1415 |
1354 |
0.07 |
|
|
|
| 11 |
A' |
1298 |
1243 |
5.24 |
|
|
|
| 12 |
A' |
1273 |
1218 |
2.21 |
|
|
|
| 13 |
A' |
1231 |
1178 |
16.48 |
|
|
|
| 14 |
A' |
1111 |
1063 |
19.47 |
|
|
|
| 15 |
A' |
1073 |
1027 |
17.57 |
|
|
|
| 16 |
A' |
1017 |
973 |
6.19 |
|
|
|
| 17 |
A' |
981 |
939 |
1.67 |
|
|
|
| 18 |
A' |
892 |
854 |
22.65 |
|
|
|
| 19 |
A' |
648 |
620 |
2.28 |
|
|
|
| 20 |
A' |
315 |
302 |
0.85 |
|
|
|
| 21 |
A" |
3112 |
2978 |
16.42 |
|
|
|
| 22 |
A" |
1484 |
1420 |
6.36 |
|
|
|
| 23 |
A" |
1063 |
1017 |
1.81 |
|
|
|
| 24 |
A" |
877 |
840 |
0.21 |
|
|
|
| 25 |
A" |
803 |
768 |
55.36 |
|
|
|
| 26 |
A" |
750 |
718 |
9.26 |
|
|
|
| 27 |
A" |
633 |
606 |
0.04 |
|
|
|
| 28 |
A" |
627 |
600 |
21.40 |
|
|
|
| 29 |
A" |
268 |
256 |
3.22 |
|
|
|
| 30 |
A" |
137 |
131 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21608.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20681.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.144 |
|
|
|
| 2 |
C |
-0.114 |
|
|
|
| 3 |
C |
-0.031 |
|
|
|
| 4 |
C |
0.089 |
|
|
|
| 5 |
C |
-0.236 |
|
|
|
| 6 |
C |
-0.355 |
|
|
|
| 7 |
H |
0.148 |
|
|
|
| 8 |
H |
0.150 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
| 10 |
H |
0.114 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.122 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.412 |
0.783 |
0.000 |
0.885 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.067 |
-1.958 |
0.000 |
| y |
-1.958 |
-33.285 |
0.000 |
| z |
0.000 |
0.000 |
-37.707 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.429 |
-1.958 |
0.000 |
| y |
-1.958 |
1.603 |
0.000 |
| z |
0.000 |
0.000 |
-5.031 |
|
| Polar |
|---|
| 3z2-r2 | -10.062 |
|---|
| x2-y2 | 1.217 |
|---|
| xy | -1.958 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.034 |
-0.179 |
0.000 |
| y |
-0.179 |
10.276 |
0.000 |
| z |
0.000 |
0.000 |
5.429 |
<r2> (average value of r
2) Å
2
| <r2> |
142.135 |
| (<r2>)1/2 |
11.922 |