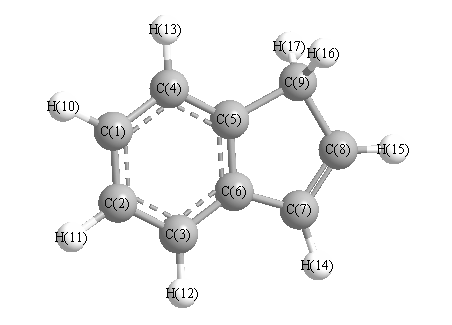
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3242 |
3103 |
14.32 |
|
|
|
| 2 |
A' |
3218 |
3080 |
5.70 |
|
|
|
| 3 |
A' |
3217 |
3079 |
19.87 |
|
|
|
| 4 |
A' |
3204 |
3067 |
27.21 |
|
|
|
| 5 |
A' |
3193 |
3056 |
2.48 |
|
|
|
| 6 |
A' |
3186 |
3050 |
4.04 |
|
|
|
| 7 |
A' |
3049 |
2918 |
11.47 |
|
|
|
| 8 |
A' |
1681 |
1609 |
2.23 |
|
|
|
| 9 |
A' |
1666 |
1594 |
0.34 |
|
|
|
| 10 |
A' |
1631 |
1561 |
0.64 |
|
|
|
| 11 |
A' |
1509 |
1444 |
10.11 |
|
|
|
| 12 |
A' |
1507 |
1442 |
5.76 |
|
|
|
| 13 |
A' |
1429 |
1368 |
13.90 |
|
|
|
| 14 |
A' |
1411 |
1351 |
1.10 |
|
|
|
| 15 |
A' |
1362 |
1304 |
4.44 |
|
|
|
| 16 |
A' |
1326 |
1269 |
0.20 |
|
|
|
| 17 |
A' |
1264 |
1210 |
2.82 |
|
|
|
| 18 |
A' |
1241 |
1188 |
2.57 |
|
|
|
| 19 |
A' |
1193 |
1142 |
0.92 |
|
|
|
| 20 |
A' |
1178 |
1127 |
0.03 |
|
|
|
| 21 |
A' |
1138 |
1090 |
0.52 |
|
|
|
| 22 |
A' |
1093 |
1046 |
1.25 |
|
|
|
| 23 |
A' |
1056 |
1011 |
3.98 |
|
|
|
| 24 |
A' |
975 |
934 |
10.48 |
|
|
|
| 25 |
A' |
876 |
839 |
3.10 |
|
|
|
| 26 |
A' |
849 |
812 |
1.27 |
|
|
|
| 27 |
A' |
746 |
714 |
2.54 |
|
|
|
| 28 |
A' |
600 |
574 |
1.48 |
|
|
|
| 29 |
A' |
541 |
518 |
0.14 |
|
|
|
| 30 |
A' |
386 |
370 |
1.28 |
|
|
|
| 31 |
A" |
3077 |
2945 |
6.17 |
|
|
|
| 32 |
A" |
1148 |
1099 |
1.82 |
|
|
|
| 33 |
A" |
1003 |
960 |
0.03 |
|
|
|
| 34 |
A" |
978 |
936 |
1.39 |
|
|
|
| 35 |
A" |
966 |
924 |
0.12 |
|
|
|
| 36 |
A" |
942 |
901 |
7.07 |
|
|
|
| 37 |
A" |
881 |
843 |
0.00 |
|
|
|
| 38 |
A" |
793 |
759 |
51.00 |
|
|
|
| 39 |
A" |
740 |
708 |
25.99 |
|
|
|
| 40 |
A" |
716 |
685 |
10.31 |
|
|
|
| 41 |
A" |
567 |
542 |
4.57 |
|
|
|
| 42 |
A" |
427 |
408 |
8.21 |
|
|
|
| 43 |
A" |
396 |
379 |
3.58 |
|
|
|
| 44 |
A" |
212 |
203 |
3.47 |
|
|
|
| 45 |
A" |
196 |
187 |
0.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31004.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29674.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.142 |
|
|
|
| 2 |
C |
-0.144 |
|
|
|
| 3 |
C |
-0.163 |
|
|
|
| 4 |
C |
-0.156 |
|
|
|
| 5 |
C |
0.082 |
|
|
|
| 6 |
C |
0.079 |
|
|
|
| 7 |
C |
-0.193 |
|
|
|
| 8 |
C |
-0.181 |
|
|
|
| 9 |
C |
-0.195 |
|
|
|
| 10 |
H |
0.126 |
|
|
|
| 11 |
H |
0.126 |
|
|
|
| 12 |
H |
0.120 |
|
|
|
| 13 |
H |
0.119 |
|
|
|
| 14 |
H |
0.132 |
|
|
|
| 15 |
H |
0.142 |
|
|
|
| 16 |
H |
0.123 |
|
|
|
| 17 |
H |
0.123 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.449 |
0.567 |
0.000 |
0.724 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.883 |
0.576 |
0.000 |
| y |
0.576 |
-46.465 |
0.000 |
| z |
0.000 |
0.000 |
-56.355 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.527 |
0.576 |
0.000 |
| y |
0.576 |
5.654 |
0.000 |
| z |
0.000 |
0.000 |
-9.181 |
|
| Polar |
|---|
| 3z2-r2 | -18.362 |
|---|
| x2-y2 | -1.418 |
|---|
| xy | 0.576 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.987 |
1.650 |
0.000 |
| y |
1.650 |
16.169 |
0.000 |
| z |
0.000 |
0.000 |
7.496 |
<r2> (average value of r
2) Å
2
| <r2> |
289.666 |
| (<r2>)1/2 |
17.020 |