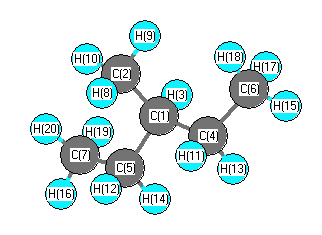
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3132 |
2998 |
78.17 |
|
|
|
| 2 |
A' |
3121 |
2987 |
38.15 |
|
|
|
| 3 |
A' |
3117 |
2983 |
54.82 |
|
|
|
| 4 |
A' |
3073 |
2941 |
44.68 |
|
|
|
| 5 |
A' |
3053 |
2922 |
14.07 |
|
|
|
| 6 |
A' |
3046 |
2915 |
27.56 |
|
|
|
| 7 |
A' |
3034 |
2904 |
66.07 |
|
|
|
| 8 |
A' |
3004 |
2875 |
8.47 |
|
|
|
| 9 |
A' |
1524 |
1459 |
4.96 |
|
|
|
| 10 |
A' |
1505 |
1441 |
12.56 |
|
|
|
| 11 |
A' |
1494 |
1430 |
13.42 |
|
|
|
| 12 |
A' |
1486 |
1422 |
0.81 |
|
|
|
| 13 |
A' |
1412 |
1352 |
3.05 |
|
|
|
| 14 |
A' |
1409 |
1348 |
6.68 |
|
|
|
| 15 |
A' |
1375 |
1316 |
1.05 |
|
|
|
| 16 |
A' |
1362 |
1303 |
0.38 |
|
|
|
| 17 |
A' |
1300 |
1245 |
1.20 |
|
|
|
| 18 |
A' |
1199 |
1147 |
0.12 |
|
|
|
| 19 |
A' |
1186 |
1135 |
0.66 |
|
|
|
| 20 |
A' |
1084 |
1037 |
0.08 |
|
|
|
| 21 |
A' |
979 |
937 |
2.82 |
|
|
|
| 22 |
A' |
954 |
913 |
1.56 |
|
|
|
| 23 |
A' |
821 |
786 |
0.80 |
|
|
|
| 24 |
A' |
773 |
739 |
4.21 |
|
|
|
| 25 |
A' |
424 |
406 |
0.14 |
|
|
|
| 26 |
A' |
385 |
368 |
0.10 |
|
|
|
| 27 |
A' |
244 |
234 |
0.01 |
|
|
|
| 28 |
A' |
193 |
185 |
0.00 |
|
|
|
| 29 |
A' |
77 |
74 |
0.00 |
|
|
|
| 30 |
A" |
3139 |
3004 |
14.19 |
|
|
|
| 31 |
A" |
3125 |
2991 |
38.89 |
|
|
|
| 32 |
A" |
3120 |
2986 |
7.21 |
|
|
|
| 33 |
A" |
3065 |
2934 |
0.20 |
|
|
|
| 34 |
A" |
3052 |
2921 |
40.68 |
|
|
|
| 35 |
A" |
3028 |
2898 |
0.46 |
|
|
|
| 36 |
A" |
1508 |
1443 |
5.20 |
|
|
|
| 37 |
A" |
1499 |
1435 |
0.27 |
|
|
|
| 38 |
A" |
1496 |
1432 |
1.56 |
|
|
|
| 39 |
A" |
1477 |
1413 |
0.07 |
|
|
|
| 40 |
A" |
1408 |
1348 |
4.10 |
|
|
|
| 41 |
A" |
1385 |
1325 |
0.00 |
|
|
|
| 42 |
A" |
1308 |
1252 |
0.25 |
|
|
|
| 43 |
A" |
1273 |
1218 |
1.95 |
|
|
|
| 44 |
A" |
1178 |
1128 |
4.13 |
|
|
|
| 45 |
A" |
1060 |
1014 |
1.96 |
|
|
|
| 46 |
A" |
1046 |
1001 |
0.99 |
|
|
|
| 47 |
A" |
990 |
947 |
6.33 |
|
|
|
| 48 |
A" |
900 |
861 |
0.01 |
|
|
|
| 49 |
A" |
764 |
731 |
0.21 |
|
|
|
| 50 |
A" |
445 |
426 |
0.20 |
|
|
|
| 51 |
A" |
346 |
331 |
0.02 |
|
|
|
| 52 |
A" |
268 |
257 |
0.01 |
|
|
|
| 53 |
A" |
211 |
202 |
0.02 |
|
|
|
| 54 |
A" |
76 |
73 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41465.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 39686.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.060 |
|
|
|
| 2 |
C |
-0.389 |
|
|
|
| 3 |
H |
0.113 |
|
|
|
| 4 |
C |
-0.207 |
|
|
|
| 5 |
C |
-0.207 |
|
|
|
| 6 |
C |
-0.381 |
|
|
|
| 7 |
C |
-0.381 |
|
|
|
| 8 |
H |
0.115 |
|
|
|
| 9 |
H |
0.123 |
|
|
|
| 10 |
H |
0.123 |
|
|
|
| 11 |
H |
0.112 |
|
|
|
| 12 |
H |
0.112 |
|
|
|
| 13 |
H |
0.111 |
|
|
|
| 14 |
H |
0.111 |
|
|
|
| 15 |
H |
0.120 |
|
|
|
| 16 |
H |
0.120 |
|
|
|
| 17 |
H |
0.114 |
|
|
|
| 18 |
H |
0.117 |
|
|
|
| 19 |
H |
0.114 |
|
|
|
| 20 |
H |
0.117 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.086 |
0.044 |
0.000 |
0.097 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.509 |
0.207 |
0.000 |
| y |
0.207 |
-41.853 |
0.000 |
| z |
0.000 |
0.000 |
-42.380 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.608 |
0.207 |
0.000 |
| y |
0.207 |
0.092 |
0.000 |
| z |
0.000 |
0.000 |
-0.699 |
|
| Polar |
|---|
| 3z2-r2 | -1.399 |
|---|
| x2-y2 | 0.344 |
|---|
| xy | 0.207 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.121 |
-0.386 |
0.000 |
| y |
-0.386 |
10.167 |
0.000 |
| z |
0.000 |
0.000 |
12.584 |
<r2> (average value of r
2) Å
2
| <r2> |
235.370 |
| (<r2>)1/2 |
15.342 |