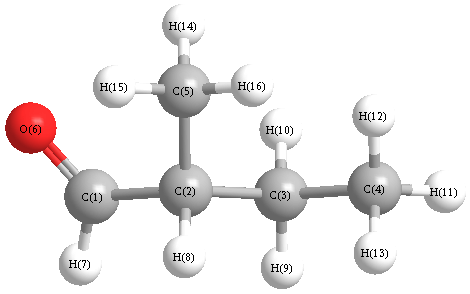
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3153 |
3018 |
16.52 |
|
|
|
| 2 |
A |
3141 |
3007 |
21.90 |
|
|
|
| 3 |
A |
3135 |
3000 |
29.47 |
|
|
|
| 4 |
A |
3127 |
2993 |
34.85 |
|
|
|
| 5 |
A |
3078 |
2946 |
17.39 |
|
|
|
| 6 |
A |
3070 |
2938 |
15.20 |
|
|
|
| 7 |
A |
3056 |
2925 |
24.41 |
|
|
|
| 8 |
A |
3041 |
2911 |
26.98 |
|
|
|
| 9 |
A |
3001 |
2872 |
16.30 |
|
|
|
| 10 |
A |
2882 |
2758 |
120.91 |
|
|
|
| 11 |
A |
1844 |
1765 |
187.93 |
|
|
|
| 12 |
A |
1504 |
1440 |
5.79 |
|
|
|
| 13 |
A |
1499 |
1435 |
9.34 |
|
|
|
| 14 |
A |
1494 |
1430 |
5.38 |
|
|
|
| 15 |
A |
1489 |
1425 |
9.06 |
|
|
|
| 16 |
A |
1483 |
1419 |
0.37 |
|
|
|
| 17 |
A |
1427 |
1366 |
8.66 |
|
|
|
| 18 |
A |
1410 |
1350 |
2.82 |
|
|
|
| 19 |
A |
1404 |
1343 |
4.54 |
|
|
|
| 20 |
A |
1373 |
1314 |
5.36 |
|
|
|
| 21 |
A |
1359 |
1301 |
1.48 |
|
|
|
| 22 |
A |
1305 |
1249 |
2.39 |
|
|
|
| 23 |
A |
1270 |
1216 |
0.60 |
|
|
|
| 24 |
A |
1194 |
1143 |
1.79 |
|
|
|
| 25 |
A |
1173 |
1123 |
3.48 |
|
|
|
| 26 |
A |
1124 |
1076 |
1.74 |
|
|
|
| 27 |
A |
1053 |
1008 |
0.12 |
|
|
|
| 28 |
A |
1016 |
972 |
15.12 |
|
|
|
| 29 |
A |
976 |
934 |
7.28 |
|
|
|
| 30 |
A |
940 |
900 |
6.97 |
|
|
|
| 31 |
A |
914 |
874 |
14.93 |
|
|
|
| 32 |
A |
798 |
764 |
7.07 |
|
|
|
| 33 |
A |
777 |
744 |
2.67 |
|
|
|
| 34 |
A |
659 |
631 |
7.51 |
|
|
|
| 35 |
A |
393 |
376 |
0.84 |
|
|
|
| 36 |
A |
384 |
367 |
0.69 |
|
|
|
| 37 |
A |
276 |
264 |
5.67 |
|
|
|
| 38 |
A |
255 |
245 |
0.97 |
|
|
|
| 39 |
A |
199 |
191 |
0.87 |
|
|
|
| 40 |
A |
185 |
177 |
1.18 |
|
|
|
| 41 |
A |
96 |
92 |
3.35 |
|
|
|
| 42 |
A |
75 |
72 |
3.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31016.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29685.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.090 |
|
|
|
| 2 |
C |
-0.069 |
|
|
|
| 3 |
C |
-0.216 |
|
|
|
| 4 |
C |
-0.369 |
|
|
|
| 5 |
C |
-0.379 |
|
|
|
| 6 |
O |
-0.251 |
|
|
|
| 7 |
H |
0.076 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
| 9 |
H |
0.117 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
H |
0.126 |
|
|
|
| 12 |
H |
0.123 |
|
|
|
| 13 |
H |
0.115 |
|
|
|
| 14 |
H |
0.127 |
|
|
|
| 15 |
H |
0.134 |
|
|
|
| 16 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.631 |
-0.467 |
0.787 |
2.785 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.009 |
-0.076 |
-1.773 |
| y |
-0.076 |
-37.157 |
-0.301 |
| z |
-1.773 |
-0.301 |
-37.382 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.740 |
-0.076 |
-1.773 |
| y |
-0.076 |
4.539 |
-0.301 |
| z |
-1.773 |
-0.301 |
4.201 |
|
| Polar |
|---|
| 3z2-r2 | 8.401 |
|---|
| x2-y2 | -8.853 |
|---|
| xy | -0.076 |
|---|
| xz | -1.773 |
|---|
| yz | -0.301 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.674 |
-0.215 |
0.237 |
| y |
-0.215 |
9.232 |
-0.236 |
| z |
0.237 |
-0.236 |
7.783 |
<r2> (average value of r
2) Å
2
| <r2> |
201.509 |
| (<r2>)1/2 |
14.195 |