return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (3-Pentanone)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-271.313513
Energy at 298.15K 
HF Energy-270.130396
Nuclear repulsion energy240.870008
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3184 3023 19.14      
2 A1 3109 2952 2.30      
3 A1 3089 2933 19.37      
4 A1 1779 1689 78.90      
5 A1 1526 1449 5.74      
6 A1 1481 1406 17.62      
7 A1 1429 1357 11.96      
8 A1 1366 1297 4.32      
9 A1 1140 1082 1.08      
10 A1 1045 992 2.03      
11 A1 803 763 2.63      
12 A1 418 397 0.42      
13 A1 199 189 0.66      
14 A2 3186 3024 0.00      
15 A2 3108 2950 0.00      
16 A2 1522 1445 0.00      
17 A2 1279 1215 0.00      
18 A2 1013 962 0.00      
19 A2 716 679 0.00      
20 A2 239 227 0.00      
21 A2 27 25 0.00      
22 B1 3186 3025 29.86      
23 B1 3118 2960 11.15      
24 B1 1522 1445 14.84      
25 B1 1323 1256 0.74      
26 B1 1150 1092 0.24      
27 B1 829 787 8.03      
28 B1 479 454 0.02      
29 B1 218 207 0.31      
30 B1 70 67 0.22      
31 B2 3183 3022 11.99      
32 B2 3109 2951 36.22      
33 B2 3081 2925 10.57      
34 B2 1526 1449 11.43      
35 B2 1467 1393 2.36      
36 B2 1429 1357 0.07      
37 B2 1410 1339 42.99      
38 B2 1165 1106 49.88      
39 B2 1033 981 13.04      
40 B2 997 946 14.77      
41 B2 634 602 4.12      
42 B2 313 298 11.77      

Unscaled Zero Point Vibrational Energy (zpe) 31449.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 29857.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.30030 0.06653 0.05670

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 1.278
C2 0.000 0.000 0.063
C3 0.000 1.279 -0.732
C4 0.000 -1.279 -0.732
C5 0.000 2.518 0.133
C6 0.000 -2.518 0.133
H7 0.867 1.256 -1.391
H8 -0.867 1.256 -1.391
H9 -0.867 -1.256 -1.391
H10 0.867 -1.256 -1.391
H11 0.000 3.416 -0.476
H12 -0.873 2.535 0.776
H13 0.873 2.535 0.776
H14 0.000 -3.416 -0.476
H15 0.873 -2.535 0.776
H16 -0.873 -2.535 0.776

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
O11.21492.38322.38322.76662.76663.07513.07513.07513.07513.83992.72802.72803.83992.72802.7280
C21.21491.50661.50662.51952.51952.10842.10842.10842.10843.45792.77432.77433.45792.77432.7743
C32.38321.50662.55861.51153.89521.08901.08902.75932.75932.15152.14792.14794.70184.19374.1937
C42.38321.50662.55863.89521.51152.75932.75931.08901.08904.70184.19374.19372.15152.14792.1479
C52.76662.51951.51153.89525.03702.16072.16074.16204.16201.08451.08391.08395.96525.16875.1687
C62.76662.51953.89521.51155.03704.16204.16202.16072.16075.96525.16875.16871.08451.08391.0839
H73.07512.10841.08902.75932.16074.16201.73333.05212.51222.50033.05912.51644.83864.36704.7006
H83.07512.10841.08902.75932.16074.16201.73332.51223.05212.50032.51643.05914.83864.70064.3670
H93.07512.10842.75931.08904.16202.16073.05212.51221.73334.83864.36704.70062.50033.05912.5164
H103.07512.10842.75931.08904.16202.16072.51223.05211.73334.83864.70064.36702.50032.51643.0591
H113.83993.45792.15154.70181.08455.96522.50032.50034.83864.83861.76181.76186.83106.14346.1434
H122.72802.77432.14794.19371.08395.16873.05912.51644.36704.70061.76181.74566.14345.36265.0706
H132.72802.77432.14794.19371.08395.16872.51643.05914.70064.36701.76181.74566.14345.07065.3626
H143.83993.45794.70182.15155.96521.08454.83864.83862.50032.50036.83106.14346.14341.76181.7618
H152.72802.77434.19372.14795.16871.08394.36704.70063.05912.51646.14345.36265.07061.76181.7456
H162.72802.77434.19372.14795.16871.08394.70064.36702.51643.05916.14345.07065.36261.76181.7456

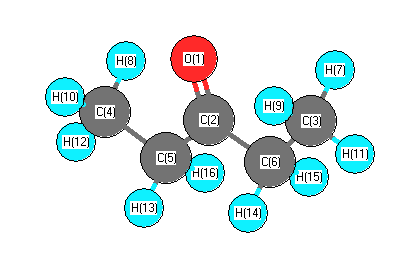
picture of 3-Pentanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 121.884 O1 C2 C4 121.884
C2 C3 C5 113.184 C2 C3 H7 107.550
C2 C3 H8 107.550 C2 C4 C6 113.184
C2 C4 H9 107.550 C2 C4 H10 107.550
C3 C2 C4 116.232 C3 C5 H11 110.875
C3 C5 H12 110.618 C3 C5 H13 110.618
C4 C6 H14 110.875 C4 C6 H15 110.618
C4 C6 H16 110.618 C5 C3 H7 111.345
C5 C3 H8 111.345 C6 C4 H9 111.345
C6 C4 H10 111.345 H7 C3 H8 105.468
H9 C4 H10 105.468 H11 C5 H12 108.682
H11 C5 H13 108.682 H12 C5 H13 107.262
H14 C6 H15 108.682 H14 C6 H16 108.682
H15 C6 H16 107.262
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability