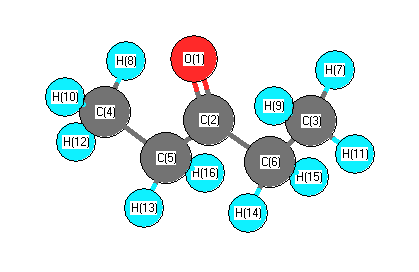
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3148 |
3013 |
27.70 |
|
|
|
| 2 |
A1 |
3073 |
2942 |
2.33 |
|
|
|
| 3 |
A1 |
3052 |
2921 |
24.31 |
|
|
|
| 4 |
A1 |
1823 |
1744 |
128.23 |
|
|
|
| 5 |
A1 |
1498 |
1433 |
6.91 |
|
|
|
| 6 |
A1 |
1459 |
1397 |
18.06 |
|
|
|
| 7 |
A1 |
1414 |
1353 |
11.73 |
|
|
|
| 8 |
A1 |
1351 |
1293 |
5.87 |
|
|
|
| 9 |
A1 |
1123 |
1075 |
1.58 |
|
|
|
| 10 |
A1 |
1027 |
983 |
2.23 |
|
|
|
| 11 |
A1 |
793 |
759 |
2.37 |
|
|
|
| 12 |
A1 |
408 |
390 |
0.47 |
|
|
|
| 13 |
A1 |
185 |
177 |
0.74 |
|
|
|
| 14 |
A2 |
3151 |
3016 |
0.00 |
|
|
|
| 15 |
A2 |
3072 |
2940 |
0.00 |
|
|
|
| 16 |
A2 |
1490 |
1426 |
0.00 |
|
|
|
| 17 |
A2 |
1261 |
1207 |
0.00 |
|
|
|
| 18 |
A2 |
997 |
954 |
0.00 |
|
|
|
| 19 |
A2 |
705 |
675 |
0.00 |
|
|
|
| 20 |
A2 |
199 |
190 |
0.00 |
|
|
|
| 21 |
A2 |
38 |
36 |
0.00 |
|
|
|
| 22 |
B1 |
3152 |
3017 |
40.94 |
|
|
|
| 23 |
B1 |
3082 |
2950 |
15.23 |
|
|
|
| 24 |
B1 |
1490 |
1426 |
15.09 |
|
|
|
| 25 |
B1 |
1303 |
1247 |
0.45 |
|
|
|
| 26 |
B1 |
1140 |
1091 |
0.11 |
|
|
|
| 27 |
B1 |
819 |
783 |
10.33 |
|
|
|
| 28 |
B1 |
466 |
446 |
0.02 |
|
|
|
| 29 |
B1 |
176 |
169 |
0.32 |
|
|
|
| 30 |
B1 |
70 |
67 |
0.30 |
|
|
|
| 31 |
B2 |
3147 |
3012 |
13.63 |
|
|
|
| 32 |
B2 |
3073 |
2941 |
43.76 |
|
|
|
| 33 |
B2 |
3044 |
2913 |
15.09 |
|
|
|
| 34 |
B2 |
1498 |
1434 |
12.76 |
|
|
|
| 35 |
B2 |
1446 |
1384 |
2.63 |
|
|
|
| 36 |
B2 |
1413 |
1353 |
0.59 |
|
|
|
| 37 |
B2 |
1395 |
1335 |
43.15 |
|
|
|
| 38 |
B2 |
1147 |
1098 |
56.61 |
|
|
|
| 39 |
B2 |
1014 |
971 |
18.46 |
|
|
|
| 40 |
B2 |
982 |
940 |
13.86 |
|
|
|
| 41 |
B2 |
627 |
600 |
4.18 |
|
|
|
| 42 |
B2 |
301 |
288 |
13.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31024.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29693.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.268 |
|
|
|
| 2 |
C |
0.196 |
|
|
|
| 3 |
C |
-0.222 |
|
|
|
| 4 |
C |
-0.222 |
|
|
|
| 5 |
C |
-0.360 |
|
|
|
| 6 |
C |
-0.360 |
|
|
|
| 7 |
H |
0.125 |
|
|
|
| 8 |
H |
0.125 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.125 |
|
|
|
| 11 |
H |
0.113 |
|
|
|
| 12 |
H |
0.127 |
|
|
|
| 13 |
H |
0.127 |
|
|
|
| 14 |
H |
0.113 |
|
|
|
| 15 |
H |
0.127 |
|
|
|
| 16 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.668 |
2.668 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.884 |
0.000 |
0.000 |
| y |
0.000 |
-36.854 |
0.000 |
| z |
0.000 |
0.000 |
-40.991 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.039 |
0.000 |
0.000 |
| y |
0.000 |
2.083 |
0.000 |
| z |
0.000 |
0.000 |
-4.122 |
|
| Polar |
|---|
| 3z2-r2 | -8.244 |
|---|
| x2-y2 | -0.030 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.502 |
0.000 |
0.000 |
| y |
0.000 |
10.879 |
0.000 |
| z |
0.000 |
0.000 |
9.175 |
<r2> (average value of r
2) Å
2
| <r2> |
212.106 |
| (<r2>)1/2 |
14.564 |