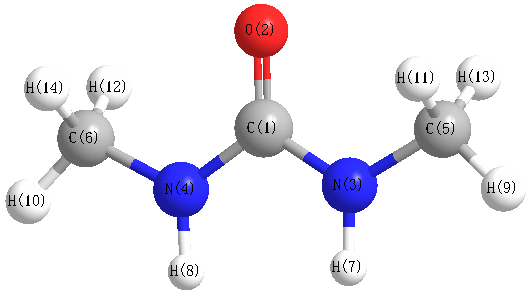
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3659 |
3510 |
21.55 |
|
|
|
| 2 |
A |
3169 |
3040 |
4.99 |
|
|
|
| 3 |
A |
3111 |
2984 |
7.38 |
|
|
|
| 4 |
A |
3034 |
2910 |
1.92 |
|
|
|
| 5 |
A |
1796 |
1723 |
260.59 |
|
|
|
| 6 |
A |
1537 |
1475 |
24.57 |
|
|
|
| 7 |
A |
1499 |
1438 |
0.01 |
|
|
|
| 8 |
A |
1487 |
1426 |
2.19 |
|
|
|
| 9 |
A |
1442 |
1383 |
12.55 |
|
|
|
| 10 |
A |
1212 |
1162 |
0.69 |
|
|
|
| 11 |
A |
1190 |
1141 |
3.23 |
|
|
|
| 12 |
A |
1150 |
1103 |
5.79 |
|
|
|
| 13 |
A |
933 |
895 |
3.35 |
|
|
|
| 14 |
A |
523 |
502 |
30.99 |
|
|
|
| 15 |
A |
401 |
385 |
52.31 |
|
|
|
| 16 |
A |
228 |
218 |
3.23 |
|
|
|
| 17 |
A |
178 |
171 |
0.00 |
|
|
|
| 18 |
A |
75 |
72 |
0.22 |
|
|
|
| 19 |
B |
3655 |
3506 |
8.51 |
|
|
|
| 20 |
B |
3169 |
3040 |
1.54 |
|
|
|
| 21 |
B |
3111 |
2984 |
54.65 |
|
|
|
| 22 |
B |
3033 |
2909 |
131.46 |
|
|
|
| 23 |
B |
1586 |
1521 |
378.27 |
|
|
|
| 24 |
B |
1511 |
1449 |
8.69 |
|
|
|
| 25 |
B |
1491 |
1431 |
75.30 |
|
|
|
| 26 |
B |
1446 |
1387 |
6.14 |
|
|
|
| 27 |
B |
1271 |
1219 |
224.07 |
|
|
|
| 28 |
B |
1159 |
1112 |
18.15 |
|
|
|
| 29 |
B |
1145 |
1098 |
12.79 |
|
|
|
| 30 |
B |
1067 |
1024 |
0.77 |
|
|
|
| 31 |
B |
779 |
748 |
11.37 |
|
|
|
| 32 |
B |
728 |
698 |
18.84 |
|
|
|
| 33 |
B |
490 |
470 |
115.68 |
|
|
|
| 34 |
B |
330 |
316 |
45.28 |
|
|
|
| 35 |
B |
121 |
116 |
10.60 |
|
|
|
| 36 |
B |
76 |
73 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26396.6 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 25319.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.308 |
|
|
|
| 2 |
O |
-0.397 |
|
|
|
| 3 |
N |
-0.181 |
|
|
|
| 4 |
N |
-0.181 |
|
|
|
| 5 |
C |
-0.164 |
|
|
|
| 6 |
C |
-0.164 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.079 |
|
|
|
| 10 |
H |
0.079 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
H |
0.115 |
|
|
|
| 13 |
H |
0.076 |
|
|
|
| 14 |
H |
0.076 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.725 |
3.725 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.442 |
-1.492 |
0.000 |
| y |
-1.492 |
-30.461 |
0.000 |
| z |
0.000 |
0.000 |
-36.856 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.784 |
-1.492 |
0.000 |
| y |
-1.492 |
6.688 |
0.000 |
| z |
0.000 |
0.000 |
-2.904 |
|
| Polar |
|---|
| 3z2-r2 | -5.809 |
|---|
| x2-y2 | -6.982 |
|---|
| xy | -1.492 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.080 |
-0.447 |
0.000 |
| y |
-0.447 |
10.561 |
0.000 |
| z |
0.000 |
0.000 |
7.842 |
<r2> (average value of r
2) Å
2
| <r2> |
191.176 |
| (<r2>)1/2 |
13.827 |