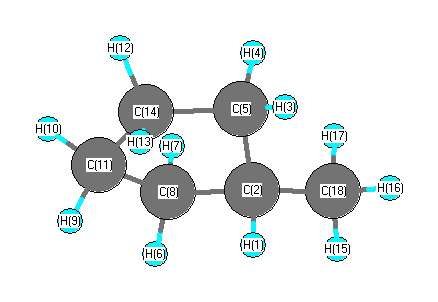
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3128 |
2994 |
87.78 |
|
|
|
| 2 |
A |
3120 |
2986 |
31.02 |
|
|
|
| 3 |
A |
3113 |
2980 |
34.21 |
|
|
|
| 4 |
A |
3109 |
2976 |
14.12 |
|
|
|
| 5 |
A |
3105 |
2972 |
5.57 |
|
|
|
| 6 |
A |
3097 |
2965 |
47.54 |
|
|
|
| 7 |
A |
3080 |
2948 |
48.59 |
|
|
|
| 8 |
A |
3068 |
2936 |
17.26 |
|
|
|
| 9 |
A |
3048 |
2917 |
80.75 |
|
|
|
| 10 |
A |
3042 |
2912 |
11.27 |
|
|
|
| 11 |
A |
3039 |
2909 |
21.13 |
|
|
|
| 12 |
A |
3017 |
2888 |
5.71 |
|
|
|
| 13 |
A |
1518 |
1453 |
1.66 |
|
|
|
| 14 |
A |
1500 |
1435 |
11.08 |
|
|
|
| 15 |
A |
1494 |
1430 |
1.58 |
|
|
|
| 16 |
A |
1493 |
1429 |
4.66 |
|
|
|
| 17 |
A |
1492 |
1428 |
2.24 |
|
|
|
| 18 |
A |
1482 |
1418 |
0.21 |
|
|
|
| 19 |
A |
1410 |
1349 |
4.82 |
|
|
|
| 20 |
A |
1391 |
1331 |
1.19 |
|
|
|
| 21 |
A |
1350 |
1292 |
0.68 |
|
|
|
| 22 |
A |
1338 |
1280 |
0.14 |
|
|
|
| 23 |
A |
1329 |
1272 |
0.04 |
|
|
|
| 24 |
A |
1319 |
1263 |
1.35 |
|
|
|
| 25 |
A |
1300 |
1244 |
0.02 |
|
|
|
| 26 |
A |
1266 |
1212 |
0.33 |
|
|
|
| 27 |
A |
1249 |
1196 |
0.48 |
|
|
|
| 28 |
A |
1219 |
1166 |
0.34 |
|
|
|
| 29 |
A |
1204 |
1152 |
0.27 |
|
|
|
| 30 |
A |
1171 |
1120 |
0.84 |
|
|
|
| 31 |
A |
1116 |
1068 |
0.00 |
|
|
|
| 32 |
A |
1063 |
1018 |
0.15 |
|
|
|
| 33 |
A |
1023 |
979 |
0.97 |
|
|
|
| 34 |
A |
1005 |
962 |
2.59 |
|
|
|
| 35 |
A |
974 |
933 |
0.81 |
|
|
|
| 36 |
A |
933 |
893 |
0.09 |
|
|
|
| 37 |
A |
926 |
887 |
0.22 |
|
|
|
| 38 |
A |
907 |
869 |
0.84 |
|
|
|
| 39 |
A |
866 |
829 |
1.15 |
|
|
|
| 40 |
A |
819 |
784 |
0.20 |
|
|
|
| 41 |
A |
757 |
725 |
0.68 |
|
|
|
| 42 |
A |
621 |
594 |
0.27 |
|
|
|
| 43 |
A |
527 |
505 |
0.47 |
|
|
|
| 44 |
A |
413 |
395 |
0.12 |
|
|
|
| 45 |
A |
297 |
284 |
0.03 |
|
|
|
| 46 |
A |
230 |
220 |
0.00 |
|
|
|
| 47 |
A |
179 |
171 |
0.01 |
|
|
|
| 48 |
A |
38 |
36 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37091.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 35500.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.113 |
|
|
|
| 2 |
C |
-0.069 |
|
|
|
| 3 |
H |
0.110 |
|
|
|
| 4 |
H |
0.105 |
|
|
|
| 5 |
C |
-0.214 |
|
|
|
| 6 |
H |
0.110 |
|
|
|
| 7 |
H |
0.105 |
|
|
|
| 8 |
C |
-0.214 |
|
|
|
| 9 |
H |
0.112 |
|
|
|
| 10 |
H |
0.111 |
|
|
|
| 11 |
C |
-0.231 |
|
|
|
| 12 |
H |
0.111 |
|
|
|
| 13 |
H |
0.112 |
|
|
|
| 14 |
C |
-0.232 |
|
|
|
| 15 |
H |
0.117 |
|
|
|
| 16 |
H |
0.117 |
|
|
|
| 17 |
H |
0.109 |
|
|
|
| 18 |
C |
-0.372 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.009 |
-0.000 |
-0.095 |
0.096 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.837 |
-0.000 |
-0.285 |
| y |
-0.000 |
-40.278 |
-0.001 |
| z |
-0.285 |
-0.001 |
-39.151 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.123 |
-0.000 |
-0.285 |
| y |
-0.000 |
-0.284 |
-0.001 |
| z |
-0.285 |
-0.001 |
1.407 |
|
| Polar |
|---|
| 3z2-r2 | 2.813 |
|---|
| x2-y2 | -0.559 |
|---|
| xy | -0.000 |
|---|
| xz | -0.285 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.345 |
0.000 |
0.049 |
| y |
0.000 |
10.200 |
0.000 |
| z |
0.049 |
0.000 |
9.088 |
<r2> (average value of r
2) Å
2
| <r2> |
176.824 |
| (<r2>)1/2 |
13.298 |