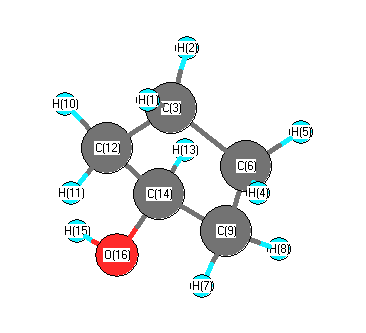
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3879 |
3713 |
28.08 |
|
|
|
| 2 |
A |
3136 |
3002 |
66.90 |
|
|
|
| 3 |
A |
3123 |
2989 |
14.82 |
|
|
|
| 4 |
A |
3110 |
2977 |
9.39 |
|
|
|
| 5 |
A |
3106 |
2973 |
41.85 |
|
|
|
| 6 |
A |
3086 |
2953 |
37.97 |
|
|
|
| 7 |
A |
3074 |
2942 |
26.34 |
|
|
|
| 8 |
A |
3064 |
2932 |
23.60 |
|
|
|
| 9 |
A |
3056 |
2925 |
30.08 |
|
|
|
| 10 |
A |
2986 |
2858 |
45.43 |
|
|
|
| 11 |
A |
1521 |
1455 |
3.55 |
|
|
|
| 12 |
A |
1495 |
1431 |
6.06 |
|
|
|
| 13 |
A |
1492 |
1428 |
2.44 |
|
|
|
| 14 |
A |
1484 |
1421 |
1.52 |
|
|
|
| 15 |
A |
1430 |
1368 |
4.09 |
|
|
|
| 16 |
A |
1380 |
1321 |
29.40 |
|
|
|
| 17 |
A |
1340 |
1283 |
0.02 |
|
|
|
| 18 |
A |
1331 |
1274 |
3.79 |
|
|
|
| 19 |
A |
1319 |
1262 |
7.83 |
|
|
|
| 20 |
A |
1299 |
1244 |
5.93 |
|
|
|
| 21 |
A |
1276 |
1221 |
5.89 |
|
|
|
| 22 |
A |
1267 |
1213 |
14.45 |
|
|
|
| 23 |
A |
1210 |
1158 |
22.77 |
|
|
|
| 24 |
A |
1205 |
1153 |
1.94 |
|
|
|
| 25 |
A |
1190 |
1139 |
3.33 |
|
|
|
| 26 |
A |
1128 |
1080 |
61.83 |
|
|
|
| 27 |
A |
1090 |
1043 |
35.89 |
|
|
|
| 28 |
A |
1063 |
1018 |
6.46 |
|
|
|
| 29 |
A |
989 |
947 |
0.37 |
|
|
|
| 30 |
A |
975 |
933 |
9.41 |
|
|
|
| 31 |
A |
959 |
918 |
2.67 |
|
|
|
| 32 |
A |
911 |
872 |
0.08 |
|
|
|
| 33 |
A |
899 |
861 |
0.05 |
|
|
|
| 34 |
A |
821 |
786 |
1.40 |
|
|
|
| 35 |
A |
771 |
738 |
1.16 |
|
|
|
| 36 |
A |
613 |
587 |
0.54 |
|
|
|
| 37 |
A |
539 |
516 |
6.46 |
|
|
|
| 38 |
A |
458 |
439 |
5.39 |
|
|
|
| 39 |
A |
359 |
344 |
13.07 |
|
|
|
| 40 |
A |
271 |
260 |
102.73 |
|
|
|
| 41 |
A |
183 |
175 |
1.12 |
|
|
|
| 42 |
A |
45 |
43 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31966.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 30594.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.113 |
|
|
|
| 2 |
H |
0.112 |
|
|
|
| 3 |
C |
-0.243 |
|
|
|
| 4 |
H |
0.114 |
|
|
|
| 5 |
H |
0.111 |
|
|
|
| 6 |
C |
-0.222 |
|
|
|
| 7 |
H |
0.116 |
|
|
|
| 8 |
H |
0.116 |
|
|
|
| 9 |
C |
-0.227 |
|
|
|
| 10 |
H |
0.104 |
|
|
|
| 11 |
H |
0.111 |
|
|
|
| 12 |
C |
-0.221 |
|
|
|
| 13 |
H |
0.084 |
|
|
|
| 14 |
C |
0.064 |
|
|
|
| 15 |
H |
0.206 |
|
|
|
| 16 |
O |
-0.337 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.535 |
-1.143 |
0.780 |
1.483 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.689 |
3.568 |
-2.000 |
| y |
3.568 |
-37.619 |
-0.266 |
| z |
-2.000 |
-0.266 |
-37.530 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.115 |
3.568 |
-2.000 |
| y |
3.568 |
0.491 |
-0.266 |
| z |
-2.000 |
-0.266 |
0.624 |
|
| Polar |
|---|
| 3z2-r2 | 1.248 |
|---|
| x2-y2 | -1.071 |
|---|
| xy | 3.568 |
|---|
| xz | -2.000 |
|---|
| yz | -0.266 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.680 |
0.171 |
0.017 |
| y |
0.171 |
9.166 |
-0.064 |
| z |
0.017 |
-0.064 |
8.004 |
<r2> (average value of r
2) Å
2
| <r2> |
164.019 |
| (<r2>)1/2 |
12.807 |