Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -341.915651 |
| Energy at 298.15K | |
| HF Energy | -340.655843 |
| Nuclear repulsion energy | 246.577037 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3127 |
2968 |
56.16 |
|
|
|
| 2 |
A1 |
1935 |
1837 |
571.45 |
|
|
|
| 3 |
A1 |
1581 |
1501 |
2.10 |
|
|
|
| 4 |
A1 |
1415 |
1343 |
0.07 |
|
|
|
| 5 |
A1 |
1231 |
1169 |
241.00 |
|
|
|
| 6 |
A1 |
983 |
933 |
12.95 |
|
|
|
| 7 |
A1 |
840 |
798 |
43.73 |
|
|
|
| 8 |
A1 |
731 |
694 |
0.12 |
|
|
|
| 9 |
A2 |
3149 |
2989 |
0.00 |
|
|
|
| 10 |
A2 |
1284 |
1219 |
0.00 |
|
|
|
| 11 |
A2 |
1187 |
1127 |
0.00 |
|
|
|
| 12 |
A2 |
188i |
178i |
0.00 |
|
|
|
| 13 |
B1 |
3172 |
3011 |
31.03 |
|
|
|
| 14 |
B1 |
1299 |
1233 |
1.97 |
|
|
|
| 15 |
B1 |
872 |
828 |
0.72 |
|
|
|
| 16 |
B1 |
776 |
737 |
19.11 |
|
|
|
| 17 |
B1 |
143 |
136 |
1.81 |
|
|
|
| 18 |
B2 |
3120 |
2962 |
19.49 |
|
|
|
| 19 |
B2 |
1566 |
1487 |
4.69 |
|
|
|
| 20 |
B2 |
1432 |
1359 |
12.01 |
|
|
|
| 21 |
B2 |
1222 |
1160 |
6.71 |
|
|
|
| 22 |
B2 |
1055 |
1002 |
302.45 |
|
|
|
| 23 |
B2 |
757 |
719 |
1.52 |
|
|
|
| 24 |
B2 |
517 |
491 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16602.7 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 15762.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.861 |
| O2 |
0.000 |
0.000 |
2.044 |
| O3 |
0.000 |
1.128 |
0.057 |
| O4 |
0.000 |
-1.128 |
0.057 |
| C5 |
0.000 |
0.769 |
-1.281 |
| C6 |
0.000 |
-0.769 |
-1.281 |
| H7 |
-0.882 |
1.173 |
-1.763 |
| H8 |
0.882 |
1.173 |
-1.763 |
| H9 |
0.882 |
-1.173 |
-1.763 |
| H10 |
-0.882 |
-1.173 |
-1.763 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1823 | 1.3854 | 1.3854 | 2.2766 | 2.2766 | 3.0076 | 3.0076 | 3.0076 | 3.0076 |
O2 | 1.1823 | | 2.2847 | 2.2847 | 3.4128 | 3.4128 | 4.0805 | 4.0805 | 4.0805 | 4.0805 | O3 | 1.3854 | 2.2847 | | 2.2557 | 1.3854 | 2.3213 | 2.0234 | 2.0234 | 3.0640 | 3.0640 | O4 | 1.3854 | 2.2847 | 2.2557 | | 2.3213 | 1.3854 | 3.0640 | 3.0640 | 2.0234 | 2.0234 | C5 | 2.2766 | 3.4128 | 1.3854 | 2.3213 | | 1.5379 | 1.0839 | 1.0839 | 2.1873 | 2.1873 | C6 | 2.2766 | 3.4128 | 2.3213 | 1.3854 | 1.5379 | | 2.1873 | 2.1873 | 1.0839 | 1.0839 | H7 | 3.0076 | 4.0805 | 2.0234 | 3.0640 | 1.0839 | 2.1873 | | 1.7649 | 2.9365 | 2.3470 | H8 | 3.0076 | 4.0805 | 2.0234 | 3.0640 | 1.0839 | 2.1873 | 1.7649 | | 2.3470 | 2.9365 | H9 | 3.0076 | 4.0805 | 3.0640 | 2.0234 | 2.1873 | 1.0839 | 2.9365 | 2.3470 | | 1.7649 | H10 | 3.0076 | 4.0805 | 3.0640 | 2.0234 | 2.1873 | 1.0839 | 2.3470 | 2.9365 | 1.7649 | |
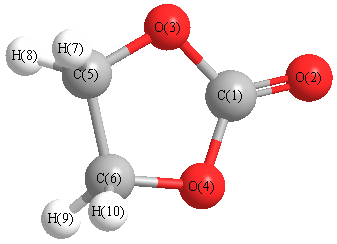 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
110.491 |
|
C1 |
O4 |
C6 |
110.491 |
| O2 |
C1 |
O3 |
125.505 |
|
O2 |
C1 |
O4 |
125.505 |
| O3 |
C1 |
O4 |
108.989 |
|
O3 |
C5 |
C6 |
105.014 |
| O3 |
C5 |
H7 |
109.446 |
|
O3 |
C5 |
H8 |
109.446 |
| O4 |
C6 |
C5 |
105.014 |
|
O4 |
C6 |
H9 |
109.446 |
| O4 |
C6 |
H10 |
109.446 |
|
C5 |
C6 |
H9 |
111.916 |
| C5 |
C6 |
H10 |
111.916 |
|
C6 |
C5 |
H7 |
111.916 |
| C6 |
C5 |
H8 |
111.916 |
|
H7 |
C5 |
H8 |
109.006 |
| H9 |
C6 |
H10 |
109.006 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -341.916805 |
| Energy at 298.15K | -341.923275 |
| HF Energy | -340.655660 |
| Nuclear repulsion energy | 247.287071 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3178 |
3017 |
16.72 |
|
|
|
| 2 |
A |
3098 |
2942 |
28.11 |
|
|
|
| 3 |
A |
1937 |
1839 |
552.68 |
|
|
|
| 4 |
A |
1564 |
1485 |
2.55 |
|
|
|
| 5 |
A |
1422 |
1350 |
4.70 |
|
|
|
| 6 |
A |
1287 |
1222 |
18.50 |
|
|
|
| 7 |
A |
1228 |
1166 |
164.13 |
|
|
|
| 8 |
A |
1167 |
1108 |
48.76 |
|
|
|
| 9 |
A |
984 |
935 |
5.70 |
|
|
|
| 10 |
A |
835 |
793 |
47.84 |
|
|
|
| 11 |
A |
727 |
691 |
0.30 |
|
|
|
| 12 |
A |
159 |
151 |
0.52 |
|
|
|
| 13 |
B |
3187 |
3026 |
15.93 |
|
|
|
| 14 |
B |
3105 |
2948 |
39.09 |
|
|
|
| 15 |
B |
1555 |
1476 |
5.91 |
|
|
|
| 16 |
B |
1419 |
1348 |
9.55 |
|
|
|
| 17 |
B |
1297 |
1232 |
5.30 |
|
|
|
| 18 |
B |
1195 |
1134 |
5.55 |
|
|
|
| 19 |
B |
1048 |
995 |
276.94 |
|
|
|
| 20 |
B |
917 |
871 |
11.49 |
|
|
|
| 21 |
B |
785 |
745 |
19.77 |
|
|
|
| 22 |
B |
679 |
644 |
0.80 |
|
|
|
| 23 |
B |
517 |
491 |
0.27 |
|
|
|
| 24 |
B |
152 |
144 |
1.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16720.2 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 15874.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.855 |
| O2 |
0.000 |
0.000 |
2.037 |
| O3 |
0.000 |
1.128 |
0.044 |
| O4 |
0.000 |
-1.128 |
0.044 |
| C5 |
0.215 |
0.729 |
-1.269 |
| C6 |
-0.215 |
-0.729 |
-1.269 |
| H7 |
-0.375 |
1.344 |
-1.936 |
| H8 |
1.269 |
0.832 |
-1.517 |
| H9 |
0.375 |
-1.344 |
-1.936 |
| H10 |
-1.269 |
-0.832 |
-1.517 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1819 | 1.3894 | 1.3894 | 2.2567 | 2.2567 | 3.1202 | 2.8161 | 3.1202 | 2.8161 |
O2 | 1.1819 | | 2.2903 | 2.2903 | 3.3930 | 3.3930 | 4.2107 | 3.8646 | 4.2107 | 3.8646 | O3 | 1.3894 | 2.2903 | | 2.2557 | 1.3894 | 2.2845 | 2.0263 | 2.0332 | 3.1887 | 2.8086 | O4 | 1.3894 | 2.2903 | 2.2557 | | 2.2845 | 1.3894 | 3.1887 | 2.8086 | 2.0263 | 2.0332 | C5 | 2.2567 | 3.3930 | 1.3894 | 2.2845 | | 1.5203 | 1.0817 | 1.0871 | 2.1829 | 2.1685 | C6 | 2.2567 | 3.3930 | 2.2845 | 1.3894 | 1.5203 | | 2.1829 | 2.1685 | 1.0817 | 1.0871 | H7 | 3.1202 | 4.2107 | 2.0263 | 3.1887 | 1.0817 | 2.1829 | | 1.7716 | 2.7900 | 2.3895 | H8 | 2.8161 | 3.8646 | 2.0332 | 2.8086 | 1.0871 | 2.1685 | 1.7716 | | 2.3895 | 3.0351 | H9 | 3.1202 | 4.2107 | 3.1887 | 2.0263 | 2.1829 | 1.0817 | 2.7900 | 2.3895 | | 1.7716 | H10 | 2.8161 | 3.8646 | 2.8086 | 2.0332 | 2.1685 | 1.0871 | 2.3895 | 3.0351 | 1.7716 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
108.602 |
|
C1 |
O4 |
C6 |
108.602 |
| O2 |
C1 |
O3 |
125.733 |
|
O2 |
C1 |
O4 |
125.733 |
| O3 |
C1 |
O4 |
108.535 |
|
O3 |
C5 |
C6 |
103.377 |
| O3 |
C5 |
H7 |
109.535 |
|
O3 |
C5 |
H8 |
109.766 |
| O4 |
C6 |
C5 |
103.377 |
|
O4 |
C6 |
H9 |
109.535 |
| O4 |
C6 |
H10 |
109.766 |
|
C5 |
C6 |
H9 |
112.976 |
| C5 |
C6 |
H10 |
111.459 |
|
C6 |
C5 |
H7 |
112.976 |
| C6 |
C5 |
H8 |
111.459 |
|
H7 |
C5 |
H8 |
109.547 |
| H9 |
C6 |
H10 |
109.547 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
