Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -342.402560 |
| Energy at 298.15K | |
| HF Energy | -342.402560 |
| Nuclear repulsion energy | 247.289949 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3076 |
2944 |
61.50 |
|
|
|
| 2 |
A1 |
1960 |
1876 |
643.13 |
|
|
|
| 3 |
A1 |
1555 |
1488 |
1.75 |
|
|
|
| 4 |
A1 |
1401 |
1341 |
0.14 |
|
|
|
| 5 |
A1 |
1238 |
1184 |
249.63 |
|
|
|
| 6 |
A1 |
964 |
923 |
8.00 |
|
|
|
| 7 |
A1 |
855 |
818 |
35.78 |
|
|
|
| 8 |
A1 |
730 |
699 |
0.99 |
|
|
|
| 9 |
A2 |
3099 |
2966 |
0.00 |
|
|
|
| 10 |
A2 |
1264 |
1210 |
0.00 |
|
|
|
| 11 |
A2 |
1173 |
1122 |
0.00 |
|
|
|
| 12 |
A2 |
171i |
164i |
0.00 |
|
|
|
| 13 |
B1 |
3122 |
2988 |
41.94 |
|
|
|
| 14 |
B1 |
1278 |
1223 |
1.95 |
|
|
|
| 15 |
B1 |
863 |
826 |
0.52 |
|
|
|
| 16 |
B1 |
777 |
744 |
22.07 |
|
|
|
| 17 |
B1 |
142 |
136 |
1.34 |
|
|
|
| 18 |
B2 |
3069 |
2937 |
25.02 |
|
|
|
| 19 |
B2 |
1542 |
1476 |
5.60 |
|
|
|
| 20 |
B2 |
1420 |
1359 |
16.49 |
|
|
|
| 21 |
B2 |
1221 |
1169 |
3.79 |
|
|
|
| 22 |
B2 |
1066 |
1020 |
303.07 |
|
|
|
| 23 |
B2 |
755 |
723 |
1.00 |
|
|
|
| 24 |
B2 |
511 |
489 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16454.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 15748.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.861 |
| O2 |
0.000 |
0.000 |
2.037 |
| O3 |
0.000 |
1.121 |
0.057 |
| O4 |
0.000 |
-1.121 |
0.057 |
| C5 |
0.000 |
0.770 |
-1.277 |
| C6 |
0.000 |
-0.770 |
-1.277 |
| H7 |
-0.883 |
1.180 |
-1.763 |
| H8 |
0.883 |
1.180 |
-1.763 |
| H9 |
0.883 |
-1.180 |
-1.763 |
| H10 |
-0.883 |
-1.180 |
-1.763 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1764 | 1.3794 | 1.3794 | 2.2719 | 2.2719 | 3.0090 | 3.0090 | 3.0090 | 3.0090 |
O2 | 1.1764 | | 2.2755 | 2.2755 | 3.4022 | 3.4022 | 4.0756 | 4.0756 | 4.0756 | 4.0756 | O3 | 1.3794 | 2.2755 | | 2.2421 | 1.3794 | 2.3138 | 2.0235 | 2.0235 | 3.0635 | 3.0635 | O4 | 1.3794 | 2.2755 | 2.2421 | | 2.3138 | 1.3794 | 3.0635 | 3.0635 | 2.0235 | 2.0235 | C5 | 2.2719 | 3.4022 | 1.3794 | 2.3138 | | 1.5390 | 1.0884 | 1.0884 | 2.1946 | 2.1946 | C6 | 2.2719 | 3.4022 | 2.3138 | 1.3794 | 1.5390 | | 2.1946 | 2.1946 | 1.0884 | 1.0884 | H7 | 3.0090 | 4.0756 | 2.0235 | 3.0635 | 1.0884 | 2.1946 | | 1.7668 | 2.9478 | 2.3596 | H8 | 3.0090 | 4.0756 | 2.0235 | 3.0635 | 1.0884 | 2.1946 | 1.7668 | | 2.3596 | 2.9478 | H9 | 3.0090 | 4.0756 | 3.0635 | 2.0235 | 2.1946 | 1.0884 | 2.9478 | 2.3596 | | 1.7668 | H10 | 3.0090 | 4.0756 | 3.0635 | 2.0235 | 2.1946 | 1.0884 | 2.3596 | 2.9478 | 1.7668 | |
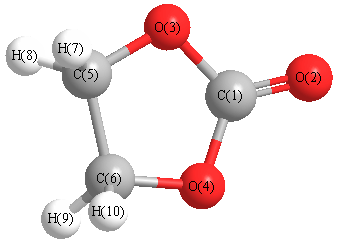 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
110.875 |
|
C1 |
O4 |
C6 |
110.875 |
| O2 |
C1 |
O3 |
125.639 |
|
O2 |
C1 |
O4 |
125.639 |
| O3 |
C1 |
O4 |
108.721 |
|
O3 |
C5 |
C6 |
104.764 |
| O3 |
C5 |
H7 |
109.600 |
|
O3 |
C5 |
H8 |
109.600 |
| O4 |
C6 |
C5 |
104.764 |
|
O4 |
C6 |
H9 |
109.600 |
| O4 |
C6 |
H10 |
109.600 |
|
C5 |
C6 |
H9 |
112.146 |
| C5 |
C6 |
H10 |
112.146 |
|
C6 |
C5 |
H7 |
112.146 |
| C6 |
C5 |
H8 |
112.146 |
|
H7 |
C5 |
H8 |
108.512 |
| H9 |
C6 |
H10 |
108.512 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.323 |
|
|
|
| 2 |
O |
-0.256 |
|
|
|
| 3 |
O |
-0.197 |
|
|
|
| 4 |
O |
-0.197 |
|
|
|
| 5 |
C |
-0.031 |
|
|
|
| 6 |
C |
-0.031 |
|
|
|
| 7 |
H |
0.097 |
|
|
|
| 8 |
H |
0.097 |
|
|
|
| 9 |
H |
0.097 |
|
|
|
| 10 |
H |
0.097 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.757 |
4.757 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.897 |
0.000 |
0.000 |
| y |
0.000 |
-35.781 |
0.000 |
| z |
0.000 |
0.000 |
-35.807 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.897 |
0.000 |
0.000 |
| y |
0.000 |
-1.929 |
0.000 |
| z |
0.000 |
0.000 |
-1.968 |
|
| Polar |
|---|
| 3z2-r2 | -3.936 |
|---|
| x2-y2 | 3.884 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.664 |
0.000 |
0.000 |
| y |
0.000 |
5.752 |
0.000 |
| z |
0.000 |
0.000 |
7.793 |
<r2> (average value of r
2) Å
2
| <r2> |
127.138 |
| (<r2>)1/2 |
11.276 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -342.403138 |
| Energy at 298.15K | -342.409522 |
| HF Energy | -342.403138 |
| Nuclear repulsion energy | 247.807707 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3128 |
2994 |
20.15 |
|
|
|
| 2 |
A |
3051 |
2920 |
32.25 |
|
|
|
| 3 |
A |
1964 |
1879 |
628.58 |
|
|
|
| 4 |
A |
1541 |
1475 |
2.08 |
|
|
|
| 5 |
A |
1405 |
1345 |
3.67 |
|
|
|
| 6 |
A |
1268 |
1214 |
27.98 |
|
|
|
| 7 |
A |
1232 |
1179 |
185.67 |
|
|
|
| 8 |
A |
1159 |
1109 |
27.92 |
|
|
|
| 9 |
A |
968 |
927 |
3.36 |
|
|
|
| 10 |
A |
850 |
814 |
38.90 |
|
|
|
| 11 |
A |
727 |
696 |
1.24 |
|
|
|
| 12 |
A |
115 |
110 |
0.31 |
|
|
|
| 13 |
B |
3138 |
3004 |
24.89 |
|
|
|
| 14 |
B |
3057 |
2926 |
46.13 |
|
|
|
| 15 |
B |
1533 |
1467 |
6.70 |
|
|
|
| 16 |
B |
1412 |
1351 |
14.32 |
|
|
|
| 17 |
B |
1280 |
1225 |
5.23 |
|
|
|
| 18 |
B |
1198 |
1146 |
3.64 |
|
|
|
| 19 |
B |
1058 |
1013 |
285.54 |
|
|
|
| 20 |
B |
901 |
863 |
5.86 |
|
|
|
| 21 |
B |
784 |
750 |
22.49 |
|
|
|
| 22 |
B |
690 |
661 |
0.53 |
|
|
|
| 23 |
B |
512 |
490 |
0.10 |
|
|
|
| 24 |
B |
147 |
141 |
1.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16559.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 15848.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.856 |
| O2 |
0.000 |
0.000 |
2.032 |
| O3 |
0.000 |
1.121 |
0.048 |
| O4 |
0.000 |
-1.121 |
0.048 |
| C5 |
0.185 |
0.740 |
-1.268 |
| C6 |
-0.185 |
-0.740 |
-1.268 |
| H7 |
-0.453 |
1.335 |
-1.915 |
| H8 |
1.226 |
0.898 |
-1.555 |
| H9 |
0.453 |
-1.335 |
-1.915 |
| H10 |
-1.226 |
-0.898 |
-1.555 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1759 | 1.3825 | 1.3825 | 2.2574 | 2.2574 | 3.1092 | 2.8507 | 3.1092 | 2.8507 |
O2 | 1.1759 | | 2.2795 | 2.2795 | 3.3875 | 3.3875 | 4.1914 | 3.8964 | 4.1914 | 3.8964 | O3 | 1.3825 | 2.2795 | | 2.2426 | 1.3825 | 2.2869 | 2.0254 | 2.0305 | 3.1765 | 2.8550 | O4 | 1.3825 | 2.2795 | 2.2426 | | 2.2869 | 1.3825 | 3.1765 | 2.8550 | 2.0254 | 2.0305 | C5 | 2.2574 | 3.3875 | 1.3825 | 2.2869 | | 1.5254 | 1.0861 | 1.0911 | 2.1896 | 2.1811 | C6 | 2.2574 | 3.3875 | 2.2869 | 1.3825 | 1.5254 | | 2.1896 | 2.1811 | 1.0861 | 1.0911 | H7 | 3.1092 | 4.1914 | 2.0254 | 3.1765 | 1.0861 | 2.1896 | | 1.7716 | 2.8194 | 2.3903 | H8 | 2.8507 | 3.8964 | 2.0305 | 2.8550 | 1.0911 | 2.1811 | 1.7716 | | 2.3903 | 3.0395 | H9 | 3.1092 | 4.1914 | 3.1765 | 2.0254 | 2.1896 | 1.0861 | 2.8194 | 2.3903 | | 1.7716 | H10 | 2.8507 | 3.8964 | 2.8550 | 2.0305 | 2.1811 | 1.0911 | 2.3903 | 3.0395 | 1.7716 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
109.448 |
|
C1 |
O4 |
C6 |
109.448 |
| O2 |
C1 |
O3 |
125.800 |
|
O2 |
C1 |
O4 |
125.800 |
| O3 |
C1 |
O4 |
108.399 |
|
O3 |
C5 |
C6 |
103.597 |
| O3 |
C5 |
H7 |
109.674 |
|
O3 |
C5 |
H8 |
109.778 |
| O4 |
C6 |
C5 |
103.597 |
|
O4 |
C6 |
H9 |
109.674 |
| O4 |
C6 |
H10 |
109.778 |
|
C5 |
C6 |
H9 |
112.879 |
| C5 |
C6 |
H10 |
111.866 |
|
C6 |
C5 |
H7 |
112.879 |
| C6 |
C5 |
H8 |
111.866 |
|
H7 |
C5 |
H8 |
108.919 |
| H9 |
C6 |
H10 |
108.919 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.328 |
|
|
|
| 2 |
O |
-0.256 |
|
|
|
| 3 |
O |
-0.202 |
|
|
|
| 4 |
O |
-0.202 |
|
|
|
| 5 |
C |
-0.033 |
|
|
|
| 6 |
C |
-0.033 |
|
|
|
| 7 |
H |
0.103 |
|
|
|
| 8 |
H |
0.096 |
|
|
|
| 9 |
H |
0.103 |
|
|
|
| 10 |
H |
0.096 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.721 |
4.721 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.981 |
0.194 |
0.000 |
| y |
0.194 |
-35.798 |
0.000 |
| z |
0.000 |
0.000 |
-35.612 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.724 |
0.194 |
0.000 |
| y |
0.194 |
-2.002 |
0.000 |
| z |
0.000 |
0.000 |
-1.723 |
|
| Polar |
|---|
| 3z2-r2 | -3.445 |
|---|
| x2-y2 | 3.817 |
|---|
| xy | 0.194 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.756 |
0.163 |
0.000 |
| y |
0.163 |
5.690 |
0.000 |
| z |
0.000 |
0.000 |
7.759 |
<r2> (average value of r
2) Å
2
| <r2> |
126.170 |
| (<r2>)1/2 |
11.233 |
