Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -343.338910 |
| Energy at 298.15K | -343.343822 |
| HF Energy | -343.338910 |
| Nuclear repulsion energy | 273.470664 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3300 |
3158 |
0.12 |
163.95 |
0.13 |
0.23 |
| 2 |
A' |
3281 |
3140 |
0.07 |
43.38 |
0.49 |
0.65 |
| 3 |
A' |
3270 |
3130 |
1.20 |
83.97 |
0.68 |
0.81 |
| 4 |
A' |
2928 |
2803 |
101.74 |
144.18 |
0.31 |
0.47 |
| 5 |
A' |
1809 |
1732 |
346.20 |
136.19 |
0.34 |
0.50 |
| 6 |
A' |
1631 |
1561 |
9.95 |
10.27 |
0.45 |
0.62 |
| 7 |
A' |
1530 |
1464 |
46.34 |
102.11 |
0.28 |
0.44 |
| 8 |
A' |
1453 |
1390 |
43.27 |
43.03 |
0.40 |
0.58 |
| 9 |
A' |
1402 |
1342 |
0.02 |
19.81 |
0.11 |
0.21 |
| 10 |
A' |
1324 |
1267 |
45.48 |
7.55 |
0.39 |
0.57 |
| 11 |
A' |
1259 |
1205 |
2.15 |
10.75 |
0.39 |
0.56 |
| 12 |
A' |
1215 |
1163 |
7.56 |
3.57 |
0.10 |
0.19 |
| 13 |
A' |
1125 |
1077 |
10.53 |
11.77 |
0.28 |
0.44 |
| 14 |
A' |
1045 |
1001 |
43.49 |
4.42 |
0.38 |
0.55 |
| 15 |
A' |
961 |
920 |
18.33 |
5.66 |
0.09 |
0.16 |
| 16 |
A' |
900 |
861 |
11.67 |
6.91 |
0.72 |
0.84 |
| 17 |
A' |
773 |
740 |
65.99 |
1.38 |
0.74 |
0.85 |
| 18 |
A' |
503 |
481 |
1.00 |
7.56 |
0.29 |
0.45 |
| 19 |
A' |
197 |
189 |
5.97 |
0.93 |
0.65 |
0.79 |
| 20 |
A" |
1019 |
975 |
0.09 |
2.51 |
0.75 |
0.86 |
| 21 |
A" |
918 |
879 |
1.15 |
0.74 |
0.75 |
0.86 |
| 22 |
A" |
862 |
825 |
4.00 |
0.17 |
0.75 |
0.86 |
| 23 |
A" |
790 |
756 |
63.37 |
0.05 |
0.75 |
0.86 |
| 24 |
A" |
665 |
636 |
0.58 |
1.77 |
0.75 |
0.86 |
| 25 |
A" |
615 |
588 |
7.60 |
0.15 |
0.75 |
0.86 |
| 26 |
A" |
297 |
284 |
12.71 |
0.48 |
0.75 |
0.86 |
| 27 |
A" |
136 |
131 |
1.50 |
1.36 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17604.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 16849.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.508 |
-0.891 |
0.000 |
| C2 |
-0.531 |
-1.733 |
0.000 |
| C3 |
-1.709 |
-1.059 |
0.000 |
| C4 |
0.000 |
0.363 |
0.000 |
| C5 |
-1.362 |
0.310 |
0.000 |
| C6 |
0.898 |
1.499 |
0.000 |
| O7 |
2.100 |
1.438 |
0.000 |
| H8 |
-0.290 |
-2.780 |
0.000 |
| H9 |
-2.692 |
-1.491 |
0.000 |
| H10 |
-2.025 |
1.157 |
0.000 |
| H11 |
0.364 |
2.467 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3380 | 2.2236 | 1.3533 | 2.2231 | 2.4218 | 2.8212 | 2.0505 | 3.2564 | 3.2576 | 3.3615 |
C2 | 1.3380 | | 1.3567 | 2.1624 | 2.2060 | 3.5341 | 4.1209 | 1.0742 | 2.1745 | 3.2530 | 4.2947 | C3 | 2.2236 | 1.3567 | | 2.2231 | 1.4129 | 3.6523 | 4.5545 | 2.2303 | 1.0740 | 2.2383 | 4.0903 | C4 | 1.3533 | 2.1624 | 2.2231 | | 1.3628 | 1.4481 | 2.3593 | 3.1562 | 3.2687 | 2.1749 | 2.1354 | C5 | 2.2231 | 2.2060 | 1.4129 | 1.3628 | | 2.5531 | 3.6409 | 3.2710 | 2.2394 | 1.0751 | 2.7621 | C6 | 2.4218 | 3.5341 | 3.6523 | 1.4481 | 2.5531 | | 1.2038 | 4.4408 | 4.6721 | 2.9427 | 1.1055 | O7 | 2.8212 | 4.1209 | 4.5545 | 2.3593 | 3.6409 | 1.2038 | | 4.8481 | 5.6165 | 4.1346 | 2.0182 | H8 | 2.0505 | 1.0742 | 2.2303 | 3.1562 | 3.2710 | 4.4408 | 4.8481 | | 2.7264 | 4.3020 | 5.2876 | H9 | 3.2564 | 2.1745 | 1.0740 | 3.2687 | 2.2394 | 4.6721 | 5.6165 | 2.7264 | | 2.7303 | 5.0005 | H10 | 3.2576 | 3.2530 | 2.2383 | 2.1749 | 1.0751 | 2.9427 | 4.1346 | 4.3020 | 2.7303 | | 2.7246 | H11 | 3.3615 | 4.2947 | 4.0903 | 2.1354 | 2.7621 | 1.1055 | 2.0182 | 5.2876 | 5.0005 | 2.7246 | |
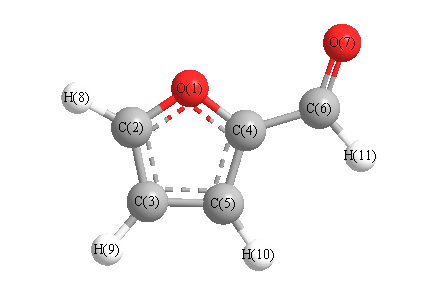 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.213 |
|
O1 |
C2 |
H8 |
116.007 |
| O1 |
C4 |
C5 |
109.864 |
|
O1 |
C4 |
C6 |
119.615 |
| C2 |
O1 |
C4 |
106.928 |
|
C2 |
C3 |
C5 |
105.573 |
| C2 |
C3 |
H9 |
126.515 |
|
C3 |
C2 |
H8 |
132.780 |
| C3 |
C5 |
C4 |
106.422 |
|
C3 |
C5 |
H10 |
127.699 |
| C4 |
C5 |
H10 |
125.879 |
|
C4 |
C6 |
O7 |
125.414 |
| C4 |
C6 |
H11 |
112.806 |
|
C5 |
C3 |
H9 |
127.912 |
| C5 |
C4 |
C6 |
130.521 |
|
O7 |
C6 |
H11 |
121.780 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.101 |
|
|
|
| 2 |
C |
-0.022 |
|
|
|
| 3 |
C |
-0.183 |
|
|
|
| 4 |
C |
0.152 |
|
|
|
| 5 |
C |
-0.198 |
|
|
|
| 6 |
C |
0.074 |
|
|
|
| 7 |
O |
-0.263 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
| 9 |
H |
0.153 |
|
|
|
| 10 |
H |
0.155 |
|
|
|
| 11 |
H |
0.071 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.936 |
-1.171 |
0.000 |
4.107 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.594 |
-2.968 |
0.000 |
| y |
-2.968 |
-33.654 |
0.000 |
| z |
0.000 |
0.000 |
-40.790 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.372 |
-2.968 |
0.000 |
| y |
-2.968 |
8.537 |
0.000 |
| z |
0.000 |
0.000 |
-2.166 |
|
| Polar |
|---|
| 3z2-r2 | -4.331 |
|---|
| x2-y2 | -9.939 |
|---|
| xy | -2.968 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.160 |
1.993 |
0.000 |
| y |
1.993 |
11.151 |
0.000 |
| z |
0.000 |
0.000 |
4.792 |
<r2> (average value of r
2) Å
2
| <r2> |
188.550 |
| (<r2>)1/2 |
13.731 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -343.339974 |
| Energy at 298.15K | -343.344868 |
| HF Energy | -343.339974 |
| Nuclear repulsion energy | 272.630582 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3300 |
3158 |
0.31 |
173.75 |
0.13 |
0.23 |
| 2 |
A' |
3289 |
3148 |
0.51 |
22.52 |
0.57 |
0.73 |
| 3 |
A' |
3274 |
3133 |
0.78 |
86.15 |
0.74 |
0.85 |
| 4 |
A' |
2943 |
2817 |
94.79 |
153.79 |
0.31 |
0.47 |
| 5 |
A' |
1804 |
1727 |
287.57 |
104.99 |
0.36 |
0.53 |
| 6 |
A' |
1638 |
1568 |
60.07 |
26.32 |
0.36 |
0.53 |
| 7 |
A' |
1520 |
1455 |
115.18 |
182.30 |
0.27 |
0.43 |
| 8 |
A' |
1461 |
1399 |
3.48 |
10.90 |
0.72 |
0.83 |
| 9 |
A' |
1410 |
1350 |
12.60 |
25.19 |
0.34 |
0.51 |
| 10 |
A' |
1280 |
1225 |
26.79 |
5.25 |
0.33 |
0.50 |
| 11 |
A' |
1259 |
1205 |
1.14 |
4.33 |
0.40 |
0.57 |
| 12 |
A' |
1201 |
1149 |
13.93 |
4.92 |
0.15 |
0.25 |
| 13 |
A' |
1132 |
1083 |
24.13 |
15.21 |
0.27 |
0.43 |
| 14 |
A' |
1040 |
995 |
40.99 |
3.22 |
0.32 |
0.48 |
| 15 |
A' |
976 |
934 |
7.88 |
6.13 |
0.13 |
0.24 |
| 16 |
A' |
900 |
861 |
7.15 |
5.90 |
0.75 |
0.86 |
| 17 |
A' |
760 |
727 |
75.54 |
2.67 |
0.64 |
0.78 |
| 18 |
A' |
501 |
480 |
0.71 |
5.94 |
0.33 |
0.49 |
| 19 |
A' |
202 |
193 |
6.90 |
0.33 |
0.37 |
0.54 |
| 20 |
A" |
1027 |
983 |
0.03 |
3.80 |
0.75 |
0.86 |
| 21 |
A" |
925 |
886 |
1.45 |
0.86 |
0.75 |
0.86 |
| 22 |
A" |
869 |
831 |
5.43 |
0.02 |
0.75 |
0.86 |
| 23 |
A" |
789 |
756 |
59.10 |
0.19 |
0.75 |
0.86 |
| 24 |
A" |
654 |
626 |
0.66 |
0.72 |
0.75 |
0.86 |
| 25 |
A" |
616 |
590 |
10.25 |
0.28 |
0.75 |
0.86 |
| 26 |
A" |
254 |
243 |
12.77 |
1.52 |
0.75 |
0.86 |
| 27 |
A" |
159 |
152 |
2.83 |
1.10 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17591.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 16836.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.235 |
-0.284 |
0.000 |
| C2 |
1.055 |
-1.610 |
0.000 |
| C3 |
-0.267 |
-1.920 |
0.000 |
| C4 |
0.000 |
0.284 |
0.000 |
| C5 |
-0.954 |
-0.686 |
0.000 |
| C6 |
-0.059 |
1.731 |
0.000 |
| O7 |
-1.087 |
2.360 |
0.000 |
| H8 |
1.946 |
-2.209 |
0.000 |
| H9 |
-0.689 |
-2.908 |
0.000 |
| H10 |
-2.015 |
-0.513 |
0.000 |
| H11 |
0.929 |
2.226 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3383 | 2.2209 | 1.3594 | 2.2263 | 2.3950 | 3.5195 | 2.0521 | 3.2536 | 3.2578 | 2.5285 |
C2 | 1.3383 | | 1.3575 | 2.1676 | 2.2111 | 3.5219 | 4.5113 | 1.0741 | 2.1735 | 3.2592 | 3.8380 | C3 | 2.2209 | 1.3575 | | 2.2197 | 1.4119 | 3.6569 | 4.3580 | 2.2320 | 1.0743 | 2.2431 | 4.3146 | C4 | 1.3594 | 2.1676 | 2.2197 | | 1.3610 | 1.4486 | 2.3440 | 3.1625 | 3.2650 | 2.1666 | 2.1527 | C5 | 2.2263 | 2.2111 | 1.4119 | 1.3610 | | 2.5781 | 3.0497 | 3.2759 | 2.2371 | 1.0742 | 3.4680 | C6 | 2.3950 | 3.5219 | 3.6569 | 1.4486 | 2.5781 | | 1.2055 | 4.4211 | 4.6815 | 2.9770 | 1.1046 | O7 | 3.5195 | 4.5113 | 4.3580 | 2.3440 | 3.0497 | 1.2055 | | 5.4846 | 5.2832 | 3.0197 | 2.0203 | H8 | 2.0521 | 1.0741 | 2.2320 | 3.1625 | 3.2759 | 4.4211 | 5.4846 | | 2.7261 | 4.3084 | 4.5502 | H9 | 3.2536 | 2.1735 | 1.0743 | 3.2650 | 2.2371 | 4.6815 | 5.2832 | 2.7261 | | 2.7368 | 5.3824 | H10 | 3.2578 | 3.2592 | 2.2431 | 2.1666 | 1.0742 | 2.9770 | 3.0197 | 4.3084 | 2.7368 | | 4.0207 | H11 | 2.5285 | 3.8380 | 4.3146 | 2.1527 | 3.4680 | 1.1046 | 2.0203 | 4.5502 | 5.3824 | 4.0207 | |
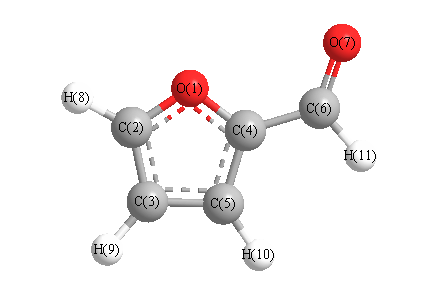 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
110.946 |
|
O1 |
C2 |
H8 |
116.145 |
| O1 |
C4 |
C5 |
109.842 |
|
O1 |
C4 |
C6 |
117.021 |
| C2 |
O1 |
C4 |
106.931 |
|
C2 |
C3 |
C5 |
105.942 |
| C2 |
C3 |
H9 |
126.314 |
|
C3 |
C2 |
H8 |
132.909 |
| C3 |
C5 |
C4 |
106.339 |
|
C3 |
C5 |
H10 |
128.401 |
| C4 |
C5 |
H10 |
125.260 |
|
C4 |
C6 |
O7 |
123.797 |
| C4 |
C6 |
H11 |
114.271 |
|
C5 |
C3 |
H9 |
127.744 |
| C5 |
C4 |
C6 |
133.137 |
|
O7 |
C6 |
H11 |
121.931 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.127 |
|
|
|
| 2 |
C |
-0.033 |
|
|
|
| 3 |
C |
-0.170 |
|
|
|
| 4 |
C |
0.147 |
|
|
|
| 5 |
C |
-0.185 |
|
|
|
| 6 |
C |
0.079 |
|
|
|
| 7 |
O |
-0.271 |
|
|
|
| 8 |
H |
0.161 |
|
|
|
| 9 |
H |
0.153 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
H |
0.080 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.498 |
-3.227 |
0.000 |
3.558 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.226 |
3.940 |
0.000 |
| y |
3.940 |
-39.882 |
0.000 |
| z |
0.000 |
0.000 |
-40.761 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.095 |
3.940 |
0.000 |
| y |
3.940 |
-0.388 |
0.000 |
| z |
0.000 |
0.000 |
-1.707 |
|
| Polar |
|---|
| 3z2-r2 | -3.413 |
|---|
| x2-y2 | 1.656 |
|---|
| xy | 3.940 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.290 |
-1.085 |
0.000 |
| y |
-1.085 |
12.995 |
0.000 |
| z |
0.000 |
0.000 |
4.772 |
<r2> (average value of r
2) Å
2
| <r2> |
190.909 |
| (<r2>)1/2 |
13.817 |
