Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -384.243217 |
| Energy at 298.15K | -384.251698 |
| HF Energy | -382.604161 |
| Nuclear repulsion energy | 403.848038 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3259 |
3094 |
5.94 |
|
|
|
| 2 |
A |
3254 |
3090 |
8.21 |
|
|
|
| 3 |
A |
3243 |
3079 |
6.24 |
|
|
|
| 4 |
A |
3229 |
3066 |
5.28 |
|
|
|
| 5 |
A |
3214 |
3051 |
0.04 |
|
|
|
| 6 |
A |
3209 |
3047 |
6.86 |
|
|
|
| 7 |
A |
3168 |
3008 |
3.75 |
|
|
|
| 8 |
A |
3098 |
2941 |
1.78 |
|
|
|
| 9 |
A |
1751 |
1662 |
107.24 |
|
|
|
| 10 |
A |
1663 |
1579 |
10.92 |
|
|
|
| 11 |
A |
1641 |
1558 |
7.76 |
|
|
|
| 12 |
A |
1535 |
1458 |
2.32 |
|
|
|
| 13 |
A |
1505 |
1429 |
10.43 |
|
|
|
| 14 |
A |
1494 |
1418 |
13.29 |
|
|
|
| 15 |
A |
1492 |
1417 |
5.39 |
|
|
|
| 16 |
A |
1481 |
1406 |
11.31 |
|
|
|
| 17 |
A |
1397 |
1326 |
73.04 |
|
|
|
| 18 |
A |
1344 |
1276 |
4.53 |
|
|
|
| 19 |
A |
1315 |
1248 |
131.18 |
|
|
|
| 20 |
A |
1213 |
1152 |
7.70 |
|
|
|
| 21 |
A |
1198 |
1137 |
1.01 |
|
|
|
| 22 |
A |
1116 |
1060 |
6.70 |
|
|
|
| 23 |
A |
1109 |
1052 |
1.68 |
|
|
|
| 24 |
A |
1057 |
1003 |
7.17 |
|
|
|
| 25 |
A |
1056 |
1002 |
0.05 |
|
|
|
| 26 |
A |
1028 |
976 |
0.70 |
|
|
|
| 27 |
A |
1021 |
969 |
0.50 |
|
|
|
| 28 |
A |
1006 |
955 |
0.00 |
|
|
|
| 29 |
A |
973 |
924 |
19.54 |
|
|
|
| 30 |
A |
964 |
916 |
3.13 |
|
|
|
| 31 |
A |
886 |
841 |
0.08 |
|
|
|
| 32 |
A |
800 |
760 |
36.38 |
|
|
|
| 33 |
A |
749 |
711 |
1.95 |
|
|
|
| 34 |
A |
727 |
690 |
33.83 |
|
|
|
| 35 |
A |
625 |
593 |
0.61 |
|
|
|
| 36 |
A |
611 |
580 |
11.47 |
|
|
|
| 37 |
A |
598 |
567 |
23.75 |
|
|
|
| 38 |
A |
469 |
445 |
0.47 |
|
|
|
| 39 |
A |
438 |
416 |
0.40 |
|
|
|
| 40 |
A |
418 |
397 |
0.01 |
|
|
|
| 41 |
A |
369 |
351 |
0.76 |
|
|
|
| 42 |
A |
221 |
210 |
4.65 |
|
|
|
| 43 |
A |
188 |
178 |
0.19 |
|
|
|
| 44 |
A |
160 |
152 |
0.00 |
|
|
|
| 45 |
A |
61 |
58 |
3.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30675.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 29123.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.444 |
0.846 |
0.716 |
| C2 |
1.695 |
-0.141 |
-0.126 |
| C3 |
0.211 |
-0.054 |
-0.056 |
| O4 |
2.247 |
-1.004 |
-0.780 |
| C5 |
-0.443 |
1.143 |
-0.320 |
| C6 |
-0.527 |
-1.187 |
0.259 |
| C7 |
-1.827 |
1.204 |
-0.280 |
| C8 |
-1.910 |
-1.120 |
0.323 |
| C9 |
-2.561 |
0.074 |
0.050 |
| H10 |
3.513 |
0.707 |
0.606 |
| H11 |
2.179 |
1.862 |
0.439 |
| H12 |
2.165 |
0.720 |
1.759 |
| H13 |
0.125 |
2.022 |
-0.576 |
| H14 |
-0.016 |
-2.116 |
0.447 |
| H15 |
-2.331 |
2.129 |
-0.502 |
| H16 |
-2.478 |
-1.999 |
0.575 |
| H17 |
-3.635 |
0.124 |
0.092 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
C5 |
C6 |
C7 |
C8 |
C9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.4982 | 2.5282 | 2.3869 | 3.0819 | 3.6289 | 4.4006 | 4.7934 | 5.1085 | 1.0832 | 1.0869 | 1.0870 | 2.9034 | 3.8594 | 5.0923 | 5.6865 | 6.1542 |
C2 | 1.4982 | | 1.4877 | 1.2151 | 2.5014 | 2.4854 | 3.7731 | 3.7619 | 4.2654 | 2.1356 | 2.1373 | 2.1248 | 2.7100 | 2.6743 | 4.6367 | 4.6207 | 5.3413 | C3 | 2.5282 | 1.4877 | | 2.3601 | 1.3899 | 1.3881 | 2.4058 | 2.4037 | 2.7778 | 3.4523 | 2.7912 | 2.7764 | 2.1419 | 2.1339 | 3.3804 | 3.3778 | 3.8537 | O4 | 2.3869 | 1.2151 | 2.3601 | | 3.4720 | 2.9677 | 4.6604 | 4.3020 | 4.9972 | 2.5397 | 3.1151 | 3.0697 | 3.7008 | 2.8038 | 5.5538 | 5.0148 | 6.0527 | C5 | 3.0819 | 2.5014 | 1.3899 | 3.4720 | | 2.4026 | 1.3861 | 2.7727 | 2.4018 | 4.0860 | 2.8228 | 3.3619 | 1.0770 | 3.3750 | 2.1374 | 3.8489 | 3.3767 | C6 | 3.6289 | 2.4854 | 1.3881 | 2.9677 | 2.4026 | | 2.7744 | 1.3859 | 2.4029 | 4.4754 | 4.0810 | 3.6237 | 3.3791 | 1.0765 | 3.8506 | 2.1364 | 3.3778 | C7 | 4.4006 | 3.7731 | 2.4058 | 4.6604 | 1.3861 | 2.7744 | | 2.4023 | 1.3873 | 5.4357 | 4.1232 | 4.5087 | 2.1372 | 3.8507 | 1.0763 | 3.3781 | 2.1390 | C8 | 4.7934 | 3.7619 | 2.4037 | 4.3020 | 2.7727 | 1.3859 | 2.4023 | | 1.3876 | 5.7294 | 5.0625 | 4.6958 | 3.8496 | 2.1433 | 3.3782 | 1.0762 | 2.1398 | C9 | 5.1085 | 4.2654 | 2.7778 | 4.9972 | 2.4018 | 2.4029 | 1.3873 | 1.3876 | | 6.1326 | 5.0817 | 5.0671 | 3.3769 | 3.3813 | 2.1399 | 2.1400 | 1.0759 | H10 | 1.0832 | 2.1356 | 3.4523 | 2.5397 | 4.0860 | 4.4754 | 5.4357 | 5.7294 | 6.1326 | | 1.7723 | 1.7742 | 3.8209 | 4.5219 | 6.1152 | 6.5735 | 7.1906 | H11 | 1.0869 | 2.1373 | 2.7912 | 3.1151 | 2.8228 | 4.0810 | 4.1232 | 5.0625 | 5.0817 | 1.7723 | | 1.7464 | 2.2962 | 4.5436 | 4.6147 | 6.0510 | 6.0789 | H12 | 1.0870 | 2.1248 | 2.7764 | 3.0697 | 3.3619 | 3.6237 | 4.5087 | 4.6958 | 5.0671 | 1.7742 | 1.7464 | | 3.3625 | 3.8103 | 5.2258 | 5.5086 | 6.0645 | H13 | 2.9034 | 2.7100 | 2.1419 | 3.7008 | 1.0770 | 3.3791 | 2.1372 | 3.8496 | 3.3769 | 3.8209 | 2.2962 | 3.3625 | | 4.2642 | 2.4595 | 4.9259 | 4.2651 | H14 | 3.8594 | 2.6743 | 2.1339 | 2.8038 | 3.3750 | 1.0765 | 3.8507 | 2.1433 | 3.3813 | 4.5219 | 4.5436 | 3.8103 | 4.2642 | | 4.9269 | 2.4680 | 4.2712 | H15 | 5.0923 | 4.6367 | 3.3804 | 5.5538 | 2.1374 | 3.8506 | 1.0763 | 3.3782 | 2.1399 | 6.1152 | 4.6147 | 5.2258 | 2.4595 | 4.9269 | | 4.2682 | 2.4647 | H16 | 5.6865 | 4.6207 | 3.3778 | 5.0148 | 3.8489 | 2.1364 | 3.3781 | 1.0762 | 2.1400 | 6.5735 | 6.0510 | 5.5086 | 4.9259 | 2.4680 | 4.2682 | | 2.4657 | H17 | 6.1542 | 5.3413 | 3.8537 | 6.0527 | 3.3767 | 3.3778 | 2.1390 | 2.1398 | 1.0759 | 7.1906 | 6.0789 | 6.0645 | 4.2651 | 4.2712 | 2.4647 | 2.4657 | |
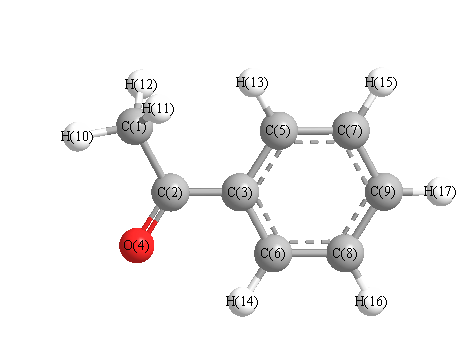 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
115.712 |
|
C1 |
C2 |
O4 |
122.867 |
| C2 |
C1 |
H10 |
110.619 |
|
C2 |
C1 |
H11 |
110.532 |
| C2 |
C1 |
H12 |
109.526 |
|
C2 |
C3 |
C5 |
120.711 |
| C2 |
C3 |
C6 |
119.555 |
|
C3 |
C2 |
O4 |
121.334 |
| C3 |
C5 |
C7 |
120.145 |
|
C3 |
C5 |
H13 |
119.980 |
| C3 |
C6 |
C8 |
120.109 |
|
C3 |
C6 |
H14 |
119.405 |
| C5 |
C3 |
C6 |
119.728 |
|
C5 |
C7 |
C9 |
119.996 |
| C5 |
C7 |
H15 |
119.938 |
|
C6 |
C8 |
C9 |
120.077 |
| C6 |
C8 |
H16 |
119.860 |
|
C7 |
C5 |
H13 |
119.864 |
| C7 |
C9 |
C8 |
119.928 |
|
C7 |
C9 |
H17 |
120.008 |
| C8 |
C6 |
H14 |
120.486 |
|
C8 |
C9 |
H17 |
120.063 |
| C9 |
C7 |
H15 |
120.065 |
|
C9 |
C8 |
H16 |
120.060 |
| H10 |
C1 |
H11 |
109.508 |
|
H10 |
C1 |
H12 |
109.677 |
| H11 |
C1 |
H12 |
106.904 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
