return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5NO2 (Nitrobenzene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-436.083495
Energy at 298.15K-436.090690
HF Energy-434.325056
Nuclear repulsion energy414.653595
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3281 3115 5.84      
2 A1 3254 3089 3.69      
3 A1 3222 3059 0.23      
4 A1 1652 1568 0.19      
5 A1 1523 1446 2.75      
6 A1 1411 1340 151.16      
7 A1 1210 1149 1.19      
8 A1 1151 1092 8.02      
9 A1 1053 1000 6.70      
10 A1 1024 972 0.38      
11 A1 870 826 36.59      
12 A1 696 661 12.09      
13 A1 404 384 1.96      
14 A2 1007 956 0.00      
15 A2 871 827 0.00      
16 A2 423 401 0.00      
17 A2 50 47 0.00      
18 B1 1017 965 0.29      
19 B1 966 917 5.14      
20 B1 834 792 12.73      
21 B1 735 698 82.48      
22 B1 705 669 0.55      
23 B1 458 434 0.69      
24 B1 176 167 0.59      
25 B2 3280 3114 1.42      
26 B2 3239 3075 6.73      
27 B2 1770 1680 104.90      
28 B2 1664 1580 13.90      
29 B2 1499 1423 1.03      
30 B2 1491 1416 7.65      
31 B2 1332 1265 0.40      
32 B2 1199 1139 0.02      
33 B2 1109 1053 6.93      
34 B2 618 586 0.00      
35 B2 529 502 0.49      
36 B2 260 247 0.67      

Unscaled Zero Point Vibrational Energy (zpe) 22990.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 21827.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.13375 0.04350 0.03282

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.246
C2 0.000 1.211 -0.420
C3 0.000 1.202 -1.806
C4 0.000 0.000 -2.500
C5 0.000 -1.202 -1.806
C6 0.000 -1.211 -0.420
N7 0.000 0.000 1.714
O8 0.000 -1.087 2.263
O9 0.000 1.087 2.263
H10 0.000 2.125 0.143
H11 0.000 2.135 -2.342
H12 0.000 0.000 -3.576
H13 0.000 -2.135 -2.342
H14 0.000 -2.125 0.143

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 O8 O9 H10 H11 H12 H13 H14
C11.38172.37852.74622.37851.38171.46722.29082.29082.12783.35563.82273.35562.1278
C21.38171.38632.40692.78302.42132.45283.53192.68521.07382.13353.38093.85883.3831
C32.37851.38631.38842.40492.78303.71924.66864.07032.15621.07592.14023.38023.8564
C42.74622.40691.38841.38842.40694.21354.88514.88513.39132.14071.07642.14073.3913
C52.37852.78302.40491.38841.38633.71924.07034.66863.85643.38022.14021.07592.1562
C61.38172.42132.78302.40691.38632.45282.68523.53193.38313.85883.38092.13351.0738
N71.46722.45283.71924.21353.71922.45281.21791.21792.64274.58365.28994.58362.6427
O82.29083.53194.66864.88514.07032.68521.21792.17413.84885.62055.93944.72302.3605
O92.29082.68524.07034.88514.66863.53191.21792.17412.36054.72305.93945.62053.8488
H102.12781.07382.15623.39133.85643.38312.64273.84882.36052.48544.28364.93224.2506
H113.35562.13351.07592.14073.38023.85884.58365.62054.72302.48542.46584.26984.9322
H123.82273.38092.14021.07642.14023.38095.28995.93945.93944.28362.46582.46584.2836
H133.35563.85883.38022.14071.07592.13354.58364.72305.62054.93224.26982.46582.4854
H142.12783.38313.85643.39132.15621.07382.64272.36053.84884.25064.93224.28362.4854

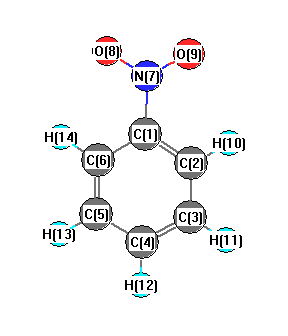
picture of Nitrobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 118.475 C1 C2 H10 119.594
C1 C6 C5 118.475 C1 C6 H14 119.594
C1 N7 O8 116.805 C1 N7 O9 116.805
C2 C1 C6 122.372 C2 C1 N7 118.814
C2 C3 C4 120.332 C2 C3 H11 119.581
C3 C2 H10 121.931 C3 C4 C5 120.015
C3 C4 H12 119.993 C4 C3 H11 120.087
C4 C5 C6 120.332 C4 C5 H13 120.087
C5 C4 H12 119.993 C5 C6 H14 121.931
C6 C1 N7 118.814 C6 C5 H13 119.581
O8 N7 O9 126.391
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability