return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NHCHNH2 (aminomethanimine)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-149.641068
Energy at 298.15K-149.646528
HF Energy-149.109692
Nuclear repulsion energy72.039353
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3750 3552 23.85      
2 A 3621 3429 20.73      
3 A 3570 3381 2.55      
4 A 3107 2942 57.03      
5 A 1762 1669 217.98      
6 A 1684 1595 30.40      
7 A 1441 1365 26.50      
8 A 1373 1300 21.67      
9 A 1153 1092 78.51      
10 A 1122 1063 40.88      
11 A 1066 1010 2.84      
12 A 839 795 25.06      
13 A 668 632 372.02      
14 A 565 535 6.18      
15 A 441 418 98.31      

Unscaled Zero Point Vibrational Energy (zpe) 13080.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12388.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
2.19119 0.36025 0.31180

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
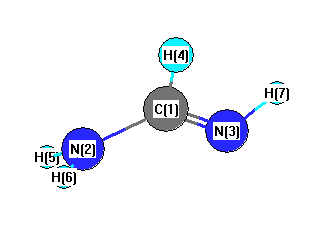
C1 -0.121 0.398 -0.002
N2 1.141 -0.148 -0.080
N3 -1.170 -0.336 0.018
H4 -0.117 1.494 -0.017
H5 1.884 0.382 0.342
H6 1.161 -1.136 0.123
H7 -1.997 0.253 -0.001

Atom - Atom Distances (Å)
  C1 N2 N3 H4 H5 H6 H7
C11.37751.28001.09612.03502.00311.8816
N21.37752.32082.06951.00581.00893.1648
N31.28002.32082.11113.15412.46681.0154
H41.09612.06952.11112.31782.92732.2529
H52.03501.00583.15412.31781.69533.8988
H62.00311.00892.46682.92731.69533.4523
H71.88163.16481.01542.25293.89883.4523

picture of aminomethanimine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H5 116.400 C1 N2 H6 113.248
C1 N3 H7 109.582 N2 C1 N3 121.644
N2 C1 H4 113.077 N3 C1 H4 125.195
H5 N2 H6 114.596
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-311G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.224      
2 N -0.802      
3 N -0.622      
4 H 0.193      
5 H 0.351      
6 H 0.366      
7 H 0.290      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000