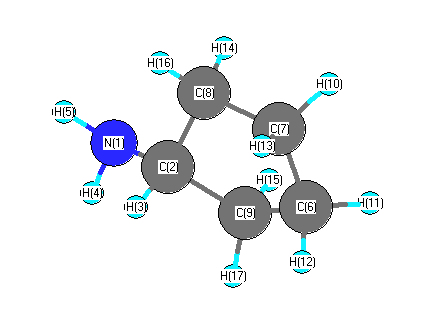
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3735 |
3378 |
1.70 |
|
|
|
| 2 |
A |
3248 |
2938 |
168.70 |
|
|
|
| 3 |
A |
3223 |
2914 |
36.87 |
|
|
|
| 4 |
A |
3207 |
2900 |
24.61 |
|
|
|
| 5 |
A |
3186 |
2882 |
61.46 |
|
|
|
| 6 |
A |
3084 |
2789 |
83.30 |
|
|
|
| 7 |
A |
1855 |
1677 |
43.61 |
|
|
|
| 8 |
A |
1662 |
1503 |
2.04 |
|
|
|
| 9 |
A |
1635 |
1479 |
3.62 |
|
|
|
| 10 |
A |
1552 |
1403 |
22.03 |
|
|
|
| 11 |
A |
1467 |
1327 |
0.64 |
|
|
|
| 12 |
A |
1438 |
1300 |
0.37 |
|
|
|
| 13 |
A |
1360 |
1230 |
1.78 |
|
|
|
| 14 |
A |
1317 |
1191 |
5.39 |
|
|
|
| 15 |
A |
1204 |
1089 |
20.61 |
|
|
|
| 16 |
A |
1124 |
1017 |
1.81 |
|
|
|
| 17 |
A |
1071 |
969 |
23.50 |
|
|
|
| 18 |
A |
987 |
892 |
89.38 |
|
|
|
| 19 |
A |
950 |
859 |
16.16 |
|
|
|
| 20 |
A |
926 |
837 |
22.77 |
|
|
|
| 21 |
A |
817 |
738 |
0.45 |
|
|
|
| 22 |
A |
576 |
521 |
3.28 |
|
|
|
| 23 |
A |
496 |
449 |
10.44 |
|
|
|
| 24 |
A |
196 |
177 |
1.21 |
|
|
|
| 25 |
A |
3804 |
3440 |
0.07 |
|
|
|
| 26 |
A |
3228 |
2919 |
8.70 |
|
|
|
| 27 |
A |
3209 |
2902 |
69.92 |
|
|
|
| 28 |
A |
3188 |
2883 |
81.93 |
|
|
|
| 29 |
A |
3179 |
2876 |
9.68 |
|
|
|
| 30 |
A |
1634 |
1478 |
2.14 |
|
|
|
| 31 |
A |
1619 |
1465 |
0.10 |
|
|
|
| 32 |
A |
1534 |
1387 |
2.32 |
|
|
|
| 33 |
A |
1467 |
1327 |
0.07 |
|
|
|
| 34 |
A |
1429 |
1292 |
0.11 |
|
|
|
| 35 |
A |
1394 |
1260 |
0.82 |
|
|
|
| 36 |
A |
1337 |
1209 |
0.01 |
|
|
|
| 37 |
A |
1314 |
1189 |
0.79 |
|
|
|
| 38 |
A |
1120 |
1013 |
0.00 |
|
|
|
| 39 |
A |
1061 |
959 |
0.01 |
|
|
|
| 40 |
A |
1009 |
912 |
3.19 |
|
|
|
| 41 |
A |
885 |
800 |
0.01 |
|
|
|
| 42 |
A |
678 |
613 |
0.00 |
|
|
|
| 43 |
A |
374 |
338 |
11.66 |
|
|
|
| 44 |
A |
263 |
238 |
40.79 |
|
|
|
| 45 |
A |
43 |
39 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37040.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 33499.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.841 |
-1.132 |
|
-1.131 |
| 2 |
C |
-0.008 |
0.581 |
|
0.544 |
| 3 |
H |
0.174 |
-0.123 |
|
-0.089 |
| 4 |
H |
0.329 |
0.385 |
|
0.389 |
| 5 |
H |
0.329 |
0.385 |
|
0.389 |
| 6 |
C |
-0.424 |
-0.014 |
|
-0.079 |
| 7 |
C |
-0.424 |
-0.014 |
|
-0.079 |
| 8 |
C |
-0.419 |
-0.023 |
|
-0.076 |
| 9 |
C |
-0.419 |
-0.023 |
|
-0.076 |
| 10 |
H |
0.216 |
-0.000 |
|
0.029 |
| 11 |
H |
0.216 |
-0.000 |
|
0.029 |
| 12 |
H |
0.209 |
-0.007 |
|
0.023 |
| 13 |
H |
0.209 |
-0.007 |
|
0.023 |
| 14 |
H |
0.221 |
0.016 |
|
0.047 |
| 15 |
H |
0.221 |
0.016 |
|
0.047 |
| 16 |
H |
0.205 |
-0.019 |
|
0.005 |
| 17 |
H |
0.205 |
-0.019 |
|
0.005 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.169 |
0.698 |
0.000 |
1.361 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.169 |
0.684 |
0.000 |
1.355 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.694 |
1.785 |
0.000 |
| y |
1.785 |
-35.775 |
0.000 |
| z |
0.000 |
0.000 |
-38.478 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.568 |
1.785 |
0.000 |
| y |
1.785 |
4.811 |
0.000 |
| z |
0.000 |
0.000 |
0.757 |
|
| Polar |
|---|
| 3z2-r2 | 1.514 |
|---|
| x2-y2 | -6.919 |
|---|
| xy | 1.785 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.882 |
-0.649 |
0.000 |
| y |
-0.649 |
8.998 |
0.000 |
| z |
0.000 |
0.000 |
8.731 |
<r2> (average value of r
2) Å
2
| <r2> |
170.976 |
| (<r2>)1/2 |
13.076 |