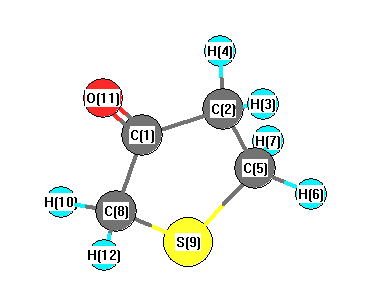
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3160 |
3016 |
7.79 |
|
|
|
| 2 |
A |
3152 |
3008 |
1.38 |
|
|
|
| 3 |
A |
3137 |
2994 |
11.57 |
|
|
|
| 4 |
A |
3087 |
2946 |
29.50 |
|
|
|
| 5 |
A |
3077 |
2937 |
17.37 |
|
|
|
| 6 |
A |
3068 |
2928 |
5.87 |
|
|
|
| 7 |
A |
1859 |
1775 |
240.22 |
|
|
|
| 8 |
A |
1510 |
1441 |
3.98 |
|
|
|
| 9 |
A |
1462 |
1395 |
6.69 |
|
|
|
| 10 |
A |
1459 |
1392 |
14.80 |
|
|
|
| 11 |
A |
1322 |
1262 |
10.36 |
|
|
|
| 12 |
A |
1316 |
1256 |
17.04 |
|
|
|
| 13 |
A |
1248 |
1191 |
28.73 |
|
|
|
| 14 |
A |
1241 |
1184 |
5.72 |
|
|
|
| 15 |
A |
1183 |
1129 |
4.12 |
|
|
|
| 16 |
A |
1158 |
1105 |
55.14 |
|
|
|
| 17 |
A |
1118 |
1067 |
6.29 |
|
|
|
| 18 |
A |
1015 |
969 |
9.57 |
|
|
|
| 19 |
A |
991 |
946 |
0.86 |
|
|
|
| 20 |
A |
886 |
845 |
11.50 |
|
|
|
| 21 |
A |
842 |
804 |
2.10 |
|
|
|
| 22 |
A |
801 |
764 |
0.76 |
|
|
|
| 23 |
A |
738 |
705 |
6.05 |
|
|
|
| 24 |
A |
691 |
660 |
0.91 |
|
|
|
| 25 |
A |
564 |
539 |
4.31 |
|
|
|
| 26 |
A |
493 |
471 |
5.00 |
|
|
|
| 27 |
A |
450 |
429 |
4.88 |
|
|
|
| 28 |
A |
432 |
412 |
4.22 |
|
|
|
| 29 |
A |
195 |
186 |
2.29 |
|
|
|
| 30 |
A |
70 |
66 |
11.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20861.9 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 19910.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.345 |
|
|
|
| 2 |
C |
-0.527 |
|
|
|
| 3 |
H |
0.262 |
|
|
|
| 4 |
H |
0.273 |
|
|
|
| 5 |
C |
-0.614 |
|
|
|
| 6 |
H |
0.264 |
|
|
|
| 7 |
H |
0.269 |
|
|
|
| 8 |
C |
-0.673 |
|
|
|
| 9 |
S |
0.128 |
|
|
|
| 10 |
H |
0.285 |
|
|
|
| 11 |
O |
-0.294 |
|
|
|
| 12 |
H |
0.280 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.028 |
1.336 |
0.418 |
1.736 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-53.811 |
1.227 |
0.352 |
| y |
1.227 |
-38.521 |
-0.021 |
| z |
0.352 |
-0.021 |
-42.239 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.431 |
1.227 |
0.352 |
| y |
1.227 |
9.503 |
-0.021 |
| z |
0.352 |
-0.021 |
3.927 |
|
| Polar |
|---|
| 3z2-r2 | 7.855 |
|---|
| x2-y2 | -15.289 |
|---|
| xy | 1.227 |
|---|
| xz | 0.352 |
|---|
| yz | -0.021 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
184.614 |
| (<r2>)1/2 |
13.587 |