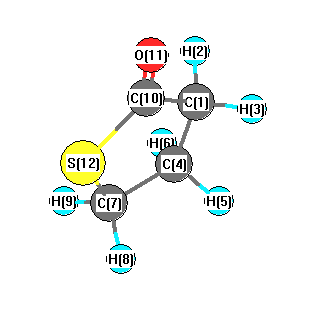
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3135 |
3018 |
14.83 |
|
|
|
| 2 |
A |
3123 |
3007 |
10.62 |
|
|
|
| 3 |
A |
3105 |
2989 |
22.55 |
|
|
|
| 4 |
A |
3068 |
2954 |
22.33 |
|
|
|
| 5 |
A |
3058 |
2944 |
28.04 |
|
|
|
| 6 |
A |
3041 |
2927 |
8.74 |
|
|
|
| 7 |
A |
1832 |
1763 |
393.92 |
|
|
|
| 8 |
A |
1508 |
1452 |
0.91 |
|
|
|
| 9 |
A |
1493 |
1438 |
8.27 |
|
|
|
| 10 |
A |
1466 |
1411 |
9.24 |
|
|
|
| 11 |
A |
1368 |
1317 |
0.09 |
|
|
|
| 12 |
A |
1322 |
1272 |
10.94 |
|
|
|
| 13 |
A |
1314 |
1265 |
3.15 |
|
|
|
| 14 |
A |
1259 |
1212 |
2.18 |
|
|
|
| 15 |
A |
1198 |
1153 |
2.04 |
|
|
|
| 16 |
A |
1128 |
1086 |
7.25 |
|
|
|
| 17 |
A |
1074 |
1034 |
54.80 |
|
|
|
| 18 |
A |
1037 |
999 |
3.58 |
|
|
|
| 19 |
A |
1034 |
995 |
57.24 |
|
|
|
| 20 |
A |
928 |
894 |
9.94 |
|
|
|
| 21 |
A |
876 |
844 |
9.80 |
|
|
|
| 22 |
A |
842 |
810 |
16.98 |
|
|
|
| 23 |
A |
700 |
674 |
2.29 |
|
|
|
| 24 |
A |
626 |
603 |
37.48 |
|
|
|
| 25 |
A |
601 |
579 |
16.06 |
|
|
|
| 26 |
A |
481 |
463 |
1.28 |
|
|
|
| 27 |
A |
475 |
457 |
1.93 |
|
|
|
| 28 |
A |
402 |
387 |
3.64 |
|
|
|
| 29 |
A |
233 |
224 |
1.35 |
|
|
|
| 30 |
A |
131 |
126 |
0.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20928.6 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 20148.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.493 |
|
|
|
| 2 |
H |
0.263 |
|
|
|
| 3 |
H |
0.258 |
|
|
|
| 4 |
C |
-0.465 |
|
|
|
| 5 |
H |
0.241 |
|
|
|
| 6 |
H |
0.250 |
|
|
|
| 7 |
C |
-0.605 |
|
|
|
| 8 |
H |
0.259 |
|
|
|
| 9 |
H |
0.269 |
|
|
|
| 10 |
C |
0.176 |
|
|
|
| 11 |
O |
-0.298 |
|
|
|
| 12 |
S |
0.145 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.707 |
1.825 |
0.392 |
4.151 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.905 |
-0.400 |
-0.041 |
| y |
-0.400 |
-42.616 |
0.156 |
| z |
-0.041 |
0.156 |
-42.315 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.439 |
-0.400 |
-0.041 |
| y |
-0.400 |
1.994 |
0.156 |
| z |
-0.041 |
0.156 |
2.445 |
|
| Polar |
|---|
| 3z2-r2 | 4.891 |
|---|
| x2-y2 | -4.289 |
|---|
| xy | -0.400 |
|---|
| xz | -0.041 |
|---|
| yz | 0.156 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
178.331 |
| (<r2>)1/2 |
13.354 |