return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2CH3 (Ethylbenzene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-310.050110
Energy at 298.15K-310.060237
HF Energy-308.825719
Nuclear repulsion energy340.057782
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3229 3058 17.09      
2 A' 3207 3038 3.05      
3 A' 3190 3021 9.25      
4 A' 3166 2999 37.90      
5 A' 3082 2919 29.23      
6 A' 3076 2914 27.82      
7 A' 1662 1574 3.09      
8 A' 1545 1464 12.26      
9 A' 1537 1456 8.12      
10 A' 1525 1445 1.29      
11 A' 1442 1366 5.50      
12 A' 1376 1303 1.43      
13 A' 1247 1181 0.33      
14 A' 1214 1150 0.11      
15 A' 1105 1047 9.22      
16 A' 1057 1001 2.78      
17 A' 1016 962 0.54      
18 A' 998 945 3.39      
19 A' 864 818 0.04      
20 A' 847 803 0.69      
21 A' 787 745 0.89      
22 A' 724 686 78.01      
23 A' 582 551 15.40      
24 A' 501 475 4.86      
25 A' 442 419 1.52      
26 A' 302 286 0.15      
27 A' 132 125 0.99      
28 A" 3216 3046 31.22      
29 A" 3191 3023 8.24      
30 A" 3174 3006 24.84      
31 A" 3132 2966 7.81      
32 A" 1640 1553 0.44      
33 A" 1535 1454 12.19      
34 A" 1491 1413 8.07      
35 A" 1462 1385 0.66      
36 A" 1361 1289 0.40      
37 A" 1297 1229 0.06      
38 A" 1194 1131 0.03      
39 A" 1134 1074 3.06      
40 A" 1071 1015 3.46      
41 A" 860 815 0.02      
42 A" 819 776 0.11      
43 A" 798 756 0.67      
44 A" 630 597 0.06      
45 A" 392 371 0.00      
46 A" 360 341 0.09      
47 A" 227 215 0.03      
48 A" 40 38 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 34441.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 32619.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.14986 0.04921 0.04080

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.250 0.505 0.000
C2 -0.244 -0.213 1.204
C3 -0.244 -1.610 1.208
C4 -0.240 -2.313 0.000
C5 -0.244 -1.610 -1.208
C6 -0.244 -0.213 -1.204
C7 -0.181 2.010 0.000
C8 1.269 2.505 0.000
H9 -0.253 0.329 2.148
H10 -0.247 -2.149 2.152
H11 -0.242 -3.399 0.000
H12 -0.247 -2.149 -2.152
H13 -0.253 0.329 -2.148
H14 -0.705 2.399 -0.879
H15 -0.705 2.399 0.879
H16 1.798 2.138 0.883
H17 1.798 2.138 -0.883
H18 1.317 3.597 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.40192.43552.81792.43551.40191.50612.51072.15573.41653.90453.41652.15572.13732.13732.76432.76433.4661
C21.40191.39702.42082.78742.40842.52843.33491.08872.15543.40653.87433.39623.37282.67233.13073.74864.2895
C32.43551.39701.39752.41562.78743.81614.54692.15501.08702.15903.40253.87604.54324.04894.28074.75275.5682
C42.81792.42081.39751.39752.42084.32285.04813.40522.15801.08652.15803.40524.81594.81594.97454.97456.1114
C52.43552.78742.41561.39751.39703.81614.54693.87603.40252.15901.08702.15504.04894.54324.75274.28075.5682
C61.40192.40842.78742.42081.39702.52843.33493.39623.87433.40652.15541.08872.67233.37283.74863.13074.2895
C71.50612.52843.81614.32283.81612.52841.53182.72864.68255.40934.68252.72861.09521.09522.17112.17112.1825
C82.51073.33494.54695.04814.54693.33491.53183.41515.34606.09415.34603.41512.16322.16321.09261.09261.0934
H92.15571.08872.15503.40523.87603.39622.72863.41512.47774.30314.96304.29693.69562.47023.01374.08254.2141
H103.41652.15541.08702.15803.40253.87434.68255.34602.47772.48884.30354.96305.48464.74494.91655.63636.3315
H113.90453.40652.15901.08652.15903.40655.40936.09414.30312.48882.48884.30315.88325.88325.96715.96717.1678
H123.41653.87433.40252.15801.08702.15544.68255.34604.96304.30352.48882.47774.74495.48465.63634.91656.3315
H132.15573.39623.87603.40522.15501.08872.72863.41514.29694.96304.30312.47772.47023.69564.08253.01374.2141
H142.13733.37284.54324.81594.04892.67231.09522.16323.69565.48465.88324.74492.47021.75833.07232.51712.5091
H152.13732.67234.04894.81594.54323.37281.09522.16322.47024.74495.88325.48463.69561.75832.51713.07232.5091
H162.76433.13074.28074.97454.75273.74862.17111.09263.01374.91655.96715.63634.08253.07232.51711.76501.7716
H172.76433.74864.75274.97454.28073.13072.17111.09264.08255.63635.96714.91653.01372.51713.07231.76501.7716
H183.46614.28955.56826.11145.56824.28952.18251.09344.21416.33157.16786.33154.21412.50912.50911.77161.7716

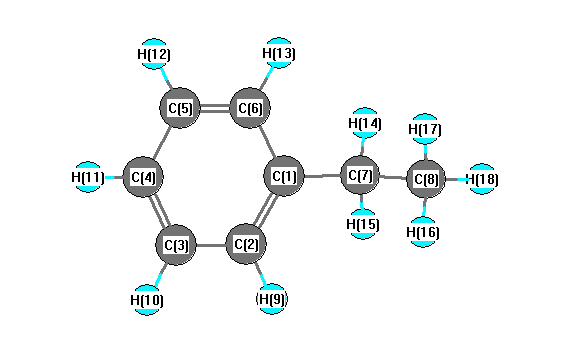
picture of Ethylbenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.948 C1 C2 H9 119.351
C1 C6 C5 120.948 C1 C6 H13 119.351
C1 C7 C8 111.473 C1 C7 H14 109.486
C1 C7 H15 109.486 C2 C1 C6 118.398
C2 C1 C7 120.751 C2 C3 C4 120.054
C2 C3 H10 119.872 C3 C2 H9 119.696
C3 C4 C5 119.594 C3 C4 H11 120.202
C4 C3 H10 120.074 C4 C5 C6 120.054
C4 C5 H12 120.074 C5 C4 H11 120.202
C5 C6 H13 119.696 C6 C1 C7 120.751
C6 C5 H12 119.872 C7 C8 H16 110.522
C7 C8 H17 110.522 C7 C8 H18 111.383
C8 C7 H14 109.748 C8 C7 H15 109.748
H14 C7 H15 106.785 H16 C8 H17 107.747
H16 C8 H18 108.274 H17 C8 H18 108.274
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability