return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CN (phenyl cyanide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-323.727547
Energy at 298.15K-323.732175
Nuclear repulsion energy298.768737
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3240 3068 5.57      
2 A1 3227 3057 6.44      
3 A1 3212 3042 0.00      
4 A1 2192 2077 0.00      
5 A1 1658 1570 0.26      
6 A1 1531 1450 12.80      
7 A1 1228 1163 0.93      
8 A1 1217 1152 0.23      
9 A1 1055 999 3.96      
10 A1 1015 961 0.56      
11 A1 771 730 1.14      
12 A1 462 437 0.00      
13 A2 883 836 0.00      
14 A2 822 779 0.00      
15 A2 388 367 0.00      
16 B1 879 832 0.33      
17 B1 866 820 1.65      
18 B1 730 691 93.09      
19 B1 561 532 17.99      
20 B1 475 450 2.53      
21 B1 368 348 0.39      
22 B1 141 133 1.18      
23 B2 3235 3064 10.00      
24 B2 3221 3051 1.72      
25 B2 1632 1546 3.40      
26 B2 1489 1410 8.69      
27 B2 1449 1373 0.05      
28 B2 1351 1279 0.01      
29 B2 1200 1137 0.00      
30 B2 1111 1052 4.26      
31 B2 634 600 0.16      
32 B2 554 524 0.08      
33 B2 162 153 3.82      

Unscaled Zero Point Vibrational Energy (zpe) 21478.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 20342.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.18828 0.05128 0.04030

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 3.217
C2 0.000 0.000 2.041
C3 0.000 0.000 0.607
C4 0.000 1.219 -0.090
C5 0.000 -1.219 -0.090
C6 0.000 1.211 -1.485
C7 0.000 -1.211 -1.485
C8 0.000 0.000 -2.184
H9 0.000 2.154 0.462
H10 0.000 -2.154 0.462
H11 0.000 2.152 -2.027
H12 0.000 -2.152 -2.027
H13 0.000 0.000 -3.270

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13
N11.17612.61053.52463.52464.85544.85545.40093.49763.49765.66835.66836.4872
C21.17611.43442.45502.45503.72813.72814.22482.67112.67114.60204.60205.3111
C32.61051.43441.40391.40392.41682.41682.79042.15912.15913.40103.40103.8767
C43.52462.45501.40392.43771.39492.80172.42271.08603.41792.14993.88783.4056
C53.52462.45501.40392.43772.80171.39492.42273.41791.08603.88782.14993.4056
C64.85543.72812.41681.39492.80172.42181.39812.16313.88761.08623.40662.1571
C74.85543.72812.41682.80171.39492.42181.39813.88762.16313.40661.08622.1571
C85.40094.22482.79042.42272.42271.39811.39813.41163.41162.15802.15801.0863
H93.49762.67112.15911.08603.41792.16313.88763.41164.30852.48844.97384.3089
H103.49762.67112.15913.41791.08603.88762.16313.41164.30854.97382.48844.3089
H115.66834.60203.40102.14993.88781.08623.40662.15802.48844.97384.30462.4857
H125.66834.60203.40103.88782.14993.40661.08622.15804.97382.48844.30462.4857
H136.48725.31113.87673.40563.40562.15712.15711.08634.30894.30892.48572.4857

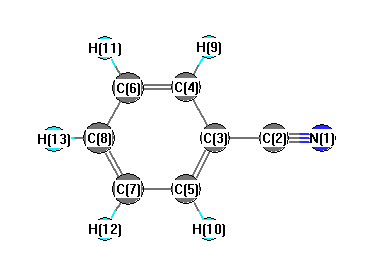
picture of phenyl cyanide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 180.000 C2 C3 C4 119.751
C2 C3 C5 119.751 C3 C4 C6 119.425
C3 C4 H9 119.715 C3 C5 C7 119.425
C3 C5 H10 119.715 C4 C3 C5 120.497
C4 C6 C8 120.317 C4 C6 H11 119.592
C5 C7 C8 120.317 C5 C7 H12 119.592
C6 C4 H9 120.860 C6 C8 C7 120.018
C6 C8 H13 119.991 C7 C5 H10 120.860
C7 C8 H13 119.991 C8 C6 H11 120.090
C8 C7 H12 120.090
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability