return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5OCH3 (Anisole)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-345.922071
Energy at 298.15K-345.930743
HF Energy-344.647308
Nuclear repulsion energy345.637475
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3248 3076 5.19      
2 A' 3235 3064 11.34      
3 A' 3228 3057 16.47      
4 A' 3213 3043 10.18      
5 A' 3210 3040 20.42      
6 A' 3205 3035 2.92      
7 A' 3061 2899 44.91      
8 A' 1667 1579 70.01      
9 A' 1644 1557 15.94      
10 A' 1554 1472 0.13      
11 A' 1545 1463 120.04      
12 A' 1512 1432 30.32      
13 A' 1496 1417 0.31      
14 A' 1462 1385 11.51      
15 A' 1351 1279 1.63      
16 A' 1310 1240 218.30      
17 A' 1231 1166 7.39      
18 A' 1212 1148 8.68      
19 A' 1191 1128 1.80      
20 A' 1113 1055 15.36      
21 A' 1095 1037 40.47      
22 A' 1052 996 2.08      
23 A' 1013 959 0.85      
24 A' 803 761 16.34      
25 A' 623 590 0.40      
26 A' 561 532 5.40      
27 A' 449 425 0.91      
28 A' 269 255 2.64      
29 A" 3141 2975 37.86      
30 A" 1535 1454 8.51      
31 A" 1200 1137 0.65      
32 A" 866 820 0.04      
33 A" 857 812 1.78      
34 A" 826 783 4.19      
35 A" 800 758 0.05      
36 A" 732 693 104.79      
37 A" 537 509 8.86      
38 A" 424 402 2.99      
39 A" 400 379 0.04      
40 A" 289 273 0.14      
41 A" 210 199 0.22      
42 A" 83 78 4.94      

Unscaled Zero Point Vibrational Energy (zpe) 29226.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 27680.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.16755 0.05240 0.04023

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.536 0.000
C2 0.936 -0.506 0.000
C3 0.488 -1.833 0.000
C4 -0.873 -2.132 0.000
C5 -1.802 -1.084 0.000
C6 -1.371 0.238 0.000
O7 0.307 1.864 0.000
C8 1.684 2.199 0.000
H9 2.002 -0.310 0.000
H10 1.221 -2.636 0.000
H11 -1.209 -3.164 0.000
H12 -2.868 -1.298 0.000
H13 -2.074 1.066 0.000
H14 1.721 3.286 0.000
H15 2.188 1.817 0.894
H16 2.188 1.817 -0.894

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 C8 H9 H10 H11 H12 H13 H14 H15 H16
C11.40072.41962.80712.42361.40291.36262.36622.17313.39933.89303.40422.14063.24422.68792.6879
C21.40071.40082.43182.79822.42382.45172.80641.08392.14903.41553.88503.39573.87272.78622.7862
C32.41961.40081.39352.40992.78383.70174.20582.14771.08692.15683.39843.86975.26604.12474.1247
C42.80712.43181.39351.40022.42184.16615.02883.40342.15421.08602.16183.41616.00695.07525.0752
C52.42362.79822.40991.40021.39123.62504.78853.88193.39852.16301.08682.16775.61395.01345.0134
C61.40292.42382.78382.42181.39122.33633.62963.41683.87073.40662.14511.08614.34183.99423.9942
O71.36262.45173.70174.16613.62502.33631.41652.75574.59175.25174.48102.51132.00562.08232.0823
C82.36622.80644.20585.02884.78853.62961.41652.52854.85696.09355.73973.92461.08791.09461.0946
H92.17311.08392.14773.40343.88193.41682.75572.52852.45384.29624.96864.30163.60662.31442.3144
H103.39932.14901.08692.15423.39853.87074.59174.85692.45382.48714.30234.95655.94324.64364.6436
H113.89303.41552.15681.08602.16303.40665.25176.09354.29622.48712.49664.31827.08486.09506.0950
H123.40423.88503.39842.16181.08682.14514.48105.73974.96864.30232.49662.49426.48666.00496.0049
H132.14063.39573.86973.41612.16771.08612.51133.92464.30164.95654.31822.49424.39694.41844.4184
H143.24423.87275.26606.00695.61394.34182.00561.08793.60665.94327.08486.48664.39691.78151.7815
H152.68792.78624.12475.07525.01343.99422.08231.09462.31444.64366.09506.00494.41841.78151.7874
H162.68792.78624.12475.07525.01343.99422.08231.09462.31444.64366.09506.00494.41841.78151.7874

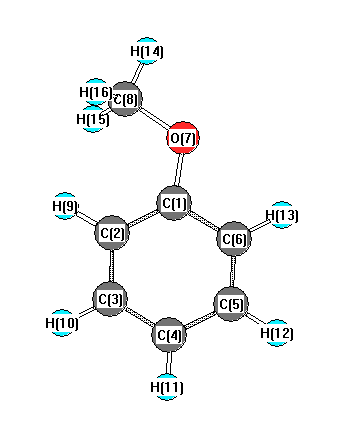
picture of Anisole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.459 C1 C2 H9 121.473
C1 C6 C5 120.314 C1 C6 H13 118.080
C1 O7 C8 116.720 C2 C1 C6 119.658
C2 C1 O7 125.047 C2 C3 C4 120.980
C2 C3 H10 118.964 C3 C2 H9 119.067
C3 C4 C5 119.226 C3 C4 H11 120.380
C4 C3 H10 120.056 C4 C5 C6 120.363
C4 C5 H12 120.223 C5 C4 H11 120.394
C5 C6 H13 121.606 C6 C1 O7 115.295
C6 C5 H12 119.413 O7 C8 H14 105.673
O7 C8 H15 111.391 O7 C8 H16 111.391
H14 C8 H15 109.419 H14 C8 H16 109.419
H15 C8 H16 109.462
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-311G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.346      
2 C -0.319      
3 C -0.200      
4 C -0.244      
5 C -0.196      
6 C -0.268      
7 O -0.468      
8 C -0.413      
9 H 0.235      
10 H 0.217      
11 H 0.214      
12 H 0.219      
13 H 0.230      
14 H 0.229      
15 H 0.209      
16 H 0.209      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000