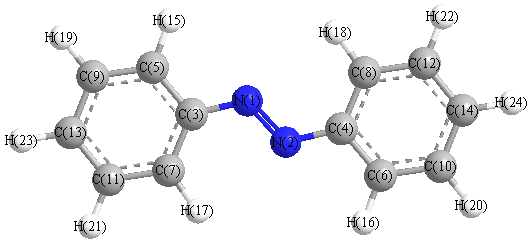
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3236 |
3106 |
0.00 |
|
|
|
| 2 |
Ag |
3224 |
3094 |
0.00 |
|
|
|
| 3 |
Ag |
3214 |
3084 |
0.00 |
|
|
|
| 4 |
Ag |
3203 |
3073 |
0.00 |
|
|
|
| 5 |
Ag |
3192 |
3063 |
0.00 |
|
|
|
| 6 |
Ag |
1679 |
1611 |
0.00 |
|
|
|
| 7 |
Ag |
1660 |
1593 |
0.00 |
|
|
|
| 8 |
Ag |
1592 |
1528 |
0.00 |
|
|
|
| 9 |
Ag |
1521 |
1460 |
0.00 |
|
|
|
| 10 |
Ag |
1498 |
1437 |
0.00 |
|
|
|
| 11 |
Ag |
1388 |
1332 |
0.00 |
|
|
|
| 12 |
Ag |
1345 |
1291 |
0.00 |
|
|
|
| 13 |
Ag |
1226 |
1176 |
0.00 |
|
|
|
| 14 |
Ag |
1186 |
1138 |
0.00 |
|
|
|
| 15 |
Ag |
1177 |
1129 |
0.00 |
|
|
|
| 16 |
Ag |
1103 |
1059 |
0.00 |
|
|
|
| 17 |
Ag |
1052 |
1010 |
0.00 |
|
|
|
| 18 |
Ag |
1023 |
981 |
0.00 |
|
|
|
| 19 |
Ag |
949 |
911 |
0.00 |
|
|
|
| 20 |
Ag |
686 |
658 |
0.00 |
|
|
|
| 21 |
Ag |
626 |
601 |
0.00 |
|
|
|
| 22 |
Ag |
309 |
297 |
0.00 |
|
|
|
| 23 |
Ag |
227 |
217 |
0.00 |
|
|
|
| 24 |
Au |
1013 |
972 |
1.67 |
|
|
|
| 25 |
Au |
991 |
951 |
0.00 |
|
|
|
| 26 |
Au |
958 |
919 |
10.19 |
|
|
|
| 27 |
Au |
863 |
828 |
0.02 |
|
|
|
| 28 |
Au |
805 |
772 |
66.73 |
|
|
|
| 29 |
Au |
708 |
679 |
98.63 |
|
|
|
| 30 |
Au |
562 |
539 |
17.26 |
|
|
|
| 31 |
Au |
419 |
402 |
0.01 |
|
|
|
| 32 |
Au |
309 |
297 |
0.51 |
|
|
|
| 33 |
Au |
65 |
62 |
1.58 |
|
|
|
| 34 |
Au |
23 |
22 |
0.02 |
|
|
|
| 35 |
Bg |
1013 |
972 |
0.00 |
|
|
|
| 36 |
Bg |
992 |
952 |
0.00 |
|
|
|
| 37 |
Bg |
953 |
915 |
0.00 |
|
|
|
| 38 |
Bg |
864 |
829 |
0.00 |
|
|
|
| 39 |
Bg |
785 |
753 |
0.00 |
|
|
|
| 40 |
Bg |
703 |
674 |
0.00 |
|
|
|
| 41 |
Bg |
492 |
472 |
0.00 |
|
|
|
| 42 |
Bg |
422 |
405 |
0.00 |
|
|
|
| 43 |
Bg |
260 |
249 |
0.00 |
|
|
|
| 44 |
Bg |
105 |
101 |
0.00 |
|
|
|
| 45 |
Bu |
3236 |
3106 |
11.10 |
|
|
|
| 46 |
Bu |
3224 |
3093 |
42.44 |
|
|
|
| 47 |
Bu |
3214 |
3084 |
53.75 |
|
|
|
| 48 |
Bu |
3203 |
3073 |
24.66 |
|
|
|
| 49 |
Bu |
3192 |
3063 |
2.61 |
|
|
|
| 50 |
Bu |
1672 |
1604 |
3.25 |
|
|
|
| 51 |
Bu |
1654 |
1587 |
5.00 |
|
|
|
| 52 |
Bu |
1535 |
1473 |
14.29 |
|
|
|
| 53 |
Bu |
1499 |
1438 |
16.06 |
|
|
|
| 54 |
Bu |
1385 |
1329 |
8.64 |
|
|
|
| 55 |
Bu |
1339 |
1285 |
1.60 |
|
|
|
| 56 |
Bu |
1282 |
1230 |
26.84 |
|
|
|
| 57 |
Bu |
1186 |
1138 |
0.27 |
|
|
|
| 58 |
Bu |
1180 |
1132 |
28.64 |
|
|
|
| 59 |
Bu |
1108 |
1063 |
17.31 |
|
|
|
| 60 |
Bu |
1051 |
1009 |
15.28 |
|
|
|
| 61 |
Bu |
1022 |
980 |
3.47 |
|
|
|
| 62 |
Bu |
849 |
814 |
0.93 |
|
|
|
| 63 |
Bu |
633 |
607 |
0.68 |
|
|
|
| 64 |
Bu |
549 |
526 |
5.38 |
|
|
|
| 65 |
Bu |
534 |
512 |
25.90 |
|
|
|
| 66 |
Bu |
87 |
83 |
1.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42122.8 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 40421.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.232 |
|
|
|
| 2 |
N |
-0.232 |
|
|
|
| 3 |
C |
0.126 |
|
|
|
| 4 |
C |
0.126 |
|
|
|
| 5 |
C |
-0.197 |
|
|
|
| 6 |
C |
-0.197 |
|
|
|
| 7 |
C |
-0.163 |
|
|
|
| 8 |
C |
-0.163 |
|
|
|
| 9 |
C |
-0.207 |
|
|
|
| 10 |
C |
-0.207 |
|
|
|
| 11 |
C |
-0.209 |
|
|
|
| 12 |
C |
-0.209 |
|
|
|
| 13 |
C |
-0.192 |
|
|
|
| 14 |
C |
-0.192 |
|
|
|
| 15 |
H |
0.216 |
|
|
|
| 16 |
H |
0.216 |
|
|
|
| 17 |
H |
0.229 |
|
|
|
| 18 |
H |
0.229 |
|
|
|
| 19 |
H |
0.209 |
|
|
|
| 20 |
H |
0.209 |
|
|
|
| 21 |
H |
0.210 |
|
|
|
| 22 |
H |
0.210 |
|
|
|
| 23 |
H |
0.209 |
|
|
|
| 24 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-65.592 |
5.137 |
0.000 |
| y |
5.137 |
-68.225 |
0.000 |
| z |
0.000 |
0.000 |
-87.166 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
12.103 |
5.137 |
0.000 |
| y |
5.137 |
8.153 |
0.000 |
| z |
0.000 |
0.000 |
-20.257 |
|
| Polar |
|---|
| 3z2-r2 | -40.514 |
|---|
| x2-y2 | 2.633 |
|---|
| xy | 5.137 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
34.586 |
10.822 |
0.000 |
| y |
10.822 |
30.270 |
0.000 |
| z |
0.000 |
0.000 |
8.461 |
<r2> (average value of r
2) Å
2
| <r2> |
1087.635 |
| (<r2>)1/2 |
32.979 |